Jump to
S1C2
Energy calculated at HSEh1PBE/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -216.989821 |
| Energy at 298.15K | |
| HF Energy | -216.989821 |
| Nuclear repulsion energy | 116.894358 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3253 |
3253 |
3.65 |
52.41 |
0.70 |
0.82 |
| 2 |
A' |
3167 |
3167 |
3.21 |
167.92 |
0.11 |
0.20 |
| 3 |
A' |
3153 |
3153 |
8.59 |
31.52 |
0.66 |
0.79 |
| 4 |
A' |
3025 |
3025 |
32.31 |
178.88 |
0.10 |
0.18 |
| 5 |
A' |
1729 |
1729 |
6.53 |
41.71 |
0.05 |
0.10 |
| 6 |
A' |
1487 |
1487 |
6.74 |
12.62 |
0.51 |
0.68 |
| 7 |
A' |
1444 |
1444 |
7.65 |
9.47 |
0.40 |
0.57 |
| 8 |
A' |
1409 |
1409 |
8.19 |
3.19 |
0.70 |
0.82 |
| 9 |
A' |
1314 |
1314 |
0.21 |
19.02 |
0.22 |
0.37 |
| 10 |
A' |
1145 |
1145 |
62.54 |
1.59 |
0.71 |
0.83 |
| 11 |
A' |
1015 |
1015 |
36.10 |
4.79 |
0.66 |
0.80 |
| 12 |
A' |
921 |
921 |
1.28 |
5.24 |
0.03 |
0.06 |
| 13 |
A' |
614 |
614 |
6.54 |
1.60 |
0.67 |
0.80 |
| 14 |
A' |
263 |
263 |
2.13 |
1.32 |
0.44 |
0.61 |
| 15 |
A" |
3061 |
3061 |
18.39 |
86.08 |
0.75 |
0.86 |
| 16 |
A" |
1268 |
1268 |
0.00 |
3.79 |
0.75 |
0.86 |
| 17 |
A" |
1046 |
1046 |
11.49 |
1.30 |
0.75 |
0.86 |
| 18 |
A" |
1023 |
1023 |
7.80 |
0.08 |
0.75 |
0.86 |
| 19 |
A" |
965 |
965 |
47.44 |
2.52 |
0.75 |
0.86 |
| 20 |
A" |
563 |
563 |
12.03 |
1.25 |
0.75 |
0.86 |
| 21 |
A" |
184 |
184 |
3.12 |
0.81 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16023.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16023.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.936 |
-0.208 |
0.000 |
| C2 |
0.000 |
0.947 |
0.000 |
| C3 |
1.320 |
0.850 |
0.000 |
| F4 |
-0.268 |
-1.412 |
0.000 |
| H5 |
1.944 |
1.734 |
0.000 |
| H6 |
1.811 |
-0.114 |
0.000 |
| H7 |
-0.484 |
1.920 |
0.000 |
| H8 |
-1.581 |
-0.184 |
0.885 |
| H9 |
-1.581 |
-0.184 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4866 | 2.4918 | 1.3776 | 3.4739 | 2.7487 | 2.1751 | 1.0957 | 1.0957 |
C2 | 1.4866 | | 1.3232 | 2.3742 | 2.0975 | 2.0988 | 1.0869 | 2.1364 | 2.1364 | C3 | 2.4918 | 1.3232 | | 2.7638 | 1.0820 | 1.0824 | 2.0973 | 3.2047 | 3.2047 | F4 | 1.3776 | 2.3742 | 2.7638 | | 3.8459 | 2.4504 | 3.3391 | 2.0043 | 2.0043 | H5 | 3.4739 | 2.0975 | 1.0820 | 3.8459 | | 1.8530 | 2.4358 | 4.1101 | 4.1101 | H6 | 2.7487 | 2.0988 | 1.0824 | 2.4504 | 1.8530 | | 3.0669 | 3.5064 | 3.5064 | H7 | 2.1751 | 1.0869 | 2.0973 | 3.3391 | 2.4358 | 3.0669 | | 2.5327 | 2.5327 | H8 | 1.0957 | 2.1364 | 3.2047 | 2.0043 | 4.1101 | 3.5064 | 2.5327 | | 1.7709 | H9 | 1.0957 | 2.1364 | 3.2047 | 2.0043 | 4.1101 | 3.5064 | 2.5327 | 1.7709 | |
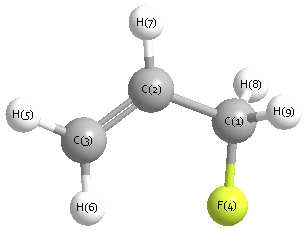 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.856 |
|
C1 |
C2 |
H7 |
114.491 |
| C2 |
C1 |
F4 |
111.922 |
|
C2 |
C1 |
H8 |
110.740 |
| C2 |
C1 |
H9 |
110.740 |
|
C2 |
C3 |
H5 |
121.071 |
| C2 |
C3 |
H6 |
121.165 |
|
C3 |
C2 |
H7 |
120.653 |
| F4 |
C1 |
H8 |
107.723 |
|
F4 |
C1 |
H9 |
107.723 |
| H5 |
C3 |
H6 |
117.764 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.277 |
|
|
|
| 2 |
C |
-0.123 |
|
|
|
| 3 |
C |
-0.308 |
|
|
|
| 4 |
F |
-0.462 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
H |
0.131 |
|
|
|
| 7 |
H |
0.104 |
|
|
|
| 8 |
H |
0.126 |
|
|
|
| 9 |
H |
0.126 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.922 |
1.489 |
0.000 |
1.752 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.223 |
-0.279 |
0.000 |
| y |
-0.279 |
-25.501 |
0.000 |
| z |
0.000 |
0.000 |
-25.363 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.209 |
-0.279 |
0.000 |
| y |
-0.279 |
-2.208 |
0.000 |
| z |
0.000 |
0.000 |
-2.001 |
|
| Polar |
|---|
| 3z2-r2 | -4.003 |
|---|
| x2-y2 | 4.278 |
|---|
| xy | -0.279 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.403 |
0.351 |
0.000 |
| y |
0.351 |
5.582 |
0.000 |
| z |
0.000 |
0.000 |
4.665 |
<r2> (average value of r
2) Å
2
| <r2> |
79.924 |
| (<r2>)1/2 |
8.940 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -216.989339 |
| Energy at 298.15K | |
| HF Energy | -216.989339 |
| Nuclear repulsion energy | 114.590311 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3236 |
3236 |
8.57 |
61.87 |
0.59 |
0.74 |
| 2 |
A |
3165 |
3165 |
4.75 |
118.42 |
0.22 |
0.36 |
| 3 |
A |
3146 |
3146 |
7.89 |
70.13 |
0.10 |
0.19 |
| 4 |
A |
3094 |
3094 |
18.08 |
65.34 |
0.69 |
0.82 |
| 5 |
A |
3040 |
3040 |
28.98 |
134.38 |
0.08 |
0.15 |
| 6 |
A |
1727 |
1727 |
0.70 |
43.41 |
0.07 |
0.13 |
| 7 |
A |
1497 |
1497 |
4.61 |
3.76 |
0.66 |
0.79 |
| 8 |
A |
1457 |
1457 |
19.23 |
9.45 |
0.52 |
0.68 |
| 9 |
A |
1386 |
1386 |
10.72 |
4.14 |
0.22 |
0.37 |
| 10 |
A |
1314 |
1314 |
0.10 |
14.32 |
0.30 |
0.46 |
| 11 |
A |
1269 |
1269 |
4.41 |
7.09 |
0.66 |
0.79 |
| 12 |
A |
1185 |
1185 |
2.73 |
1.12 |
0.60 |
0.75 |
| 13 |
A |
1060 |
1060 |
124.91 |
5.64 |
0.56 |
0.72 |
| 14 |
A |
1035 |
1035 |
25.83 |
1.85 |
0.72 |
0.84 |
| 15 |
A |
993 |
993 |
3.27 |
1.90 |
0.13 |
0.23 |
| 16 |
A |
976 |
976 |
53.85 |
2.20 |
0.54 |
0.70 |
| 17 |
A |
931 |
931 |
1.92 |
3.03 |
0.12 |
0.21 |
| 18 |
A |
662 |
662 |
7.97 |
1.46 |
0.30 |
0.46 |
| 19 |
A |
438 |
438 |
2.52 |
3.51 |
0.32 |
0.48 |
| 20 |
A |
331 |
331 |
7.18 |
1.15 |
0.73 |
0.84 |
| 21 |
A |
117 |
117 |
1.01 |
2.52 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16028.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16028.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.603 |
0.426 |
0.318 |
| C2 |
0.643 |
-0.380 |
0.241 |
| C3 |
1.793 |
0.093 |
-0.213 |
| F4 |
-1.638 |
-0.222 |
-0.343 |
| H5 |
2.693 |
-0.509 |
-0.223 |
| H6 |
1.878 |
1.108 |
-0.587 |
| H7 |
0.571 |
-1.401 |
0.605 |
| H8 |
-0.926 |
0.559 |
1.355 |
| H9 |
-0.471 |
1.408 |
-0.142 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4860 | 2.4769 | 1.3880 | 3.4684 | 2.7280 | 2.1903 | 1.0945 | 1.0925 |
C2 | 1.4860 | | 1.3242 | 2.3599 | 2.1058 | 2.1039 | 1.0862 | 2.1407 | 2.1412 | C3 | 2.4769 | 1.3242 | | 3.4481 | 1.0826 | 1.0848 | 2.0967 | 3.1730 | 2.6192 | F4 | 1.3880 | 2.3599 | 3.4481 | | 4.3419 | 3.7674 | 2.6771 | 2.0000 | 2.0146 | H5 | 3.4684 | 2.1058 | 1.0826 | 4.3419 | | 1.8466 | 2.4464 | 4.0897 | 3.7002 | H6 | 2.7280 | 2.1039 | 1.0848 | 3.7674 | 1.8466 | | 3.0701 | 3.4547 | 2.4099 | H7 | 2.1903 | 1.0862 | 2.0967 | 2.6771 | 2.4464 | 3.0701 | | 2.5774 | 3.0876 | H8 | 1.0945 | 2.1407 | 3.1730 | 2.0000 | 4.0897 | 3.4547 | 2.5774 | | 1.7803 | H9 | 1.0925 | 2.1412 | 2.6192 | 2.0146 | 3.7002 | 2.4099 | 3.0876 | 1.7803 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.525 |
|
C1 |
C2 |
H7 |
115.893 |
| C2 |
C1 |
F4 |
110.349 |
|
C2 |
C1 |
H8 |
111.204 |
| C2 |
C1 |
H9 |
111.372 |
|
C2 |
C3 |
H5 |
121.750 |
| C2 |
C3 |
H6 |
121.386 |
|
C3 |
C2 |
H7 |
120.570 |
| F4 |
C1 |
H8 |
106.750 |
|
F4 |
C1 |
H9 |
108.021 |
| H5 |
C3 |
H6 |
116.863 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.238 |
|
|
|
| 2 |
C |
-0.023 |
|
|
|
| 3 |
C |
-0.361 |
|
|
|
| 4 |
F |
-0.445 |
|
|
|
| 5 |
H |
0.131 |
|
|
|
| 6 |
H |
0.119 |
|
|
|
| 7 |
H |
0.105 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.112 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.520 |
0.741 |
0.859 |
1.897 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.794 |
-1.190 |
-2.040 |
| y |
-1.190 |
-22.575 |
-1.054 |
| z |
-2.040 |
-1.054 |
-24.993 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.010 |
-1.190 |
-2.040 |
| y |
-1.190 |
2.819 |
-1.054 |
| z |
-2.040 |
-1.054 |
-0.809 |
|
| Polar |
|---|
| 3z2-r2 | -1.618 |
|---|
| x2-y2 | -3.219 |
|---|
| xy | -1.190 |
|---|
| xz | -2.040 |
|---|
| yz | -1.054 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.423 |
0.249 |
-0.601 |
| y |
0.249 |
5.391 |
-0.204 |
| z |
-0.601 |
-0.204 |
4.973 |
<r2> (average value of r
2) Å
2
| <r2> |
88.657 |
| (<r2>)1/2 |
9.416 |
