Jump to
S1C2
Energy calculated at MP2/CEP-31G
| | hartrees |
|---|
| Energy at 0K | -21.422275 |
| Energy at 298.15K | -21.425912 |
| HF Energy | -21.270558 |
| Nuclear repulsion energy | 20.530939 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/CEP-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3556 |
3436 |
4.48 |
|
|
|
| 2 |
A' |
2510 |
2426 |
67.54 |
|
|
|
| 3 |
A' |
1663 |
1607 |
17.89 |
|
|
|
| 4 |
A' |
966 |
934 |
14.23 |
|
|
|
| 5 |
A' |
825 |
797 |
93.46 |
|
|
|
| 6 |
A' |
517 |
499 |
148.04 |
|
|
|
| 7 |
A" |
3705 |
3580 |
9.68 |
|
|
|
| 8 |
A" |
1098 |
1061 |
5.39 |
|
|
|
| 9 |
A" |
449 |
434 |
96.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7644.6 cm
-1
Scaled (by 0.9663) Zero Point Vibrational Energy (zpe) 7386.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/CEP-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.023 |
1.169 |
0.000 |
| S2 |
0.023 |
-0.657 |
0.000 |
| H3 |
-1.354 |
-0.804 |
0.000 |
| H4 |
0.418 |
1.565 |
0.859 |
| H5 |
0.418 |
1.565 |
-0.859 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.8263 | 2.4064 | 1.0247 | 1.0247 |
S2 | 1.8263 | | 1.3848 | 2.4143 | 2.4143 | H3 | 2.4064 | 1.3848 | | 3.0804 | 3.0804 | H4 | 1.0247 | 2.4143 | 3.0804 | | 1.7178 | H5 | 1.0247 | 2.4143 | 3.0804 | 1.7178 | |
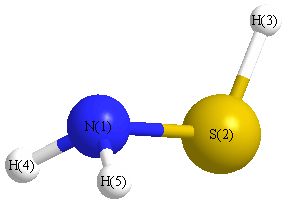 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
96.105 |
|
S2 |
N1 |
H4 |
112.687 |
| S2 |
N1 |
H5 |
112.687 |
|
H4 |
N1 |
H5 |
113.899 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/CEP-31G
| | hartrees |
|---|
| Energy at 0K | -21.422275 |
| Energy at 298.15K | -21.425914 |
| HF Energy | -21.270553 |
| Nuclear repulsion energy | 20.530992 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/CEP-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3555 |
3435 |
4.45 |
|
|
|
| 2 |
A' |
2510 |
2426 |
67.44 |
|
|
|
| 3 |
A' |
1663 |
1607 |
17.94 |
|
|
|
| 4 |
A' |
966 |
934 |
14.40 |
|
|
|
| 5 |
A' |
826 |
798 |
93.81 |
|
|
|
| 6 |
A' |
519 |
501 |
146.55 |
|
|
|
| 7 |
A" |
3704 |
3579 |
9.58 |
|
|
|
| 8 |
A" |
1099 |
1062 |
5.35 |
|
|
|
| 9 |
A" |
449 |
434 |
96.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7645.3 cm
-1
Scaled (by 0.9663) Zero Point Vibrational Energy (zpe) 7387.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/CEP-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.022 |
1.169 |
0.000 |
| S2 |
0.022 |
-0.657 |
0.000 |
| H3 |
-1.354 |
-0.804 |
0.000 |
| H4 |
0.419 |
1.564 |
0.859 |
| H5 |
0.419 |
1.564 |
-0.859 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.8263 | 2.4064 | 1.0248 | 1.0248 |
S2 | 1.8263 | | 1.3848 | 2.4141 | 2.4141 | H3 | 2.4064 | 1.3848 | | 3.0807 | 3.0807 | H4 | 1.0248 | 2.4141 | 3.0807 | | 1.7174 | H5 | 1.0248 | 2.4141 | 3.0807 | 1.7174 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
96.104 |
|
S2 |
N1 |
H4 |
112.669 |
| S2 |
N1 |
H5 |
112.669 |
|
H4 |
N1 |
H5 |
113.844 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
