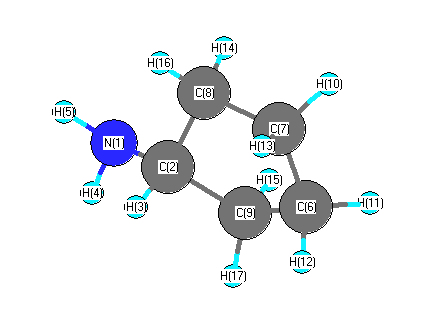
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3461 |
3342 |
4.33 |
|
|
|
| 2 |
A |
3116 |
3010 |
164.58 |
|
|
|
| 3 |
A |
3089 |
2983 |
14.82 |
|
|
|
| 4 |
A |
3064 |
2959 |
40.65 |
|
|
|
| 5 |
A |
3044 |
2939 |
42.51 |
|
|
|
| 6 |
A |
2939 |
2839 |
84.09 |
|
|
|
| 7 |
A |
1671 |
1614 |
27.45 |
|
|
|
| 8 |
A |
1523 |
1471 |
1.81 |
|
|
|
| 9 |
A |
1498 |
1447 |
4.48 |
|
|
|
| 10 |
A |
1397 |
1350 |
19.96 |
|
|
|
| 11 |
A |
1327 |
1281 |
0.46 |
|
|
|
| 12 |
A |
1303 |
1259 |
0.39 |
|
|
|
| 13 |
A |
1236 |
1194 |
2.43 |
|
|
|
| 14 |
A |
1191 |
1150 |
3.66 |
|
|
|
| 15 |
A |
1098 |
1061 |
12.99 |
|
|
|
| 16 |
A |
1044 |
1008 |
0.33 |
|
|
|
| 17 |
A |
980 |
946 |
25.91 |
|
|
|
| 18 |
A |
906 |
875 |
60.28 |
|
|
|
| 19 |
A |
882 |
852 |
23.06 |
|
|
|
| 20 |
A |
863 |
834 |
24.32 |
|
|
|
| 21 |
A |
744 |
718 |
3.69 |
|
|
|
| 22 |
A |
528 |
510 |
2.81 |
|
|
|
| 23 |
A |
452 |
436 |
11.47 |
|
|
|
| 24 |
A |
180 |
174 |
1.45 |
|
|
|
| 25 |
A |
3545 |
3424 |
0.54 |
|
|
|
| 26 |
A |
3096 |
2990 |
9.12 |
|
|
|
| 27 |
A |
3071 |
2966 |
21.43 |
|
|
|
| 28 |
A |
3049 |
2944 |
77.80 |
|
|
|
| 29 |
A |
3038 |
2934 |
32.44 |
|
|
|
| 30 |
A |
1498 |
1446 |
2.63 |
|
|
|
| 31 |
A |
1482 |
1431 |
0.03 |
|
|
|
| 32 |
A |
1386 |
1338 |
2.10 |
|
|
|
| 33 |
A |
1325 |
1280 |
0.21 |
|
|
|
| 34 |
A |
1290 |
1245 |
0.03 |
|
|
|
| 35 |
A |
1268 |
1225 |
0.77 |
|
|
|
| 36 |
A |
1212 |
1171 |
0.41 |
|
|
|
| 37 |
A |
1172 |
1132 |
0.78 |
|
|
|
| 38 |
A |
1029 |
994 |
0.06 |
|
|
|
| 39 |
A |
949 |
916 |
0.82 |
|
|
|
| 40 |
A |
928 |
896 |
5.09 |
|
|
|
| 41 |
A |
800 |
773 |
0.26 |
|
|
|
| 42 |
A |
616 |
595 |
0.09 |
|
|
|
| 43 |
A |
343 |
331 |
15.58 |
|
|
|
| 44 |
A |
246 |
238 |
35.16 |
|
|
|
| 45 |
A |
43 |
41 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34460.5 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 33278.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.523 |
|
|
|
| 2 |
C |
-0.057 |
|
|
|
| 3 |
H |
0.142 |
|
|
|
| 4 |
H |
0.254 |
|
|
|
| 5 |
H |
0.254 |
|
|
|
| 6 |
C |
-0.305 |
|
|
|
| 7 |
C |
-0.305 |
|
|
|
| 8 |
C |
-0.332 |
|
|
|
| 9 |
C |
-0.332 |
|
|
|
| 10 |
H |
0.170 |
|
|
|
| 11 |
H |
0.170 |
|
|
|
| 12 |
H |
0.125 |
|
|
|
| 13 |
H |
0.125 |
|
|
|
| 14 |
H |
0.130 |
|
|
|
| 15 |
H |
0.130 |
|
|
|
| 16 |
H |
0.177 |
|
|
|
| 17 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.229 |
0.690 |
0.000 |
1.409 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.633 |
1.902 |
0.000 |
| y |
1.902 |
-34.648 |
0.000 |
| z |
0.000 |
0.000 |
-37.576 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.522 |
1.902 |
0.000 |
| y |
1.902 |
4.956 |
0.000 |
| z |
0.000 |
0.000 |
0.565 |
|
| Polar |
|---|
| 3z2-r2 | 1.131 |
|---|
| x2-y2 | -6.985 |
|---|
| xy | 1.902 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.763 |
-1.087 |
0.000 |
| y |
-1.087 |
9.363 |
0.000 |
| z |
0.000 |
0.000 |
8.987 |
<r2> (average value of r
2) Å
2
| <r2> |
145.738 |
| (<r2>)1/2 |
12.072 |