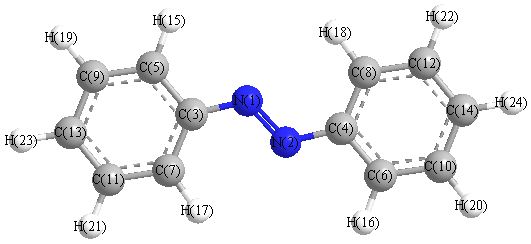
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3213 |
3103 |
0.00 |
|
|
|
| 2 |
Ag |
3197 |
3088 |
0.00 |
|
|
|
| 3 |
Ag |
3184 |
3075 |
0.00 |
|
|
|
| 4 |
Ag |
3168 |
3059 |
0.00 |
|
|
|
| 5 |
Ag |
3152 |
3044 |
0.00 |
|
|
|
| 6 |
Ag |
1647 |
1591 |
0.00 |
|
|
|
| 7 |
Ag |
1634 |
1578 |
0.00 |
|
|
|
| 8 |
Ag |
1536 |
1483 |
0.00 |
|
|
|
| 9 |
Ag |
1490 |
1439 |
0.00 |
|
|
|
| 10 |
Ag |
1464 |
1414 |
0.00 |
|
|
|
| 11 |
Ag |
1389 |
1342 |
0.00 |
|
|
|
| 12 |
Ag |
1314 |
1269 |
0.00 |
|
|
|
| 13 |
Ag |
1192 |
1151 |
0.00 |
|
|
|
| 14 |
Ag |
1158 |
1118 |
0.00 |
|
|
|
| 15 |
Ag |
1149 |
1110 |
0.00 |
|
|
|
| 16 |
Ag |
1085 |
1048 |
0.00 |
|
|
|
| 17 |
Ag |
1032 |
997 |
0.00 |
|
|
|
| 18 |
Ag |
987 |
953 |
0.00 |
|
|
|
| 19 |
Ag |
921 |
889 |
0.00 |
|
|
|
| 20 |
Ag |
658 |
636 |
0.00 |
|
|
|
| 21 |
Ag |
598 |
578 |
0.00 |
|
|
|
| 22 |
Ag |
300 |
290 |
0.00 |
|
|
|
| 23 |
Ag |
219 |
211 |
0.00 |
|
|
|
| 24 |
Au |
1016 |
981 |
0.47 |
|
|
|
| 25 |
Au |
1008 |
973 |
0.03 |
|
|
|
| 26 |
Au |
967 |
934 |
15.97 |
|
|
|
| 27 |
Au |
877 |
847 |
0.21 |
|
|
|
| 28 |
Au |
809 |
781 |
141.80 |
|
|
|
| 29 |
Au |
699 |
675 |
94.44 |
|
|
|
| 30 |
Au |
558 |
539 |
14.71 |
|
|
|
| 31 |
Au |
409 |
395 |
0.02 |
|
|
|
| 32 |
Au |
304 |
293 |
0.59 |
|
|
|
| 33 |
Au |
63 |
61 |
1.61 |
|
|
|
| 34 |
Au |
29 |
28 |
0.06 |
|
|
|
| 35 |
Bg |
1016 |
981 |
0.00 |
|
|
|
| 36 |
Bg |
1008 |
974 |
0.00 |
|
|
|
| 37 |
Bg |
961 |
928 |
0.00 |
|
|
|
| 38 |
Bg |
878 |
848 |
0.00 |
|
|
|
| 39 |
Bg |
792 |
765 |
0.00 |
|
|
|
| 40 |
Bg |
684 |
661 |
0.00 |
|
|
|
| 41 |
Bg |
492 |
475 |
0.00 |
|
|
|
| 42 |
Bg |
413 |
398 |
0.00 |
|
|
|
| 43 |
Bg |
265 |
256 |
0.00 |
|
|
|
| 44 |
Bg |
116 |
112 |
0.00 |
|
|
|
| 45 |
Bu |
3213 |
3103 |
16.30 |
|
|
|
| 46 |
Bu |
3197 |
3088 |
53.07 |
|
|
|
| 47 |
Bu |
3184 |
3075 |
50.73 |
|
|
|
| 48 |
Bu |
3168 |
3059 |
23.70 |
|
|
|
| 49 |
Bu |
3152 |
3044 |
2.62 |
|
|
|
| 50 |
Bu |
1646 |
1590 |
3.06 |
|
|
|
| 51 |
Bu |
1628 |
1572 |
4.13 |
|
|
|
| 52 |
Bu |
1505 |
1453 |
10.67 |
|
|
|
| 53 |
Bu |
1469 |
1419 |
11.94 |
|
|
|
| 54 |
Bu |
1385 |
1338 |
10.12 |
|
|
|
| 55 |
Bu |
1309 |
1264 |
0.91 |
|
|
|
| 56 |
Bu |
1244 |
1201 |
22.47 |
|
|
|
| 57 |
Bu |
1160 |
1120 |
12.25 |
|
|
|
| 58 |
Bu |
1157 |
1117 |
10.89 |
|
|
|
| 59 |
Bu |
1089 |
1052 |
16.25 |
|
|
|
| 60 |
Bu |
1033 |
997 |
12.88 |
|
|
|
| 61 |
Bu |
986 |
952 |
2.59 |
|
|
|
| 62 |
Bu |
818 |
790 |
0.72 |
|
|
|
| 63 |
Bu |
605 |
584 |
0.81 |
|
|
|
| 64 |
Bu |
527 |
509 |
0.83 |
|
|
|
| 65 |
Bu |
516 |
499 |
35.47 |
|
|
|
| 66 |
Bu |
84 |
81 |
1.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41562.6 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 40137.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
0.203 |
|
|
|
| 2 |
N |
0.203 |
|
|
|
| 3 |
C |
-0.451 |
|
|
|
| 4 |
C |
-0.451 |
|
|
|
| 5 |
C |
-0.138 |
|
|
|
| 6 |
C |
-0.138 |
|
|
|
| 7 |
C |
-0.277 |
|
|
|
| 8 |
C |
-0.277 |
|
|
|
| 9 |
C |
-0.318 |
|
|
|
| 10 |
C |
-0.318 |
|
|
|
| 11 |
C |
-0.286 |
|
|
|
| 12 |
C |
-0.286 |
|
|
|
| 13 |
C |
-0.260 |
|
|
|
| 14 |
C |
-0.260 |
|
|
|
| 15 |
H |
0.315 |
|
|
|
| 16 |
H |
0.315 |
|
|
|
| 17 |
H |
0.332 |
|
|
|
| 18 |
H |
0.332 |
|
|
|
| 19 |
H |
0.296 |
|
|
|
| 20 |
H |
0.296 |
|
|
|
| 21 |
H |
0.294 |
|
|
|
| 22 |
H |
0.294 |
|
|
|
| 23 |
H |
0.292 |
|
|
|
| 24 |
H |
0.292 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-64.107 |
5.147 |
0.000 |
| y |
5.147 |
-66.835 |
0.000 |
| z |
0.000 |
0.000 |
-86.803 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
12.713 |
5.147 |
0.000 |
| y |
5.147 |
8.619 |
0.000 |
| z |
0.000 |
0.000 |
-21.332 |
|
| Polar |
|---|
| 3z2-r2 | -42.664 |
|---|
| x2-y2 | 2.729 |
|---|
| xy | 5.147 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
36.261 |
12.123 |
0.000 |
| y |
12.123 |
31.362 |
0.000 |
| z |
0.000 |
0.000 |
8.607 |
<r2> (average value of r
2) Å
2
| <r2> |
827.988 |
| (<r2>)1/2 |
28.775 |