Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/CEP-31G*
Geometric Data calculated at B3LYP/CEP-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -38.943479 |
| Energy at 298.15K | -38.946777 |
| HF Energy | -38.943479 |
| Nuclear repulsion energy | 54.347946 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3792 |
3662 |
29.48 |
65.74 |
0.27 |
0.42 |
| 2 |
A |
3138 |
3031 |
8.66 |
79.28 |
0.41 |
0.58 |
| 3 |
A |
3048 |
2943 |
37.34 |
115.84 |
0.13 |
0.22 |
| 4 |
A |
2343 |
2262 |
0.48 |
60.88 |
0.20 |
0.33 |
| 5 |
A |
1480 |
1429 |
2.92 |
11.63 |
0.74 |
0.85 |
| 6 |
A |
1411 |
1362 |
56.28 |
7.60 |
0.75 |
0.86 |
| 7 |
A |
1361 |
1314 |
1.86 |
6.83 |
0.75 |
0.86 |
| 8 |
A |
1212 |
1171 |
22.21 |
4.55 |
0.66 |
0.80 |
| 9 |
A |
1070 |
1033 |
103.76 |
9.20 |
0.29 |
0.45 |
| 10 |
A |
981 |
947 |
22.87 |
0.37 |
0.40 |
0.58 |
| 11 |
A |
879 |
849 |
21.19 |
2.88 |
0.16 |
0.27 |
| 12 |
A |
563 |
544 |
3.73 |
2.60 |
0.24 |
0.38 |
| 13 |
A |
386 |
373 |
87.39 |
1.19 |
0.75 |
0.86 |
| 14 |
A |
303 |
292 |
77.15 |
5.15 |
0.75 |
0.86 |
| 15 |
A |
210 |
203 |
7.25 |
4.33 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11087.7 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 10707.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.588 |
0.598 |
0.039 |
| C2 |
0.838 |
0.117 |
-0.008 |
| O3 |
-1.532 |
-0.460 |
-0.112 |
| H4 |
-0.722 |
1.159 |
0.983 |
| H5 |
-0.747 |
1.288 |
-0.803 |
| H6 |
-1.409 |
-1.084 |
0.627 |
| N7 |
1.947 |
-0.282 |
-0.015 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.5053 | 1.4262 | 1.1064 | 1.1007 | 1.9623 | 2.6838 |
C2 | 1.5053 | | 2.4414 | 2.1216 | 2.1258 | 2.6257 | 1.1788 | O3 | 1.4262 | 2.4414 | | 2.1159 | 2.0372 | 0.9748 | 3.4851 | H4 | 1.1064 | 2.1216 | 2.1159 | | 1.7914 | 2.3733 | 3.1932 | H5 | 1.1007 | 2.1258 | 2.0372 | 1.7914 | | 2.8482 | 3.2169 | H6 | 1.9623 | 2.6257 | 0.9748 | 2.3733 | 2.8482 | | 3.5100 | N7 | 2.6838 | 1.1788 | 3.4851 | 3.1932 | 3.2169 | 3.5100 | |
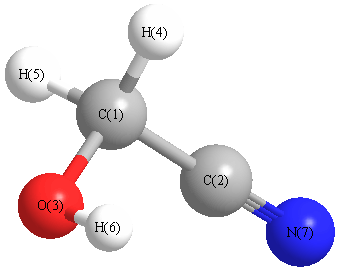 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.179 |
|
C1 |
O3 |
H6 |
108.148 |
| C2 |
C1 |
O3 |
112.743 |
|
C2 |
C1 |
H4 |
107.666 |
| C2 |
C1 |
H5 |
108.313 |
|
O3 |
C1 |
H4 |
112.719 |
| O3 |
C1 |
H5 |
106.743 |
|
H4 |
C1 |
H5 |
108.508 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.150 |
|
|
|
| 2 |
C |
-0.563 |
|
|
|
| 3 |
O |
-0.428 |
|
|
|
| 4 |
H |
0.174 |
|
|
|
| 5 |
H |
0.201 |
|
|
|
| 6 |
H |
0.378 |
|
|
|
| 7 |
N |
0.388 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.354 |
1.193 |
1.430 |
3.001 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.070 |
1.061 |
-2.063 |
| y |
1.061 |
-20.407 |
-1.844 |
| z |
-2.063 |
-1.844 |
-21.700 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.017 |
1.061 |
-2.063 |
| y |
1.061 |
6.478 |
-1.844 |
| z |
-2.063 |
-1.844 |
4.539 |
|
| Polar |
|---|
| 3z2-r2 | 9.077 |
|---|
| x2-y2 | -11.664 |
|---|
| xy | 1.061 |
|---|
| xz | -2.063 |
|---|
| yz | -1.844 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.643 |
-0.441 |
-0.002 |
| y |
-0.441 |
3.690 |
-0.138 |
| z |
-0.002 |
-0.138 |
3.129 |
<r2> (average value of r
2) Å
2
| <r2> |
65.157 |
| (<r2>)1/2 |
8.072 |
