Jump to
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -40.125091 |
| Energy at 298.15K | -40.131001 |
| Nuclear repulsion energy | 65.317637 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3801 |
3671 |
58.57 |
|
|
|
| 2 |
A |
3147 |
3039 |
26.56 |
|
|
|
| 3 |
A |
3096 |
2989 |
18.31 |
|
|
|
| 4 |
A |
3043 |
2938 |
26.93 |
|
|
|
| 5 |
A |
1713 |
1654 |
1.94 |
|
|
|
| 6 |
A |
1482 |
1431 |
10.74 |
|
|
|
| 7 |
A |
1435 |
1386 |
6.29 |
|
|
|
| 8 |
A |
1390 |
1342 |
27.36 |
|
|
|
| 9 |
A |
1270 |
1227 |
48.27 |
|
|
|
| 10 |
A |
1139 |
1100 |
4.63 |
|
|
|
| 11 |
A |
1001 |
967 |
136.57 |
|
|
|
| 12 |
A |
899 |
868 |
15.29 |
|
|
|
| 13 |
A |
551 |
532 |
15.72 |
|
|
|
| 14 |
A |
316 |
306 |
2.68 |
|
|
|
| 15 |
A |
3096 |
2990 |
32.13 |
|
|
|
| 16 |
A |
1478 |
1428 |
12.74 |
|
|
|
| 17 |
A |
1063 |
1027 |
1.75 |
|
|
|
| 18 |
A |
898 |
868 |
17.85 |
|
|
|
| 19 |
A |
393 |
380 |
150.25 |
|
|
|
| 20 |
A |
277 |
267 |
1.14 |
|
|
|
| 21 |
A |
189 |
183 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15838.3 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 15295.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.321 |
1.313 |
0.000 |
| C2 |
0.000 |
0.562 |
0.000 |
| N3 |
0.003 |
-0.729 |
0.000 |
| O4 |
1.345 |
-1.220 |
0.000 |
| H5 |
1.205 |
-2.184 |
0.000 |
| H6 |
-2.162 |
0.601 |
0.000 |
| H7 |
-1.397 |
1.963 |
0.891 |
| H8 |
-1.397 |
1.963 |
-0.891 |
| H9 |
0.950 |
1.122 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5196 | 2.4337 | 3.6770 | 4.3137 | 1.1013 | 1.1056 | 1.1056 | 2.2795 |
C2 | 1.5196 | | 1.2912 | 2.2324 | 2.9989 | 2.1620 | 2.1692 | 2.1692 | 1.1026 | N3 | 2.4337 | 1.2912 | | 1.4282 | 1.8869 | 2.5407 | 3.1620 | 3.1620 | 2.0787 | O4 | 3.6770 | 2.2324 | 1.4282 | | 0.9743 | 3.9507 | 4.2936 | 4.2936 | 2.3744 | H5 | 4.3137 | 2.9989 | 1.8869 | 0.9743 | | 4.3689 | 4.9754 | 4.9754 | 3.3154 | H6 | 1.1013 | 2.1620 | 2.5407 | 3.9507 | 4.3689 | | 1.7985 | 1.7985 | 3.1551 | H7 | 1.1056 | 2.1692 | 3.1620 | 4.2936 | 4.9754 | 1.7985 | | 1.7825 | 2.6473 | H8 | 1.1056 | 2.1692 | 3.1620 | 4.2936 | 4.9754 | 1.7985 | 1.7825 | | 2.6473 | H9 | 2.2795 | 1.1026 | 2.0787 | 2.3744 | 3.3154 | 3.1551 | 2.6473 | 2.6473 | |
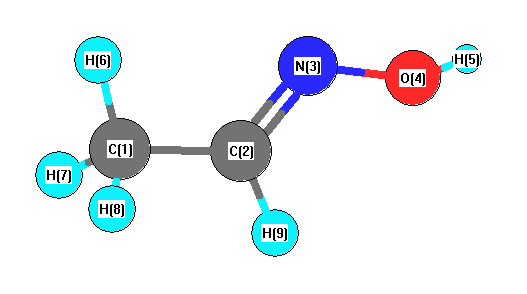 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
119.743 |
|
C1 |
C2 |
H9 |
119.921 |
| C2 |
C1 |
H6 |
110.135 |
|
C2 |
C1 |
H7 |
110.450 |
| C2 |
C1 |
H8 |
110.450 |
|
C2 |
N3 |
O4 |
110.251 |
| N3 |
C2 |
H9 |
120.336 |
|
N3 |
O4 |
H5 |
101.849 |
| H6 |
C1 |
H7 |
109.157 |
|
H6 |
C1 |
H8 |
109.157 |
| H7 |
C1 |
H8 |
107.433 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.340 |
|
|
|
| 2 |
C |
-0.457 |
|
|
|
| 3 |
N |
0.113 |
|
|
|
| 4 |
O |
-0.422 |
|
|
|
| 5 |
H |
0.382 |
|
|
|
| 6 |
H |
0.178 |
|
|
|
| 7 |
H |
0.151 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.245 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.462 |
0.799 |
0.000 |
0.923 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.081 |
-2.047 |
0.000 |
| y |
-2.047 |
-18.208 |
0.000 |
| z |
0.000 |
0.000 |
-24.921 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.516 |
-2.047 |
0.000 |
| y |
-2.047 |
6.794 |
0.000 |
| z |
0.000 |
0.000 |
-3.277 |
|
| Polar |
|---|
| 3z2-r2 | -6.555 |
|---|
| x2-y2 | -6.873 |
|---|
| xy | -2.047 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.228 |
-1.622 |
0.000 |
| y |
-1.622 |
6.830 |
0.000 |
| z |
0.000 |
0.000 |
3.070 |
<r2> (average value of r
2) Å
2
| <r2> |
78.381 |
| (<r2>)1/2 |
8.853 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -40.125799 |
| Energy at 298.15K | |
| HF Energy | -40.125799 |
| Nuclear repulsion energy | 66.752986 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3806 |
3676 |
62.82 |
|
|
|
| 2 |
A' |
3180 |
3071 |
13.53 |
|
|
|
| 3 |
A' |
3149 |
3041 |
11.27 |
|
|
|
| 4 |
A' |
3050 |
2945 |
19.12 |
|
|
|
| 5 |
A' |
1727 |
1668 |
11.56 |
|
|
|
| 6 |
A' |
1479 |
1428 |
17.21 |
|
|
|
| 7 |
A' |
1403 |
1355 |
13.64 |
|
|
|
| 8 |
A' |
1363 |
1316 |
43.05 |
|
|
|
| 9 |
A' |
1331 |
1285 |
36.91 |
|
|
|
| 10 |
A' |
1133 |
1094 |
8.96 |
|
|
|
| 11 |
A' |
928 |
896 |
139.29 |
|
|
|
| 12 |
A' |
903 |
872 |
6.67 |
|
|
|
| 13 |
A' |
660 |
637 |
10.74 |
|
|
|
| 14 |
A' |
303 |
293 |
1.54 |
|
|
|
| 15 |
A" |
3100 |
2994 |
28.41 |
|
|
|
| 16 |
A" |
1487 |
1436 |
14.41 |
|
|
|
| 17 |
A" |
1052 |
1016 |
1.22 |
|
|
|
| 18 |
A" |
854 |
824 |
20.78 |
|
|
|
| 19 |
A" |
488 |
471 |
35.21 |
|
|
|
| 20 |
A" |
385 |
371 |
103.99 |
|
|
|
| 21 |
A" |
41i |
40i |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15869.2 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 15324.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.473 |
0.490 |
0.000 |
| C2 |
0.000 |
0.871 |
0.000 |
| N3 |
1.034 |
0.095 |
0.000 |
| O4 |
0.666 |
-1.285 |
0.000 |
| H5 |
1.539 |
-1.717 |
0.000 |
| H6 |
-1.603 |
-0.600 |
0.000 |
| H7 |
-1.970 |
0.918 |
0.890 |
| H8 |
-1.970 |
0.918 |
-0.890 |
| H9 |
0.273 |
1.934 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5220 | 2.5386 | 2.7797 | 3.7344 | 1.0977 | 1.1055 | 1.1055 | 2.2666 |
C2 | 1.5220 | | 1.2932 | 2.2561 | 3.0111 | 2.1761 | 2.1622 | 2.1622 | 1.0980 | N3 | 2.5386 | 1.2932 | | 1.4274 | 1.8807 | 2.7276 | 3.2395 | 3.2395 | 1.9911 | O4 | 2.7797 | 2.2561 | 1.4274 | | 0.9741 | 2.3706 | 3.5485 | 3.5485 | 3.2429 | H5 | 3.7344 | 3.0111 | 1.8807 | 0.9741 | | 3.3351 | 4.4775 | 4.4775 | 3.8648 | H6 | 1.0977 | 2.1761 | 2.7276 | 2.3706 | 3.3351 | | 1.7975 | 1.7975 | 3.1537 | H7 | 1.1055 | 2.1622 | 3.2395 | 3.5485 | 4.4775 | 1.7975 | | 1.7804 | 2.6186 | H8 | 1.1055 | 2.1622 | 3.2395 | 3.5485 | 4.4775 | 1.7975 | 1.7804 | | 2.6186 | H9 | 2.2666 | 1.0980 | 1.9911 | 3.2429 | 3.8648 | 3.1537 | 2.6186 | 2.6186 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.601 |
|
C1 |
C2 |
H9 |
118.905 |
| C2 |
C1 |
H6 |
111.302 |
|
C2 |
C1 |
H7 |
109.741 |
| C2 |
C1 |
H8 |
109.741 |
|
C2 |
N3 |
O4 |
111.957 |
| N3 |
C2 |
H9 |
112.495 |
|
N3 |
O4 |
H5 |
101.423 |
| H6 |
C1 |
H7 |
109.347 |
|
H6 |
C1 |
H8 |
109.347 |
| H7 |
C1 |
H8 |
107.273 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.407 |
|
|
|
| 2 |
C |
-0.497 |
|
|
|
| 3 |
N |
0.176 |
|
|
|
| 4 |
O |
-0.438 |
|
|
|
| 5 |
H |
0.389 |
|
|
|
| 6 |
H |
0.212 |
|
|
|
| 7 |
H |
0.156 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
| 9 |
H |
0.253 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.763 |
0.368 |
0.000 |
0.847 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.598 |
-4.081 |
0.000 |
| y |
-4.081 |
-20.760 |
0.000 |
| z |
0.000 |
0.000 |
-25.019 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.291 |
-4.081 |
0.000 |
| y |
-4.081 |
3.048 |
0.000 |
| z |
0.000 |
0.000 |
-3.340 |
|
| Polar |
|---|
| 3z2-r2 | -6.680 |
|---|
| x2-y2 | -1.838 |
|---|
| xy | -4.081 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.870 |
-1.291 |
0.000 |
| y |
-1.291 |
5.941 |
0.000 |
| z |
0.000 |
0.000 |
3.157 |
<r2> (average value of r
2) Å
2
| <r2> |
67.965 |
| (<r2>)1/2 |
8.244 |
