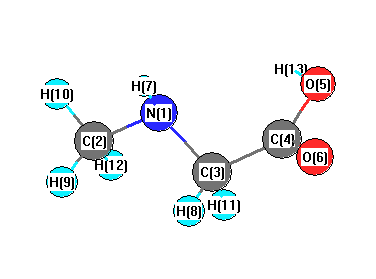
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3510 |
3390 |
0.79 |
|
|
|
| 2 |
A |
3457 |
3338 |
296.76 |
|
|
|
| 3 |
A |
3128 |
3021 |
38.48 |
|
|
|
| 4 |
A |
3106 |
2999 |
11.96 |
|
|
|
| 5 |
A |
3090 |
2984 |
46.18 |
|
|
|
| 6 |
A |
3052 |
2948 |
28.20 |
|
|
|
| 7 |
A |
3005 |
2902 |
84.06 |
|
|
|
| 8 |
A |
1851 |
1787 |
377.51 |
|
|
|
| 9 |
A |
1513 |
1462 |
13.52 |
|
|
|
| 10 |
A |
1496 |
1444 |
20.11 |
|
|
|
| 11 |
A |
1487 |
1436 |
17.29 |
|
|
|
| 12 |
A |
1472 |
1422 |
12.35 |
|
|
|
| 13 |
A |
1452 |
1402 |
1.32 |
|
|
|
| 14 |
A |
1421 |
1372 |
399.18 |
|
|
|
| 15 |
A |
1341 |
1295 |
9.93 |
|
|
|
| 16 |
A |
1278 |
1234 |
3.97 |
|
|
|
| 17 |
A |
1216 |
1174 |
17.74 |
|
|
|
| 18 |
A |
1164 |
1124 |
14.23 |
|
|
|
| 19 |
A |
1139 |
1100 |
38.92 |
|
|
|
| 20 |
A |
1118 |
1080 |
20.20 |
|
|
|
| 21 |
A |
995 |
961 |
25.91 |
|
|
|
| 22 |
A |
959 |
926 |
20.96 |
|
|
|
| 23 |
A |
896 |
865 |
67.06 |
|
|
|
| 24 |
A |
866 |
836 |
18.01 |
|
|
|
| 25 |
A |
774 |
747 |
74.84 |
|
|
|
| 26 |
A |
629 |
607 |
2.71 |
|
|
|
| 27 |
A |
564 |
544 |
7.38 |
|
|
|
| 28 |
A |
466 |
450 |
9.93 |
|
|
|
| 29 |
A |
367 |
354 |
3.59 |
|
|
|
| 30 |
A |
277 |
268 |
4.77 |
|
|
|
| 31 |
A |
204 |
197 |
2.39 |
|
|
|
| 32 |
A |
134 |
129 |
3.34 |
|
|
|
| 33 |
A |
71 |
68 |
6.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23747.5 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 22932.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.333 |
|
|
|
| 2 |
C |
-0.488 |
|
|
|
| 3 |
C |
-0.252 |
|
|
|
| 4 |
C |
-0.108 |
|
|
|
| 5 |
O |
-0.300 |
|
|
|
| 6 |
O |
-0.101 |
|
|
|
| 7 |
H |
0.272 |
|
|
|
| 8 |
H |
0.185 |
|
|
|
| 9 |
H |
0.159 |
|
|
|
| 10 |
H |
0.210 |
|
|
|
| 11 |
H |
0.173 |
|
|
|
| 12 |
H |
0.178 |
|
|
|
| 13 |
H |
0.406 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.810 |
-0.379 |
-0.625 |
5.856 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.574 |
-0.864 |
-0.539 |
| y |
-0.864 |
-37.292 |
-0.537 |
| z |
-0.539 |
-0.537 |
-33.001 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.427 |
-0.864 |
-0.539 |
| y |
-0.864 |
0.995 |
-0.537 |
| z |
-0.539 |
-0.537 |
7.432 |
|
| Polar |
|---|
| 3z2-r2 | 14.865 |
|---|
| x2-y2 | -6.282 |
|---|
| xy | -0.864 |
|---|
| xz | -0.539 |
|---|
| yz | -0.537 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.503 |
0.450 |
0.045 |
| y |
0.450 |
6.555 |
0.012 |
| z |
0.045 |
0.012 |
5.141 |
<r2> (average value of r
2) Å
2
| <r2> |
164.725 |
| (<r2>)1/2 |
12.835 |