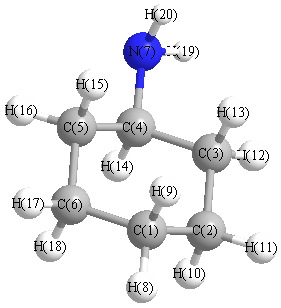
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3532 |
3410 |
1.57 |
|
|
|
| 2 |
A |
3445 |
3327 |
4.10 |
|
|
|
| 3 |
A |
3087 |
2981 |
129.10 |
|
|
|
| 4 |
A |
3076 |
2971 |
99.50 |
|
|
|
| 5 |
A |
3066 |
2961 |
35.70 |
|
|
|
| 6 |
A |
3061 |
2956 |
70.66 |
|
|
|
| 7 |
A |
3051 |
2946 |
95.10 |
|
|
|
| 8 |
A |
3024 |
2921 |
72.36 |
|
|
|
| 9 |
A |
3022 |
2918 |
4.79 |
|
|
|
| 10 |
A |
3020 |
2916 |
27.50 |
|
|
|
| 11 |
A |
3019 |
2915 |
34.87 |
|
|
|
| 12 |
A |
3007 |
2904 |
20.95 |
|
|
|
| 13 |
A |
3002 |
2899 |
31.75 |
|
|
|
| 14 |
A |
1671 |
1614 |
23.76 |
|
|
|
| 15 |
A |
1508 |
1456 |
1.65 |
|
|
|
| 16 |
A |
1491 |
1440 |
8.88 |
|
|
|
| 17 |
A |
1489 |
1438 |
7.42 |
|
|
|
| 18 |
A |
1485 |
1434 |
2.64 |
|
|
|
| 19 |
A |
1477 |
1427 |
0.22 |
|
|
|
| 20 |
A |
1417 |
1368 |
6.25 |
|
|
|
| 21 |
A |
1380 |
1332 |
3.62 |
|
|
|
| 22 |
A |
1376 |
1329 |
0.53 |
|
|
|
| 23 |
A |
1375 |
1327 |
2.45 |
|
|
|
| 24 |
A |
1358 |
1311 |
1.03 |
|
|
|
| 25 |
A |
1349 |
1302 |
3.22 |
|
|
|
| 26 |
A |
1317 |
1271 |
1.23 |
|
|
|
| 27 |
A |
1285 |
1241 |
1.08 |
|
|
|
| 28 |
A |
1272 |
1228 |
0.75 |
|
|
|
| 29 |
A |
1269 |
1225 |
4.21 |
|
|
|
| 30 |
A |
1214 |
1172 |
6.08 |
|
|
|
| 31 |
A |
1184 |
1144 |
1.21 |
|
|
|
| 32 |
A |
1113 |
1075 |
4.52 |
|
|
|
| 33 |
A |
1109 |
1071 |
12.24 |
|
|
|
| 34 |
A |
1080 |
1043 |
0.60 |
|
|
|
| 35 |
A |
1061 |
1024 |
0.48 |
|
|
|
| 36 |
A |
1037 |
1002 |
8.76 |
|
|
|
| 37 |
A |
1027 |
992 |
0.78 |
|
|
|
| 38 |
A |
968 |
934 |
2.52 |
|
|
|
| 39 |
A |
931 |
899 |
94.57 |
|
|
|
| 40 |
A |
894 |
863 |
62.49 |
|
|
|
| 41 |
A |
861 |
832 |
8.24 |
|
|
|
| 42 |
A |
856 |
827 |
11.65 |
|
|
|
| 43 |
A |
834 |
805 |
2.37 |
|
|
|
| 44 |
A |
773 |
747 |
0.25 |
|
|
|
| 45 |
A |
771 |
745 |
2.45 |
|
|
|
| 46 |
A |
534 |
516 |
1.80 |
|
|
|
| 47 |
A |
446 |
431 |
1.26 |
|
|
|
| 48 |
A |
437 |
422 |
2.18 |
|
|
|
| 49 |
A |
393 |
380 |
0.21 |
|
|
|
| 50 |
A |
333 |
322 |
15.76 |
|
|
|
| 51 |
A |
323 |
312 |
1.68 |
|
|
|
| 52 |
A |
261 |
252 |
39.32 |
|
|
|
| 53 |
A |
220 |
212 |
8.00 |
|
|
|
| 54 |
A |
155 |
150 |
0.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40870.5 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 39468.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.302 |
|
|
|
| 2 |
C |
-0.256 |
|
|
|
| 3 |
C |
-0.298 |
|
|
|
| 4 |
C |
-0.143 |
|
|
|
| 5 |
C |
-0.299 |
|
|
|
| 6 |
C |
-0.272 |
|
|
|
| 7 |
N |
-0.469 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
| 9 |
H |
0.115 |
|
|
|
| 10 |
H |
0.115 |
|
|
|
| 11 |
H |
0.166 |
|
|
|
| 12 |
H |
0.180 |
|
|
|
| 13 |
H |
0.103 |
|
|
|
| 14 |
H |
0.143 |
|
|
|
| 15 |
H |
0.116 |
|
|
|
| 16 |
H |
0.153 |
|
|
|
| 17 |
H |
0.166 |
|
|
|
| 18 |
H |
0.116 |
|
|
|
| 19 |
H |
0.253 |
|
|
|
| 20 |
H |
0.242 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.700 |
1.218 |
0.512 |
1.495 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.033 |
-4.387 |
-1.765 |
| y |
-4.387 |
-45.721 |
-0.646 |
| z |
-1.765 |
-0.646 |
-43.109 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.618 |
-4.387 |
-1.765 |
| y |
-4.387 |
-0.150 |
-0.646 |
| z |
-1.765 |
-0.646 |
3.767 |
|
| Polar |
|---|
| 3z2-r2 | 7.535 |
|---|
| x2-y2 | -2.312 |
|---|
| xy | -4.387 |
|---|
| xz | -1.765 |
|---|
| yz | -0.646 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.540 |
-0.130 |
-0.091 |
| y |
-0.130 |
10.283 |
-0.115 |
| z |
-0.091 |
-0.115 |
8.889 |
<r2> (average value of r
2) Å
2
| <r2> |
195.455 |
| (<r2>)1/2 |
13.981 |