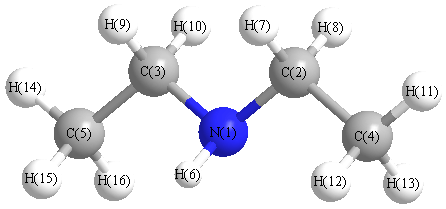
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3480 |
3360 |
3.72 |
|
|
|
| 2 |
A' |
3123 |
3016 |
91.18 |
|
|
|
| 3 |
A' |
3103 |
2996 |
120.63 |
|
|
|
| 4 |
A' |
3057 |
2952 |
62.08 |
|
|
|
| 5 |
A' |
3040 |
2935 |
17.40 |
|
|
|
| 6 |
A' |
2941 |
2840 |
211.76 |
|
|
|
| 7 |
A' |
1525 |
1473 |
2.51 |
|
|
|
| 8 |
A' |
1498 |
1446 |
3.15 |
|
|
|
| 9 |
A' |
1490 |
1438 |
14.84 |
|
|
|
| 10 |
A' |
1430 |
1381 |
1.75 |
|
|
|
| 11 |
A' |
1388 |
1340 |
1.33 |
|
|
|
| 12 |
A' |
1312 |
1267 |
2.57 |
|
|
|
| 13 |
A' |
1217 |
1175 |
1.87 |
|
|
|
| 14 |
A' |
1153 |
1114 |
15.98 |
|
|
|
| 15 |
A' |
1046 |
1010 |
5.42 |
|
|
|
| 16 |
A' |
896 |
865 |
13.19 |
|
|
|
| 17 |
A' |
813 |
785 |
4.30 |
|
|
|
| 18 |
A' |
777 |
751 |
66.41 |
|
|
|
| 19 |
A' |
414 |
400 |
0.27 |
|
|
|
| 20 |
A' |
253 |
244 |
1.48 |
|
|
|
| 21 |
A' |
183 |
177 |
1.06 |
|
|
|
| 22 |
A' |
113 |
109 |
1.49 |
|
|
|
| 23 |
A" |
3123 |
3016 |
10.15 |
|
|
|
| 24 |
A" |
3102 |
2996 |
37.50 |
|
|
|
| 25 |
A" |
3053 |
2949 |
1.18 |
|
|
|
| 26 |
A" |
3039 |
2934 |
48.64 |
|
|
|
| 27 |
A" |
2934 |
2833 |
15.60 |
|
|
|
| 28 |
A" |
1517 |
1465 |
7.05 |
|
|
|
| 29 |
A" |
1502 |
1450 |
7.52 |
|
|
|
| 30 |
A" |
1494 |
1443 |
5.65 |
|
|
|
| 31 |
A" |
1473 |
1423 |
17.13 |
|
|
|
| 32 |
A" |
1409 |
1360 |
12.76 |
|
|
|
| 33 |
A" |
1347 |
1300 |
30.80 |
|
|
|
| 34 |
A" |
1275 |
1231 |
4.72 |
|
|
|
| 35 |
A" |
1142 |
1103 |
51.17 |
|
|
|
| 36 |
A" |
1101 |
1063 |
6.69 |
|
|
|
| 37 |
A" |
1053 |
1017 |
1.64 |
|
|
|
| 38 |
A" |
930 |
898 |
0.09 |
|
|
|
| 39 |
A" |
791 |
763 |
0.67 |
|
|
|
| 40 |
A" |
413 |
399 |
0.68 |
|
|
|
| 41 |
A" |
252 |
243 |
0.49 |
|
|
|
| 42 |
A" |
119 |
115 |
1.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32657.6 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 31537.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.258 |
|
|
|
| 2 |
C |
-0.298 |
|
|
|
| 3 |
C |
-0.298 |
|
|
|
| 4 |
C |
-0.338 |
|
|
|
| 5 |
C |
-0.338 |
|
|
|
| 6 |
H |
0.218 |
|
|
|
| 7 |
H |
0.109 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.109 |
|
|
|
| 10 |
H |
0.124 |
|
|
|
| 11 |
H |
0.148 |
|
|
|
| 12 |
H |
0.121 |
|
|
|
| 13 |
H |
0.153 |
|
|
|
| 14 |
H |
0.148 |
|
|
|
| 15 |
H |
0.121 |
|
|
|
| 16 |
H |
0.153 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.910 |
0.602 |
0.000 |
1.091 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.796 |
1.636 |
0.000 |
| y |
1.636 |
-33.930 |
0.000 |
| z |
0.000 |
0.000 |
-32.883 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.610 |
1.636 |
0.000 |
| y |
1.636 |
-1.090 |
0.000 |
| z |
0.000 |
0.000 |
0.480 |
|
| Polar |
|---|
| 3z2-r2 | 0.960 |
|---|
| x2-y2 | 1.134 |
|---|
| xy | 1.636 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.493 |
0.150 |
0.000 |
| y |
0.150 |
7.023 |
0.000 |
| z |
0.000 |
0.000 |
9.462 |
<r2> (average value of r
2) Å
2
| <r2> |
158.309 |
| (<r2>)1/2 |
12.582 |