return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH3 (Propene)

using model chemistry: CCD/CEP-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/CEP-31G*
 hartrees
Energy at 0K-20.323059
Energy at 298.15K-20.328610
HF Energy-19.922251
Nuclear repulsion energy39.775959
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/CEP-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3237 3052 36.76      
2 A' 3165 2985 16.58      
3 A' 3145 2966 43.11      
4 A' 3134 2956 8.86      
5 A' 3045 2872 33.54      
6 A' 1733 1635 5.89      
7 A' 1506 1420 12.26      
8 A' 1466 1383 1.64      
9 A' 1426 1344 0.79      
10 A' 1320 1245 0.34      
11 A' 1199 1131 0.39      
12 A' 946 893 4.96      
13 A' 929 876 0.20      
14 A' 419 395 0.68      
15 A" 3113 2935 38.61      
16 A" 1490 1406 9.17      
17 A" 1077 1016 4.19      
18 A" 1005 948 30.22      
19 A" 875 825 54.71      
20 A" 581 548 15.28      
21 A" 194 183 0.39      

Unscaled Zero Point Vibrational Energy (zpe) 17501.3 cm-1
Scaled (by 0.9431) Zero Point Vibrational Energy (zpe) 16505.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/CEP-31G*
ABC
1.48418 0.30002 0.26209

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/CEP-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.165 -0.493 0.000
C2 0.000 0.494 0.000
C3 1.318 0.120 0.000
H4 1.619 -0.937 0.000
H5 2.121 0.874 0.000
H6 -0.252 1.566 0.000
H7 -0.799 -1.538 0.000
H8 -1.805 -0.345 0.894
H9 -1.805 -0.345 -0.894

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9
C11.52632.55772.81893.55842.25191.10731.11011.1101
C21.52631.37032.16072.15461.10152.18342.18252.1825
C32.55771.37031.09951.10092.13452.68993.28263.2826
H42.81892.16071.09951.87973.12522.49183.58843.5884
H53.55842.15461.10091.87972.47153.78764.20734.2073
H62.25191.10152.13453.12522.47153.15192.62012.6201
H71.10732.18342.68992.49183.78763.15191.79871.7987
H81.11012.18253.28263.58844.20732.62011.79871.7889
H91.11012.18253.28263.58844.20732.62011.79871.7889

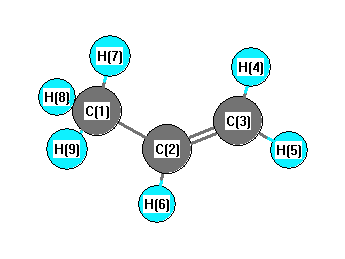
picture of Propene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.926 C1 C2 H6 117.034
C2 C1 H7 110.999 C2 C1 H8 110.760
C2 C1 H9 110.760 C2 C3 H4 121.669
C2 C3 H5 120.973 C3 C2 H6 119.040
H4 C3 H5 117.357 H7 C1 H8 108.421
H7 C1 H9 108.421 H8 C1 H9 107.360
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability