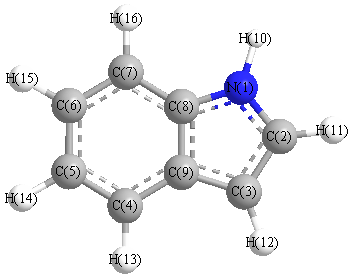
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3662 |
3537 |
59.37 |
|
|
|
| 2 |
A' |
3251 |
3140 |
4.75 |
|
|
|
| 3 |
A' |
3228 |
3117 |
0.74 |
|
|
|
| 4 |
A' |
3190 |
3080 |
25.12 |
|
|
|
| 5 |
A' |
3177 |
3068 |
33.93 |
|
|
|
| 6 |
A' |
3163 |
3054 |
6.46 |
|
|
|
| 7 |
A' |
3149 |
3041 |
0.01 |
|
|
|
| 8 |
A' |
1664 |
1607 |
1.94 |
|
|
|
| 9 |
A' |
1624 |
1569 |
0.45 |
|
|
|
| 10 |
A' |
1553 |
1499 |
9.55 |
|
|
|
| 11 |
A' |
1514 |
1462 |
2.72 |
|
|
|
| 12 |
A' |
1481 |
1430 |
22.38 |
|
|
|
| 13 |
A' |
1441 |
1392 |
5.29 |
|
|
|
| 14 |
A' |
1388 |
1341 |
43.83 |
|
|
|
| 15 |
A' |
1364 |
1317 |
6.96 |
|
|
|
| 16 |
A' |
1289 |
1245 |
8.91 |
|
|
|
| 17 |
A' |
1250 |
1207 |
8.87 |
|
|
|
| 18 |
A' |
1203 |
1161 |
2.18 |
|
|
|
| 19 |
A' |
1153 |
1113 |
0.69 |
|
|
|
| 20 |
A' |
1128 |
1089 |
0.45 |
|
|
|
| 21 |
A' |
1095 |
1058 |
28.92 |
|
|
|
| 22 |
A' |
1073 |
1036 |
7.87 |
|
|
|
| 23 |
A' |
1026 |
990 |
7.44 |
|
|
|
| 24 |
A' |
889 |
859 |
5.95 |
|
|
|
| 25 |
A' |
857 |
828 |
0.52 |
|
|
|
| 26 |
A' |
762 |
736 |
3.82 |
|
|
|
| 27 |
A' |
596 |
575 |
1.53 |
|
|
|
| 28 |
A' |
534 |
516 |
0.05 |
|
|
|
| 29 |
A' |
388 |
375 |
4.16 |
|
|
|
| 30 |
A" |
994 |
960 |
1.08 |
|
|
|
| 31 |
A" |
962 |
929 |
9.40 |
|
|
|
| 32 |
A" |
891 |
861 |
10.57 |
|
|
|
| 33 |
A" |
872 |
842 |
4.50 |
|
|
|
| 34 |
A" |
788 |
761 |
86.18 |
|
|
|
| 35 |
A" |
775 |
749 |
73.06 |
|
|
|
| 36 |
A" |
738 |
713 |
48.42 |
|
|
|
| 37 |
A" |
611 |
590 |
10.23 |
|
|
|
| 38 |
A" |
574 |
555 |
0.87 |
|
|
|
| 39 |
A" |
431 |
416 |
1.82 |
|
|
|
| 40 |
A" |
392 |
378 |
71.06 |
|
|
|
| 41 |
A" |
245 |
237 |
0.02 |
|
|
|
| 42 |
A" |
219 |
211 |
13.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28290.6 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 27320.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
0.237 |
|
|
|
| 2 |
C |
-0.526 |
|
|
|
| 3 |
C |
-0.270 |
|
|
|
| 4 |
C |
0.013 |
|
|
|
| 5 |
C |
-0.427 |
|
|
|
| 6 |
C |
-0.325 |
|
|
|
| 7 |
C |
-0.421 |
|
|
|
| 8 |
C |
-0.699 |
|
|
|
| 9 |
C |
0.222 |
|
|
|
| 10 |
H |
0.322 |
|
|
|
| 11 |
H |
0.300 |
|
|
|
| 12 |
H |
0.352 |
|
|
|
| 13 |
H |
0.331 |
|
|
|
| 14 |
H |
0.293 |
|
|
|
| 15 |
H |
0.289 |
|
|
|
| 16 |
H |
0.310 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.814 |
2.000 |
0.000 |
2.159 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.627 |
-2.209 |
0.000 |
| y |
-2.209 |
-40.773 |
0.000 |
| z |
0.000 |
0.000 |
-57.914 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.716 |
-2.209 |
0.000 |
| y |
-2.209 |
10.998 |
0.000 |
| z |
0.000 |
0.000 |
-14.714 |
|
| Polar |
|---|
| 3z2-r2 | -29.428 |
|---|
| x2-y2 | -4.854 |
|---|
| xy | -2.209 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.791 |
-1.641 |
0.000 |
| y |
-1.641 |
15.103 |
0.000 |
| z |
0.000 |
0.000 |
5.804 |
<r2> (average value of r
2) Å
2
| <r2> |
226.060 |
| (<r2>)1/2 |
15.035 |