Jump to
S1C2
S1C3
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -40.191087 |
| Energy at 298.15K | -40.196968 |
| HF Energy | -40.191087 |
| Nuclear repulsion energy | 66.391344 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3597 |
3473 |
22.00 |
|
|
|
| 2 |
A' |
3130 |
3022 |
30.09 |
|
|
|
| 3 |
A' |
3035 |
2931 |
74.15 |
|
|
|
| 4 |
A' |
2970 |
2868 |
95.48 |
|
|
|
| 5 |
A' |
1790 |
1728 |
525.62 |
|
|
|
| 6 |
A' |
1520 |
1468 |
17.56 |
|
|
|
| 7 |
A' |
1471 |
1420 |
9.13 |
|
|
|
| 8 |
A' |
1465 |
1415 |
7.97 |
|
|
|
| 9 |
A' |
1387 |
1339 |
16.61 |
|
|
|
| 10 |
A' |
1281 |
1237 |
107.12 |
|
|
|
| 11 |
A' |
1149 |
1110 |
36.20 |
|
|
|
| 12 |
A' |
996 |
961 |
40.43 |
|
|
|
| 13 |
A' |
596 |
575 |
14.38 |
|
|
|
| 14 |
A' |
334 |
323 |
8.16 |
|
|
|
| 15 |
A" |
3090 |
2984 |
48.67 |
|
|
|
| 16 |
A" |
1482 |
1431 |
5.96 |
|
|
|
| 17 |
A" |
1133 |
1094 |
0.11 |
|
|
|
| 18 |
A" |
1024 |
989 |
4.27 |
|
|
|
| 19 |
A" |
586 |
566 |
143.14 |
|
|
|
| 20 |
A" |
185 |
179 |
1.33 |
|
|
|
| 21 |
A" |
64 |
62 |
1.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16141.8 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 15588.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.287 |
-0.766 |
0.000 |
| O2 |
1.421 |
-1.248 |
0.000 |
| N3 |
0.000 |
0.580 |
0.000 |
| C4 |
-1.352 |
1.145 |
0.000 |
| H5 |
-0.636 |
-1.395 |
0.000 |
| H6 |
0.807 |
1.205 |
0.000 |
| H7 |
-2.083 |
0.319 |
0.000 |
| H8 |
-1.532 |
1.763 |
0.898 |
| H9 |
-1.532 |
1.763 |
-0.898 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2321 | 1.3765 | 2.5178 | 1.1168 | 2.0383 | 2.6067 | 3.2423 | 3.2423 |
O2 | 1.2321 | | 2.3154 | 3.6629 | 2.0623 | 2.5282 | 3.8384 | 4.3120 | 4.3120 | N3 | 1.3765 | 2.3154 | | 1.4655 | 2.0749 | 1.0205 | 2.0992 | 2.1339 | 2.1339 | C4 | 2.5178 | 3.6629 | 1.4655 | | 2.6387 | 2.1603 | 1.1023 | 1.1052 | 1.1052 | H5 | 1.1168 | 2.0623 | 2.0749 | 2.6387 | | 2.9732 | 2.2432 | 3.4031 | 3.4031 | H6 | 2.0383 | 2.5282 | 1.0205 | 2.1603 | 2.9732 | | 3.0225 | 2.5673 | 2.5673 | H7 | 2.6067 | 3.8384 | 2.0992 | 1.1023 | 2.2432 | 3.0225 | | 1.7872 | 1.7872 | H8 | 3.2423 | 4.3120 | 2.1339 | 1.1052 | 3.4031 | 2.5673 | 1.7872 | | 1.7968 | H9 | 3.2423 | 4.3120 | 2.1339 | 1.1052 | 3.4031 | 2.5673 | 1.7872 | 1.7968 | |
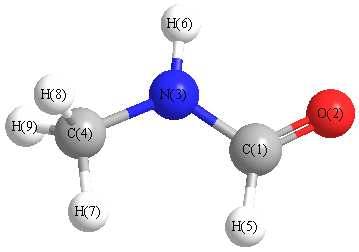 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.696 |
|
C1 |
N3 |
H6 |
115.700 |
| O2 |
C1 |
N3 |
125.051 |
|
O2 |
C1 |
H5 |
122.725 |
| N3 |
C1 |
H5 |
112.224 |
|
N3 |
C4 |
H7 |
108.846 |
| N3 |
C4 |
H8 |
111.445 |
|
N3 |
C4 |
H9 |
111.445 |
| C4 |
N3 |
H6 |
119.604 |
|
H7 |
C4 |
H8 |
108.115 |
| H7 |
C4 |
H9 |
108.115 |
|
H8 |
C4 |
H9 |
108.762 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.339 |
|
|
|
| 2 |
O |
-0.152 |
|
|
|
| 3 |
N |
-0.038 |
|
|
|
| 4 |
C |
-0.525 |
|
|
|
| 5 |
H |
0.156 |
|
|
|
| 6 |
H |
0.328 |
|
|
|
| 7 |
H |
0.188 |
|
|
|
| 8 |
H |
0.192 |
|
|
|
| 9 |
H |
0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.327 |
2.928 |
0.000 |
4.432 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.247 |
4.414 |
0.000 |
| y |
4.414 |
-24.095 |
0.000 |
| z |
0.000 |
0.000 |
-24.647 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.876 |
4.414 |
0.000 |
| y |
4.414 |
1.352 |
0.000 |
| z |
0.000 |
0.000 |
0.524 |
|
| Polar |
|---|
| 3z2-r2 | 1.049 |
|---|
| x2-y2 | -2.152 |
|---|
| xy | 4.414 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.444 |
-1.177 |
0.000 |
| y |
-1.177 |
5.563 |
0.000 |
| z |
0.000 |
0.000 |
3.135 |
<r2> (average value of r
2) Å
2
| <r2> |
75.407 |
| (<r2>)1/2 |
8.684 |
Jump to
S1C1
S1C3
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -40.191087 |
| Energy at 298.15K | -40.196968 |
| HF Energy | -40.191087 |
| Nuclear repulsion energy | 66.391344 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -40.192172 |
| Energy at 298.15K | -40.198075 |
| HF Energy | -40.192172 |
| Nuclear repulsion energy | 67.504765 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3622 |
3497 |
18.06 |
|
|
|
| 2 |
A |
3168 |
3059 |
4.04 |
|
|
|
| 3 |
A |
3098 |
2992 |
54.51 |
|
|
|
| 4 |
A |
3042 |
2937 |
59.59 |
|
|
|
| 5 |
A |
2974 |
2872 |
129.39 |
|
|
|
| 6 |
A |
1779 |
1718 |
379.37 |
|
|
|
| 7 |
A |
1538 |
1486 |
90.02 |
|
|
|
| 8 |
A |
1500 |
1448 |
9.48 |
|
|
|
| 9 |
A |
1480 |
1429 |
42.90 |
|
|
|
| 10 |
A |
1439 |
1390 |
18.60 |
|
|
|
| 11 |
A |
1408 |
1359 |
6.69 |
|
|
|
| 12 |
A |
1203 |
1162 |
88.43 |
|
|
|
| 13 |
A |
1142 |
1103 |
15.32 |
|
|
|
| 14 |
A |
1137 |
1098 |
0.59 |
|
|
|
| 15 |
A |
1006 |
971 |
0.55 |
|
|
|
| 16 |
A |
946 |
914 |
18.90 |
|
|
|
| 17 |
A |
750 |
724 |
0.50 |
|
|
|
| 18 |
A |
520 |
503 |
48.25 |
|
|
|
| 19 |
A |
288 |
278 |
12.95 |
|
|
|
| 20 |
A |
243 |
235 |
93.94 |
|
|
|
| 21 |
A |
53 |
51 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16166.8 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 15612.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.875 |
0.440 |
0.002 |
| O2 |
1.403 |
-0.677 |
0.001 |
| N3 |
-0.481 |
0.665 |
-0.020 |
| C4 |
-1.446 |
-0.447 |
0.004 |
| H5 |
1.470 |
1.383 |
0.018 |
| H6 |
-0.812 |
1.623 |
0.062 |
| H7 |
-2.455 |
-0.056 |
-0.206 |
| H8 |
-1.180 |
-1.192 |
-0.762 |
| H9 |
-1.451 |
-0.957 |
0.985 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2355 | 1.3744 | 2.4845 | 1.1158 | 2.0611 | 3.3729 | 2.7325 | 2.8849 |
O2 | 1.2355 | | 2.3131 | 2.8589 | 2.0611 | 3.1939 | 3.9136 | 2.7420 | 3.0320 | N3 | 1.3744 | 2.3131 | | 1.4722 | 2.0799 | 1.0176 | 2.1096 | 2.1176 | 2.1398 | C4 | 2.4845 | 2.8589 | 1.4722 | | 3.4433 | 2.1661 | 1.1022 | 1.1016 | 1.1051 | H5 | 1.1158 | 2.0611 | 2.0799 | 3.4433 | | 2.2951 | 4.1871 | 3.7765 | 3.8656 | H6 | 2.0611 | 3.1939 | 1.0176 | 2.1661 | 2.2951 | | 2.3650 | 2.9563 | 2.8138 | H7 | 3.3729 | 3.9136 | 2.1096 | 1.1022 | 4.1871 | 2.3650 | | 1.7962 | 1.7994 | H8 | 2.7325 | 2.7420 | 2.1176 | 1.1016 | 3.7765 | 2.9563 | 1.7962 | | 1.7834 | H9 | 2.8849 | 3.0320 | 2.1398 | 1.1051 | 3.8656 | 2.8138 | 1.7994 | 1.7834 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.536 |
|
C1 |
N3 |
H6 |
118.241 |
| O2 |
C1 |
N3 |
124.741 |
|
O2 |
C1 |
H5 |
122.386 |
| N3 |
C1 |
H5 |
112.873 |
|
N3 |
C4 |
H7 |
109.222 |
| N3 |
C4 |
H8 |
109.884 |
|
N3 |
C4 |
H9 |
111.462 |
| C4 |
N3 |
H6 |
119.800 |
|
H7 |
C4 |
H8 |
109.183 |
| H7 |
C4 |
H9 |
109.221 |
|
H8 |
C4 |
H9 |
107.831 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.324 |
|
|
|
| 2 |
O |
-0.174 |
|
|
|
| 3 |
N |
-0.071 |
|
|
|
| 4 |
C |
-0.522 |
|
|
|
| 5 |
H |
0.177 |
|
|
|
| 6 |
H |
0.302 |
|
|
|
| 7 |
H |
0.184 |
|
|
|
| 8 |
H |
0.242 |
|
|
|
| 9 |
H |
0.184 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.111 |
2.756 |
0.000 |
4.156 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.465 |
2.928 |
0.000 |
| y |
2.928 |
-21.583 |
0.000 |
| z |
0.000 |
0.000 |
-24.678 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.334 |
2.928 |
0.000 |
| y |
2.928 |
3.488 |
0.000 |
| z |
0.000 |
0.000 |
-1.154 |
|
| Polar |
|---|
| 3z2-r2 | -2.309 |
|---|
| x2-y2 | -3.882 |
|---|
| xy | 2.928 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.004 |
-0.262 |
-0.000 |
| y |
-0.262 |
4.778 |
0.000 |
| z |
-0.000 |
0.000 |
3.129 |
<r2> (average value of r
2) Å
2
| <r2> |
67.243 |
| (<r2>)1/2 |
8.200 |
