Jump to
S1C2
Energy calculated at MP2/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -31.335546 |
| Energy at 298.15K | -31.337938 |
| HF Energy | -30.971766 |
| Nuclear repulsion energy | 32.841454 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2728 |
2590 |
0.66 |
|
|
|
| 2 |
A |
913 |
867 |
0.44 |
|
|
|
| 3 |
A |
513 |
488 |
1.14 |
|
|
|
| 4 |
A |
311 |
295 |
38.22 |
|
|
|
| 5 |
A |
208 |
198 |
0.00 |
|
|
|
| 6 |
B |
2728 |
2590 |
15.54 |
|
|
|
| 7 |
B |
900 |
855 |
12.44 |
|
|
|
| 8 |
B |
517 |
491 |
27.19 |
|
|
|
| 9 |
B |
340 |
322 |
26.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4578.8 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 4347.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/CEP-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.852 |
| S2 |
0.000 |
1.678 |
-0.390 |
| S3 |
0.000 |
-1.678 |
-0.390 |
| H4 |
-1.339 |
1.774 |
-0.583 |
| H5 |
1.339 |
-1.774 |
-0.583 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0876 | 2.0876 | 2.6456 | 2.6456 |
S2 | 2.0876 | | 3.3560 | 1.3564 | 3.7076 | S3 | 2.0876 | 3.3560 | | 3.7076 | 1.3564 | H4 | 2.6456 | 1.3564 | 3.7076 | | 4.4454 | H5 | 2.6456 | 3.7076 | 1.3564 | 4.4454 | |
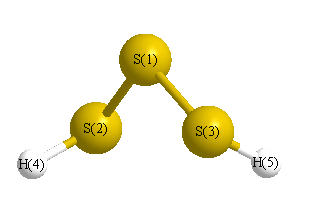 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.136 |
|
S1 |
S3 |
H5 |
98.136 |
| S2 |
S1 |
S3 |
106.991 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -31.334876 |
| Energy at 298.15K | -31.337271 |
| HF Energy | -30.971130 |
| Nuclear repulsion energy | 32.842316 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2722 |
2584 |
23.12 |
|
|
|
| 2 |
A' |
914 |
867 |
8.23 |
|
|
|
| 3 |
A' |
513 |
487 |
0.94 |
|
|
|
| 4 |
A' |
339 |
322 |
27.97 |
|
|
|
| 5 |
A' |
210 |
199 |
0.22 |
|
|
|
| 6 |
A" |
2724 |
2586 |
0.96 |
|
|
|
| 7 |
A" |
905 |
859 |
7.17 |
|
|
|
| 8 |
A" |
518 |
491 |
29.79 |
|
|
|
| 9 |
A" |
311 |
295 |
14.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4577.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 4345.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.054 |
0.849 |
0.000 |
| S2 |
-0.054 |
-0.392 |
1.678 |
| S3 |
-0.054 |
-0.392 |
-1.678 |
| H4 |
1.289 |
-0.514 |
1.830 |
| H5 |
1.289 |
-0.514 |
-1.830 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0872 | 2.0872 | 2.6472 | 2.6472 |
S2 | 2.0872 | | 3.3568 | 1.3570 | 3.7585 | S3 | 2.0872 | 3.3568 | | 3.7585 | 1.3570 | H4 | 2.6472 | 1.3570 | 3.7585 | | 3.6596 | H5 | 2.6472 | 3.7585 | 1.3570 | 3.6596 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.218 |
|
S1 |
S3 |
H5 |
98.218 |
| S2 |
S1 |
S3 |
107.057 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
