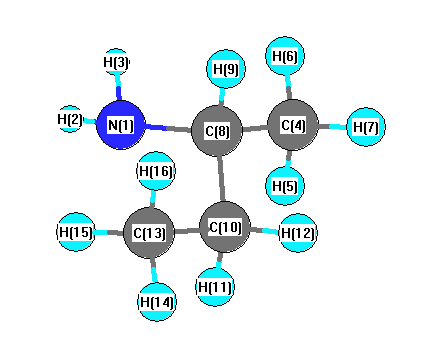
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3555 |
3433 |
1.35 |
|
|
|
| 2 |
A |
3470 |
3351 |
3.45 |
|
|
|
| 3 |
A |
3118 |
3011 |
76.20 |
|
|
|
| 4 |
A |
3106 |
2999 |
75.78 |
|
|
|
| 5 |
A |
3100 |
2994 |
85.79 |
|
|
|
| 6 |
A |
3098 |
2991 |
27.28 |
|
|
|
| 7 |
A |
3075 |
2970 |
5.12 |
|
|
|
| 8 |
A |
3036 |
2932 |
34.20 |
|
|
|
| 9 |
A |
3032 |
2928 |
60.18 |
|
|
|
| 10 |
A |
3031 |
2927 |
15.99 |
|
|
|
| 11 |
A |
2937 |
2836 |
89.91 |
|
|
|
| 12 |
A |
1677 |
1619 |
40.17 |
|
|
|
| 13 |
A |
1510 |
1458 |
9.24 |
|
|
|
| 14 |
A |
1500 |
1448 |
4.36 |
|
|
|
| 15 |
A |
1498 |
1447 |
5.33 |
|
|
|
| 16 |
A |
1491 |
1440 |
4.24 |
|
|
|
| 17 |
A |
1474 |
1423 |
1.20 |
|
|
|
| 18 |
A |
1424 |
1375 |
5.67 |
|
|
|
| 19 |
A |
1407 |
1359 |
9.48 |
|
|
|
| 20 |
A |
1399 |
1351 |
5.37 |
|
|
|
| 21 |
A |
1372 |
1325 |
14.36 |
|
|
|
| 22 |
A |
1319 |
1274 |
4.64 |
|
|
|
| 23 |
A |
1291 |
1246 |
2.09 |
|
|
|
| 24 |
A |
1250 |
1207 |
0.68 |
|
|
|
| 25 |
A |
1180 |
1140 |
2.14 |
|
|
|
| 26 |
A |
1148 |
1108 |
17.00 |
|
|
|
| 27 |
A |
1054 |
1018 |
2.03 |
|
|
|
| 28 |
A |
1020 |
985 |
3.44 |
|
|
|
| 29 |
A |
1009 |
974 |
7.27 |
|
|
|
| 30 |
A |
969 |
935 |
6.23 |
|
|
|
| 31 |
A |
935 |
903 |
21.40 |
|
|
|
| 32 |
A |
887 |
857 |
94.80 |
|
|
|
| 33 |
A |
803 |
776 |
1.65 |
|
|
|
| 34 |
A |
750 |
725 |
0.15 |
|
|
|
| 35 |
A |
467 |
451 |
8.28 |
|
|
|
| 36 |
A |
438 |
423 |
6.62 |
|
|
|
| 37 |
A |
364 |
351 |
1.28 |
|
|
|
| 38 |
A |
289 |
279 |
44.03 |
|
|
|
| 39 |
A |
250 |
242 |
0.04 |
|
|
|
| 40 |
A |
227 |
219 |
5.85 |
|
|
|
| 41 |
A |
215 |
208 |
7.63 |
|
|
|
| 42 |
A |
99 |
96 |
1.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32634.1 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 31514.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.566 |
|
|
|
| 2 |
H |
0.247 |
|
|
|
| 3 |
H |
0.249 |
|
|
|
| 4 |
C |
-0.471 |
|
|
|
| 5 |
H |
0.149 |
|
|
|
| 6 |
H |
0.158 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
| 8 |
C |
-0.034 |
|
|
|
| 9 |
H |
0.132 |
|
|
|
| 10 |
C |
-0.238 |
|
|
|
| 11 |
H |
0.134 |
|
|
|
| 12 |
H |
0.119 |
|
|
|
| 13 |
C |
-0.462 |
|
|
|
| 14 |
H |
0.179 |
|
|
|
| 15 |
H |
0.135 |
|
|
|
| 16 |
H |
0.132 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.069 |
0.116 |
1.403 |
1.410 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.054 |
0.418 |
1.229 |
| y |
0.418 |
-31.712 |
2.220 |
| z |
1.229 |
2.220 |
-35.042 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.323 |
0.418 |
1.229 |
| y |
0.418 |
2.336 |
2.220 |
| z |
1.229 |
2.220 |
-2.660 |
|
| Polar |
|---|
| 3z2-r2 | -5.319 |
|---|
| x2-y2 | -1.342 |
|---|
| xy | 0.418 |
|---|
| xz | 1.229 |
|---|
| yz | 2.220 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.831 |
-0.138 |
-0.012 |
| y |
-0.138 |
7.503 |
-0.087 |
| z |
-0.012 |
-0.087 |
6.468 |
<r2> (average value of r
2) Å
2
| <r2> |
129.538 |
| (<r2>)1/2 |
11.381 |