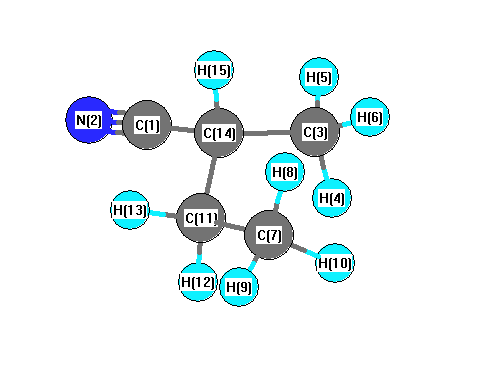
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3138 |
3030 |
32.29 |
|
|
|
| 2 |
A |
3134 |
3027 |
38.99 |
|
|
|
| 3 |
A |
3123 |
3016 |
74.60 |
|
|
|
| 4 |
A |
3113 |
3007 |
49.10 |
|
|
|
| 5 |
A |
3094 |
2988 |
3.72 |
|
|
|
| 6 |
A |
3059 |
2954 |
14.18 |
|
|
|
| 7 |
A |
3057 |
2952 |
34.63 |
|
|
|
| 8 |
A |
3051 |
2947 |
4.90 |
|
|
|
| 9 |
A |
3044 |
2940 |
26.10 |
|
|
|
| 10 |
A |
2332 |
2252 |
17.95 |
|
|
|
| 11 |
A |
1512 |
1460 |
2.98 |
|
|
|
| 12 |
A |
1505 |
1454 |
11.76 |
|
|
|
| 13 |
A |
1500 |
1448 |
6.87 |
|
|
|
| 14 |
A |
1495 |
1444 |
11.08 |
|
|
|
| 15 |
A |
1486 |
1435 |
0.14 |
|
|
|
| 16 |
A |
1415 |
1366 |
2.13 |
|
|
|
| 17 |
A |
1411 |
1363 |
4.36 |
|
|
|
| 18 |
A |
1372 |
1325 |
1.04 |
|
|
|
| 19 |
A |
1336 |
1290 |
1.32 |
|
|
|
| 20 |
A |
1306 |
1261 |
3.32 |
|
|
|
| 21 |
A |
1270 |
1226 |
1.72 |
|
|
|
| 22 |
A |
1172 |
1132 |
2.81 |
|
|
|
| 23 |
A |
1130 |
1092 |
0.22 |
|
|
|
| 24 |
A |
1103 |
1065 |
7.91 |
|
|
|
| 25 |
A |
1036 |
1000 |
1.71 |
|
|
|
| 26 |
A |
985 |
952 |
0.58 |
|
|
|
| 27 |
A |
963 |
930 |
7.56 |
|
|
|
| 28 |
A |
886 |
855 |
0.16 |
|
|
|
| 29 |
A |
799 |
771 |
1.18 |
|
|
|
| 30 |
A |
752 |
726 |
4.86 |
|
|
|
| 31 |
A |
549 |
531 |
0.49 |
|
|
|
| 32 |
A |
521 |
503 |
1.62 |
|
|
|
| 33 |
A |
385 |
372 |
0.26 |
|
|
|
| 34 |
A |
315 |
304 |
0.23 |
|
|
|
| 35 |
A |
276 |
267 |
0.06 |
|
|
|
| 36 |
A |
218 |
211 |
0.06 |
|
|
|
| 37 |
A |
199 |
192 |
1.82 |
|
|
|
| 38 |
A |
159 |
154 |
4.68 |
|
|
|
| 39 |
A |
82 |
79 |
1.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28640.9 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 27658.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.627 |
|
|
|
| 2 |
N |
0.369 |
|
|
|
| 3 |
C |
-0.538 |
|
|
|
| 4 |
H |
0.163 |
|
|
|
| 5 |
H |
0.186 |
|
|
|
| 6 |
H |
0.174 |
|
|
|
| 7 |
C |
-0.426 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
| 9 |
H |
0.189 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
C |
-0.294 |
|
|
|
| 12 |
H |
0.150 |
|
|
|
| 13 |
H |
0.165 |
|
|
|
| 14 |
C |
0.049 |
|
|
|
| 15 |
H |
0.162 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.988 |
1.418 |
0.525 |
4.265 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.038 |
-4.290 |
-0.867 |
| y |
-4.290 |
-37.280 |
-0.128 |
| z |
-0.867 |
-0.128 |
-36.191 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.303 |
-4.290 |
-0.867 |
| y |
-4.290 |
5.335 |
-0.128 |
| z |
-0.867 |
-0.128 |
6.968 |
|
| Polar |
|---|
| 3z2-r2 | 13.936 |
|---|
| x2-y2 | -11.758 |
|---|
| xy | -4.290 |
|---|
| xz | -0.867 |
|---|
| yz | -0.128 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.565 |
0.588 |
0.271 |
| y |
0.588 |
7.880 |
0.080 |
| z |
0.271 |
0.080 |
6.730 |
<r2> (average value of r
2) Å
2
| <r2> |
163.181 |
| (<r2>)1/2 |
12.774 |