Jump to
S1C2
S1C3
Energy calculated at HF/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -39.703083 |
| Energy at 298.15K | |
| HF Energy | -39.703083 |
| Nuclear repulsion energy | 49.333581 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
587 |
527 |
0.00 |
77.39 |
0.07 |
0.14 |
| 2 |
A1 |
197 |
177 |
0.00 |
4.46 |
0.72 |
0.83 |
| 3 |
B1 |
558 |
501 |
0.00 |
42.87 |
0.75 |
0.86 |
| 4 |
B2 |
361 |
324 |
0.47 |
9.38 |
0.75 |
0.86 |
| 5 |
E |
536 |
482 |
0.10 |
10.44 |
0.75 |
0.86 |
| 5 |
E |
536 |
482 |
0.10 |
10.44 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1387.8 cm
-1
Scaled (by 0.898) Zero Point Vibrational Energy (zpe) 1246.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/CEP-31G*
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.438 |
0.285 |
| S2 |
0.000 |
-1.438 |
0.285 |
| S3 |
-1.438 |
0.000 |
-0.285 |
| S4 |
1.438 |
0.000 |
-0.285 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8758 | 2.1117 | 2.1117 |
S2 | 2.8758 | | 2.1117 | 2.1117 | S3 | 2.1117 | 2.1117 | | 2.8758 | S4 | 2.1117 | 2.1117 | 2.8758 | |
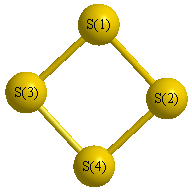 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.084 |
|
S1 |
S3 |
S4 |
47.084 |
| S2 |
S1 |
S3 |
47.084 |
|
S2 |
S4 |
S3 |
47.084 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.011 |
0.000 |
0.000 |
| y |
0.000 |
-47.011 |
0.000 |
| z |
0.000 |
0.000 |
-52.673 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.831 |
0.000 |
0.000 |
| y |
0.000 |
2.831 |
0.000 |
| z |
0.000 |
0.000 |
-5.662 |
|
| Polar |
|---|
| 3z2-r2 | -11.324 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.449 |
0.000 |
0.000 |
| y |
0.000 |
10.449 |
0.000 |
| z |
0.000 |
0.000 |
4.198 |
<r2> (average value of r
2) Å
2
| <r2> |
82.109 |
| (<r2>)1/2 |
9.061 |
Jump to
S1C1
S1C3
Energy calculated at HF/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -39.674851 |
| Energy at 298.15K | |
| HF Energy | -39.674851 |
| Nuclear repulsion energy | 46.630489 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
712 |
639 |
10.14 |
29.01 |
0.60 |
0.75 |
| 2 |
A1 |
557 |
500 |
5.56 |
387.03 |
0.26 |
0.41 |
| 3 |
A1 |
138 |
124 |
0.00 |
29.05 |
0.45 |
0.62 |
| 4 |
A2 |
215 |
194 |
0.00 |
0.60 |
0.75 |
0.86 |
| 5 |
B2 |
636 |
571 |
504.23 |
0.49 |
0.75 |
0.86 |
| 6 |
B2 |
360 |
323 |
2.00 |
14.69 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1309.2 cm
-1
Scaled (by 0.898) Zero Point Vibrational Energy (zpe) 1175.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/CEP-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.004 |
0.899 |
| S2 |
0.000 |
-1.004 |
0.899 |
| S3 |
0.000 |
1.695 |
-0.899 |
| S4 |
0.000 |
-1.695 |
-0.899 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.0072 | 1.9269 | 3.2432 |
S2 | 2.0072 | | 3.2432 | 1.9269 | S3 | 1.9269 | 3.2432 | | 3.3907 | S4 | 3.2432 | 1.9269 | 3.3907 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
111.039 |
|
S1 |
S3 |
S4 |
68.961 |
| S2 |
S1 |
S3 |
111.039 |
|
S2 |
S4 |
S3 |
68.961 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.185 |
|
|
|
| 2 |
S |
0.185 |
|
|
|
| 3 |
S |
-0.185 |
|
|
|
| 4 |
S |
-0.185 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.469 |
2.469 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.105 |
0.000 |
0.000 |
| y |
0.000 |
-53.065 |
0.000 |
| z |
0.000 |
0.000 |
-47.626 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.241 |
0.000 |
0.000 |
| y |
0.000 |
-4.700 |
0.000 |
| z |
0.000 |
0.000 |
3.459 |
|
| Polar |
|---|
| 3z2-r2 | 6.918 |
|---|
| x2-y2 | 3.961 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.537 |
0.000 |
0.000 |
| y |
0.000 |
23.610 |
0.000 |
| z |
0.000 |
0.000 |
10.780 |
<r2> (average value of r
2) Å
2
| <r2> |
97.170 |
| (<r2>)1/2 |
9.857 |
Jump to
S1C1
S1C2
Energy calculated at HF/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -39.643310 |
| Energy at 298.15K | |
| HF Energy | -39.643310 |
| Nuclear repulsion energy | 47.645000 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
821 |
737 |
0.00 |
48.26 |
0.35 |
0.52 |
| 2 |
Ag |
332 |
299 |
0.00 |
970.75 |
0.33 |
0.49 |
| 3 |
Au |
281 |
252 |
0.00 |
0.00 |
0.00 |
0.00 |
| 4 |
B1u |
709 |
637 |
631.42 |
0.00 |
0.00 |
0.00 |
| 5 |
B2u |
326i |
293i |
59.50 |
0.00 |
0.00 |
0.00 |
| 6 |
B3g |
376 |
338 |
0.00 |
18.68 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1096.4 cm
-1
Scaled (by 0.898) Zero Point Vibrational Energy (zpe) 984.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/CEP-31G*
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.941 |
1.251 |
| S2 |
0.000 |
0.941 |
-1.251 |
| S3 |
0.000 |
-0.941 |
1.251 |
| S4 |
0.000 |
-0.941 |
-1.251 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5024 | 1.8815 | 3.1308 |
S2 | 2.5024 | | 3.1308 | 1.8815 | S3 | 1.8815 | 3.1308 | | 2.5024 | S4 | 3.1308 | 1.8815 | 2.5024 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.645 |
0.000 |
0.000 |
| y |
0.000 |
-46.471 |
0.000 |
| z |
0.000 |
0.000 |
-52.603 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.108 |
0.000 |
0.000 |
| y |
0.000 |
4.653 |
0.000 |
| z |
0.000 |
0.000 |
-4.545 |
|
| Polar |
|---|
| 3z2-r2 | -9.090 |
|---|
| x2-y2 | -3.174 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.539 |
0.000 |
0.000 |
| y |
0.000 |
11.744 |
0.000 |
| z |
0.000 |
0.000 |
22.706 |
<r2> (average value of r
2) Å
2
| <r2> |
89.776 |
| (<r2>)1/2 |
9.475 |
