Jump to
S1C2
S1C3
Energy calculated at MP2/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -40.185347 |
| Energy at 298.15K | |
| HF Energy | -39.701880 |
| Nuclear repulsion energy | 48.833697 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
533 |
506 |
0.00 |
45.12 |
0.06 |
0.11 |
| 2 |
A1 |
200 |
190 |
0.00 |
5.43 |
0.72 |
0.84 |
| 3 |
B1 |
482 |
458 |
0.00 |
26.13 |
0.75 |
0.86 |
| 4 |
B2 |
311 |
295 |
0.12 |
10.12 |
0.75 |
0.86 |
| 5 |
E |
457 |
434 |
0.02 |
7.75 |
0.75 |
0.86 |
| 5 |
E |
457 |
434 |
0.02 |
7.75 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1220.8 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 1159.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/CEP-31G*
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.444 |
0.316 |
| S2 |
0.000 |
-1.444 |
0.316 |
| S3 |
-1.444 |
0.000 |
-0.316 |
| S4 |
1.444 |
0.000 |
-0.316 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8886 | 2.1379 | 2.1379 |
S2 | 2.8886 | | 2.1379 | 2.1379 | S3 | 2.1379 | 2.1379 | | 2.8886 | S4 | 2.1379 | 2.1379 | 2.8886 | |
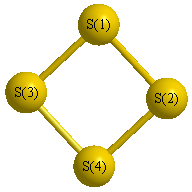 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.502 |
|
S1 |
S3 |
S4 |
47.502 |
| S2 |
S1 |
S3 |
47.502 |
|
S2 |
S4 |
S3 |
47.502 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -40.246666 |
| Energy at 298.15K | |
| HF Energy | -39.627355 |
| Nuclear repulsion energy | 44.836069 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
640 |
607 |
0.00 |
1714.41 |
0.41 |
0.58 |
| 2 |
A1 |
341 |
324 |
0.11 |
1.56 |
0.33 |
0.50 |
| 3 |
A1 |
159 |
151 |
0.00 |
486788.70 |
0.33 |
0.50 |
| 4 |
A2 |
210 |
199 |
0.00 |
0.00 |
0.75 |
0.86 |
| 5 |
B2 |
837 |
795 |
2869.58 |
0.01 |
0.75 |
0.86 |
| 6 |
B2 |
292 |
277 |
0.00 |
92.97 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1239.3 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 1176.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/CEP-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.358 |
0.980 |
| S2 |
0.000 |
-1.358 |
0.980 |
| S3 |
0.000 |
1.359 |
-0.980 |
| S4 |
0.000 |
-1.359 |
-0.980 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.7166 | 1.9600 | 3.3501 |
S2 | 2.7166 | | 3.3501 | 1.9600 | S3 | 1.9600 | 3.3501 | | 2.7171 | S4 | 3.3501 | 1.9600 | 2.7171 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.006 |
|
S1 |
S3 |
S4 |
89.994 |
| S2 |
S1 |
S3 |
90.006 |
|
S2 |
S4 |
S3 |
89.994 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -40.246666 |
| Energy at 298.15K | |
| HF Energy | -39.627358 |
| Nuclear repulsion energy | 44.836115 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
640 |
607 |
0.00 |
1707.98 |
0.41 |
0.58 |
| 2 |
Ag |
159 |
151 |
0.00 |
485060.10 |
0.33 |
0.50 |
| 3 |
Au |
210 |
199 |
0.00 |
0.00 |
0.00 |
0.00 |
| 4 |
B1u |
837 |
795 |
2862.10 |
0.00 |
0.75 |
0.86 |
| 5 |
B2u |
341 |
324 |
0.11 |
0.00 |
0.33 |
0.50 |
| 6 |
B3g |
292 |
277 |
0.00 |
92.83 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1239.2 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 1176.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/CEP-31G*
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.980 |
1.358 |
| S2 |
0.000 |
0.980 |
-1.358 |
| S3 |
0.000 |
-0.980 |
1.358 |
| S4 |
0.000 |
-0.980 |
-1.358 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.7169 | 1.9600 | 3.3501 |
S2 | 2.7169 | | 3.3501 | 1.9600 | S3 | 1.9600 | 3.3501 | | 2.7169 | S4 | 3.3501 | 1.9600 | 2.7169 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
