Jump to
S1C2
S1C3
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -15.777311 |
| Energy at 298.15K | -15.776227 |
| HF Energy | -15.777311 |
| Nuclear repulsion energy | 10.802613 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.120 |
| C2 |
0.000 |
0.000 |
-0.152 |
| N3 |
0.000 |
0.000 |
1.039 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9684 | 3.1589 |
C2 | 1.9684 | | 1.1905 | N3 | 3.1589 | 1.1905 | |
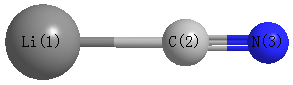 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.330 |
|
|
|
| 2 |
C |
-0.494 |
|
|
|
| 3 |
N |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-9.288 |
9.288 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.244 |
0.000 |
0.000 |
| y |
0.000 |
-14.244 |
0.000 |
| z |
0.000 |
0.000 |
2.795 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.519 |
0.000 |
0.000 |
| y |
0.000 |
-8.519 |
0.000 |
| z |
0.000 |
0.000 |
17.039 |
|
| Polar |
|---|
| 3z2-r2 | 34.078 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.512 |
0.000 |
0.000 |
| y |
0.000 |
2.512 |
0.000 |
| z |
0.000 |
0.000 |
4.608 |
<r2> (average value of r
2) Å
2
| <r2> |
15.332 |
| (<r2>)1/2 |
3.916 |
Jump to
S1C1
S1C3
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -15.783455 |
| Energy at 298.15K | |
| HF Energy | -15.783455 |
| Nuclear repulsion energy | 11.034684 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.455 |
-0.589 |
0.000 |
| C2 |
-0.727 |
-0.378 |
0.000 |
| N3 |
0.000 |
0.577 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.1923 | 1.8641 |
C2 | 2.1923 | | 1.2003 | N3 | 1.8641 | 1.2003 | |
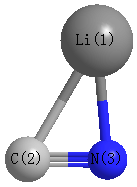 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
58.215 |
|
Li1 |
N3 |
C2 |
88.599 |
| C2 |
Li1 |
N3 |
33.186 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.333 |
|
|
|
| 2 |
C |
-0.417 |
|
|
|
| 3 |
N |
0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
8.409 |
0.128 |
0.000 |
8.410 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-1.634 |
-1.880 |
0.000 |
| y |
-1.880 |
-15.960 |
0.000 |
| z |
0.000 |
0.000 |
-14.330 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
13.511 |
-1.880 |
0.000 |
| y |
-1.880 |
-7.978 |
0.000 |
| z |
0.000 |
0.000 |
-5.533 |
|
| Polar |
|---|
| 3z2-r2 | -11.065 |
|---|
| x2-y2 | 14.326 |
|---|
| xy | -1.880 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.512 |
0.000 |
0.000 |
| y |
0.000 |
2.512 |
0.000 |
| z |
0.000 |
0.000 |
4.608 |
<r2> (average value of r
2) Å
2
| <r2> |
14.514 |
| (<r2>)1/2 |
3.810 |
Jump to
S1C1
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -15.782143 |
| Energy at 298.15K | |
| HF Energy | -15.782143 |
| Nuclear repulsion energy | 10.961567 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
2109 |
2037 |
172.71 |
|
|
|
| 2 |
Σ |
641 |
619 |
149.62 |
|
|
|
| 3 |
Π |
120i |
116i |
34.86 |
|
|
|
| 3 |
Π |
120i |
116i |
34.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1254.7 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 1211.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.926 |
| C2 |
0.000 |
0.000 |
-1.092 |
| N3 |
0.000 |
0.000 |
0.110 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 3.0182 | 1.8160 |
C2 | 3.0182 | | 1.2022 | N3 | 1.8160 | 1.2022 | |
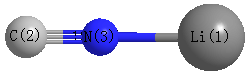 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.383 |
|
|
|
| 2 |
C |
-0.402 |
|
|
|
| 3 |
N |
0.019 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
9.079 |
9.079 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.157 |
0.000 |
0.000 |
| y |
0.000 |
-14.157 |
0.000 |
| z |
0.000 |
0.000 |
-1.232 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.462 |
0.000 |
0.000 |
| y |
0.000 |
-6.462 |
0.000 |
| z |
0.000 |
0.000 |
12.925 |
|
| Polar |
|---|
| 3z2-r2 | 25.849 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.527 |
0.000 |
0.000 |
| y |
0.000 |
2.527 |
0.000 |
| z |
0.000 |
0.000 |
4.860 |
<r2> (average value of r
2) Å
2
| <r2> |
14.692 |
| (<r2>)1/2 |
3.833 |
