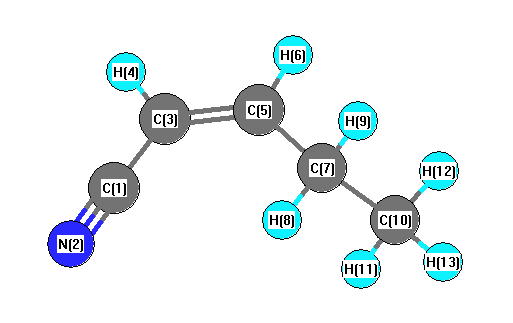
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3193 |
3084 |
7.68 |
|
|
|
| 2 |
A |
3147 |
3039 |
22.25 |
|
|
|
| 3 |
A |
3126 |
3019 |
41.66 |
|
|
|
| 4 |
A |
3119 |
3012 |
37.34 |
|
|
|
| 5 |
A |
3103 |
2997 |
5.41 |
|
|
|
| 6 |
A |
3045 |
2941 |
35.71 |
|
|
|
| 7 |
A |
3039 |
2935 |
26.73 |
|
|
|
| 8 |
A |
2318 |
2238 |
30.85 |
|
|
|
| 9 |
A |
1687 |
1629 |
17.07 |
|
|
|
| 10 |
A |
1506 |
1454 |
8.75 |
|
|
|
| 11 |
A |
1497 |
1446 |
7.40 |
|
|
|
| 12 |
A |
1483 |
1432 |
6.17 |
|
|
|
| 13 |
A |
1408 |
1360 |
1.80 |
|
|
|
| 14 |
A |
1399 |
1351 |
0.15 |
|
|
|
| 15 |
A |
1322 |
1277 |
4.00 |
|
|
|
| 16 |
A |
1282 |
1238 |
0.03 |
|
|
|
| 17 |
A |
1233 |
1190 |
0.62 |
|
|
|
| 18 |
A |
1127 |
1088 |
1.55 |
|
|
|
| 19 |
A |
1079 |
1042 |
5.29 |
|
|
|
| 20 |
A |
1017 |
982 |
4.94 |
|
|
|
| 21 |
A |
990 |
956 |
0.93 |
|
|
|
| 22 |
A |
958 |
925 |
4.90 |
|
|
|
| 23 |
A |
859 |
829 |
4.61 |
|
|
|
| 24 |
A |
791 |
764 |
13.76 |
|
|
|
| 25 |
A |
761 |
735 |
45.77 |
|
|
|
| 26 |
A |
652 |
629 |
1.64 |
|
|
|
| 27 |
A |
549 |
530 |
1.39 |
|
|
|
| 28 |
A |
385 |
372 |
0.11 |
|
|
|
| 29 |
A |
359 |
347 |
0.47 |
|
|
|
| 30 |
A |
236 |
228 |
0.26 |
|
|
|
| 31 |
A |
205 |
198 |
2.62 |
|
|
|
| 32 |
A |
143 |
138 |
4.06 |
|
|
|
| 33 |
A |
50 |
48 |
0.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23533.2 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 22726.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.647 |
|
|
|
| 2 |
N |
0.373 |
|
|
|
| 3 |
C |
-0.225 |
|
|
|
| 4 |
H |
0.306 |
|
|
|
| 5 |
C |
-0.294 |
|
|
|
| 6 |
H |
0.257 |
|
|
|
| 7 |
C |
-0.063 |
|
|
|
| 8 |
H |
0.197 |
|
|
|
| 9 |
H |
0.119 |
|
|
|
| 10 |
C |
-0.520 |
|
|
|
| 11 |
H |
0.156 |
|
|
|
| 12 |
H |
0.155 |
|
|
|
| 13 |
H |
0.184 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.561 |
2.652 |
0.081 |
4.441 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.696 |
-5.813 |
1.169 |
| y |
-5.813 |
-35.488 |
-0.791 |
| z |
1.169 |
-0.791 |
-36.881 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.512 |
-5.813 |
1.169 |
| y |
-5.813 |
4.301 |
-0.791 |
| z |
1.169 |
-0.791 |
2.211 |
|
| Polar |
|---|
| 3z2-r2 | 4.421 |
|---|
| x2-y2 | -7.208 |
|---|
| xy | -5.813 |
|---|
| xz | 1.169 |
|---|
| yz | -0.791 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.127 |
1.145 |
0.509 |
| y |
1.145 |
8.274 |
-0.189 |
| z |
0.509 |
-0.189 |
5.752 |
<r2> (average value of r
2) Å
2
| <r2> |
161.691 |
| (<r2>)1/2 |
12.716 |