return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSSCH3 (dimethyl trisulfide)

using model chemistry: MP2/CEP-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2/CEP-31G*
 hartrees
Energy at 0K-44.900424
Energy at 298.15K-44.906692
Nuclear repulsion energy89.240266
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/CEP-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3219 3056 15.71      
2 A 3205 3043 0.37      
3 A 3092 2936 7.11      
4 A 1501 1425 8.58      
5 A 1480 1405 8.59      
6 A 1393 1323 0.58      
7 A 1008 957 1.83      
8 A 1005 954 5.64      
9 A 738 701 0.06      
10 A 511 485 0.41      
11 A 273 259 0.53      
12 A 197 187 1.40      
13 A 161 153 1.81      
14 A 64 61 2.76      
15 B 3218 3056 5.94      
16 B 3204 3042 14.98      
17 B 3092 2935 48.00      
18 B 1499 1424 11.32      
19 B 1480 1405 11.15      
20 B 1396 1325 7.76      
21 B 1007 956 4.08      
22 B 1000 949 14.60      
23 B 737 700 1.62      
24 B 509 483 17.76      
25 B 279 265 0.95      
26 B 192 183 0.72      
27 B 94 89 4.03      

Unscaled Zero Point Vibrational Energy (zpe) 17775.6 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 16876.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/CEP-31G*
ABC
0.14628 0.04548 0.04524

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/CEP-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 1.095
S2 0.000 1.663 -0.158
S3 0.000 -1.663 -0.158
C4 1.762 1.734 -0.656
C5 -1.762 -1.734 -0.656
H6 1.865 2.616 -1.313
H7 -1.865 -2.616 -1.313
H8 2.409 1.850 0.228
H9 2.036 0.825 -1.216
H10 -2.409 -1.850 0.228
H11 -2.036 -0.825 -1.216

Atom - Atom Distances (Å)
  S1 S2 S3 C4 C5 H6 H7 H8 H9 H10 H11
S12.08172.08173.02963.02964.01504.01503.15833.18843.15833.1884
S22.08173.32541.83253.85912.39214.80852.44652.44234.27633.3845
S32.08173.32543.85911.83254.80852.39214.27633.38452.44652.4423
C43.02961.83253.85914.94471.10435.70221.10131.10215.56964.6139
C53.02963.85911.83254.94475.70221.10435.56964.61391.10131.1021
H64.01502.39214.80851.10435.70226.42611.80451.80146.37055.2030
H74.01504.80852.39215.70221.10436.42616.37055.20301.80451.8014
H83.15832.44654.27631.10135.56961.80456.37051.80926.07395.3845
H93.18842.44233.38451.10214.61391.80145.20301.80925.38454.3933
H103.15834.27632.44655.56961.10136.37051.80456.07395.38451.8092
H113.18843.38452.44234.61391.10215.20301.80145.38454.39331.8092

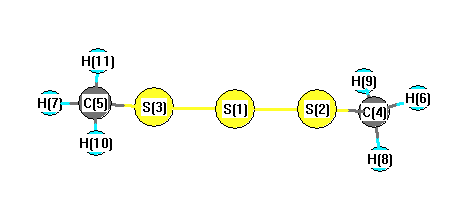
picture of dimethyl trisulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 C4 101.235 S1 S3 C5 101.235
S2 S1 S3 106.014 S2 C4 H6 106.428
S2 C4 H8 110.517 S2 C4 H9 110.163
S3 C5 H7 106.428 S3 C5 H10 110.517
S3 C5 H11 110.163 H6 C4 H8 109.800
H6 C4 H9 109.462 H7 C5 H10 109.800
H7 C5 H11 109.462 H8 C4 H9 110.388
H10 C5 H11 110.388
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability