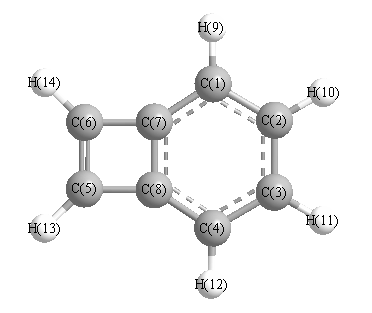
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3236 |
3125 |
10.15 |
|
|
|
| 2 |
A1 |
3194 |
3084 |
15.14 |
|
|
|
| 3 |
A1 |
3174 |
3065 |
18.90 |
|
|
|
| 4 |
A1 |
1699 |
1641 |
1.22 |
|
|
|
| 5 |
A1 |
1564 |
1511 |
4.34 |
|
|
|
| 6 |
A1 |
1506 |
1454 |
3.57 |
|
|
|
| 7 |
A1 |
1431 |
1382 |
11.53 |
|
|
|
| 8 |
A1 |
1166 |
1126 |
0.00 |
|
|
|
| 9 |
A1 |
1106 |
1068 |
0.26 |
|
|
|
| 10 |
A1 |
1050 |
1014 |
0.95 |
|
|
|
| 11 |
A1 |
978 |
944 |
5.80 |
|
|
|
| 12 |
A1 |
808 |
780 |
0.56 |
|
|
|
| 13 |
A1 |
532 |
513 |
0.06 |
|
|
|
| 14 |
A2 |
984 |
950 |
0.00 |
|
|
|
| 15 |
A2 |
931 |
899 |
0.00 |
|
|
|
| 16 |
A2 |
889 |
859 |
0.00 |
|
|
|
| 17 |
A2 |
782 |
755 |
0.00 |
|
|
|
| 18 |
A2 |
569 |
550 |
0.00 |
|
|
|
| 19 |
A2 |
294 |
284 |
0.00 |
|
|
|
| 20 |
B1 |
947 |
915 |
19.13 |
|
|
|
| 21 |
B1 |
772 |
746 |
187.45 |
|
|
|
| 22 |
B1 |
739 |
714 |
3.66 |
|
|
|
| 23 |
B1 |
364 |
352 |
4.08 |
|
|
|
| 24 |
B1 |
241 |
233 |
13.69 |
|
|
|
| 25 |
B2 |
3200 |
3091 |
0.93 |
|
|
|
| 26 |
B2 |
3187 |
3077 |
28.95 |
|
|
|
| 27 |
B2 |
3152 |
3044 |
1.07 |
|
|
|
| 28 |
B2 |
1649 |
1592 |
0.63 |
|
|
|
| 29 |
B2 |
1443 |
1393 |
3.64 |
|
|
|
| 30 |
B2 |
1284 |
1240 |
5.81 |
|
|
|
| 31 |
B2 |
1230 |
1187 |
10.81 |
|
|
|
| 32 |
B2 |
1082 |
1045 |
2.56 |
|
|
|
| 33 |
B2 |
1017 |
982 |
0.14 |
|
|
|
| 34 |
B2 |
853 |
824 |
1.51 |
|
|
|
| 35 |
B2 |
634 |
612 |
0.86 |
|
|
|
| 36 |
B2 |
405 |
391 |
4.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24044.8 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 23220.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.093 |
|
|
|
| 2 |
C |
-0.339 |
|
|
|
| 3 |
C |
-0.339 |
|
|
|
| 4 |
C |
-0.093 |
|
|
|
| 5 |
C |
-0.295 |
|
|
|
| 6 |
C |
-0.295 |
|
|
|
| 7 |
C |
-0.230 |
|
|
|
| 8 |
C |
-0.230 |
|
|
|
| 9 |
H |
0.330 |
|
|
|
| 10 |
H |
0.289 |
|
|
|
| 11 |
H |
0.289 |
|
|
|
| 12 |
H |
0.330 |
|
|
|
| 13 |
H |
0.338 |
|
|
|
| 14 |
H |
0.338 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.582 |
0.582 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.826 |
0.000 |
0.000 |
| y |
0.000 |
-39.905 |
0.000 |
| z |
0.000 |
0.000 |
-39.454 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.146 |
0.000 |
0.000 |
| y |
0.000 |
5.235 |
0.000 |
| z |
0.000 |
0.000 |
5.911 |
|
| Polar |
|---|
| 3z2-r2 | 11.822 |
|---|
| x2-y2 | -10.921 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.070 |
0.000 |
0.000 |
| y |
0.000 |
13.728 |
0.000 |
| z |
0.000 |
0.000 |
16.857 |
<r2> (average value of r
2) Å
2
| <r2> |
177.918 |
| (<r2>)1/2 |
13.339 |