Jump to
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -26.624087 |
| Energy at 298.15K | -26.626347 |
| Nuclear repulsion energy | 31.174912 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3530 |
3409 |
31.34 |
|
|
|
| 2 |
A' |
2337 |
2257 |
118.86 |
|
|
|
| 3 |
A' |
1641 |
1585 |
47.43 |
|
|
|
| 4 |
A' |
1065 |
1029 |
6.48 |
|
|
|
| 5 |
A' |
647 |
625 |
241.63 |
|
|
|
| 6 |
A' |
476 |
460 |
29.88 |
|
|
|
| 7 |
A" |
3629 |
3505 |
46.58 |
|
|
|
| 8 |
A" |
1193 |
1152 |
0.29 |
|
|
|
| 9 |
A" |
392 |
379 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7455.9 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 7200.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.221 |
0.000 |
| N2 |
-0.008 |
1.404 |
0.000 |
| N3 |
0.090 |
-1.145 |
0.000 |
| H4 |
-0.289 |
-1.570 |
0.848 |
| H5 |
-0.289 |
-1.570 |
-0.848 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1833 | 1.3685 | 2.0023 | 2.0023 |
N2 | 1.1833 | | 2.5507 | 3.1053 | 3.1053 | N3 | 1.3685 | 2.5507 | | 1.0215 | 1.0215 | H4 | 2.0023 | 3.1053 | 1.0215 | | 1.6957 | H5 | 2.0023 | 3.1053 | 1.0215 | 1.6957 | |
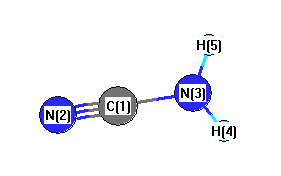 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
113.007 |
|
C1 |
N3 |
H5 |
113.007 |
| N2 |
C1 |
N3 |
176.592 |
|
H4 |
N3 |
H5 |
112.193 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.616 |
|
|
|
| 2 |
N |
0.305 |
|
|
|
| 3 |
N |
-0.368 |
|
|
|
| 4 |
H |
0.340 |
|
|
|
| 5 |
H |
0.340 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.278 |
-4.430 |
0.000 |
4.611 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.347 |
2.378 |
0.000 |
| y |
2.378 |
-18.340 |
0.000 |
| z |
0.000 |
0.000 |
-14.990 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.681 |
2.378 |
0.000 |
| y |
2.378 |
-1.672 |
0.000 |
| z |
0.000 |
0.000 |
3.353 |
|
| Polar |
|---|
| 3z2-r2 | 6.706 |
|---|
| x2-y2 | -0.006 |
|---|
| xy | 2.378 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.986 |
0.001 |
0.000 |
| y |
0.001 |
5.022 |
0.000 |
| z |
0.000 |
0.000 |
2.401 |
<r2> (average value of r
2) Å
2
| <r2> |
33.939 |
| (<r2>)1/2 |
5.826 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -26.621969 |
| Energy at 298.15K | |
| HF Energy | -26.621969 |
| Nuclear repulsion energy | 31.233119 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3609 |
3486 |
62.17 |
|
|
|
| 2 |
A1 |
2335 |
2255 |
154.77 |
|
|
|
| 3 |
A1 |
1601 |
1546 |
53.75 |
|
|
|
| 4 |
A1 |
1110 |
1072 |
11.99 |
|
|
|
| 5 |
B1 |
505 |
488 |
0.41 |
|
|
|
| 6 |
B1 |
477i |
461i |
328.40 |
|
|
|
| 7 |
B2 |
3731 |
3603 |
87.29 |
|
|
|
| 8 |
B2 |
1096 |
1059 |
2.67 |
|
|
|
| 9 |
B2 |
385 |
372 |
1.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6948.2 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 6709.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.221 |
| N2 |
0.000 |
0.000 |
1.407 |
| N3 |
0.000 |
0.000 |
-1.128 |
| H4 |
0.000 |
0.877 |
-1.639 |
| H5 |
0.000 |
-0.877 |
-1.639 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1852 | 1.3496 | 2.0567 | 2.0567 |
N2 | 1.1852 | | 2.5348 | 3.1693 | 3.1693 | N3 | 1.3496 | 2.5348 | | 1.0151 | 1.0151 | H4 | 2.0567 | 3.1693 | 1.0151 | | 1.7548 | H5 | 2.0567 | 3.1693 | 1.0151 | 1.7548 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
120.198 |
|
C1 |
N3 |
H5 |
120.198 |
| N2 |
C1 |
N3 |
180.000 |
|
H4 |
N3 |
H5 |
119.605 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.632 |
|
|
|
| 2 |
N |
0.285 |
|
|
|
| 3 |
N |
-0.366 |
|
|
|
| 4 |
H |
0.357 |
|
|
|
| 5 |
H |
0.357 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.901 |
4.901 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.725 |
0.000 |
0.000 |
| y |
0.000 |
-14.644 |
0.000 |
| z |
0.000 |
0.000 |
-17.288 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.759 |
0.000 |
0.000 |
| y |
0.000 |
3.363 |
0.000 |
| z |
0.000 |
0.000 |
-0.604 |
|
| Polar |
|---|
| 3z2-r2 | -1.208 |
|---|
| x2-y2 | -4.081 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.915 |
0.000 |
0.000 |
| y |
0.000 |
2.292 |
0.000 |
| z |
0.000 |
0.000 |
5.096 |
<r2> (average value of r
2) Å
2
| <r2> |
33.911 |
| (<r2>)1/2 |
5.823 |
