Jump to
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -33.407158 |
| Energy at 298.15K | -33.411011 |
| HF Energy | -33.407158 |
| Nuclear repulsion energy | 54.161806 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3597 |
3473 |
15.94 |
|
|
|
| 2 |
A |
3508 |
3388 |
2.98 |
|
|
|
| 3 |
A |
2362 |
2281 |
0.73 |
|
|
|
| 4 |
A |
1662 |
1605 |
14.60 |
|
|
|
| 5 |
A |
1192 |
1151 |
0.07 |
|
|
|
| 6 |
A |
812 |
785 |
3.59 |
|
|
|
| 7 |
A |
589 |
568 |
149.14 |
|
|
|
| 8 |
A |
409 |
395 |
22.24 |
|
|
|
| 9 |
A |
384 |
371 |
9.99 |
|
|
|
| 10 |
A |
166 |
160 |
27.51 |
|
|
|
| 11 |
B |
3596 |
3473 |
15.01 |
|
|
|
| 12 |
B |
3511 |
3390 |
4.60 |
|
|
|
| 13 |
B |
1661 |
1604 |
23.34 |
|
|
|
| 14 |
B |
1350 |
1303 |
113.43 |
|
|
|
| 15 |
B |
1192 |
1151 |
0.19 |
|
|
|
| 16 |
B |
654 |
631 |
348.26 |
|
|
|
| 17 |
B |
381 |
368 |
18.24 |
|
|
|
| 18 |
B |
167 |
161 |
28.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13595.8 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 13129.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.007 |
0.628 |
0.016 |
| C2 |
-0.007 |
-0.628 |
0.016 |
| N3 |
-0.007 |
2.000 |
-0.064 |
| N4 |
0.007 |
-2.000 |
-0.064 |
| H5 |
-0.335 |
2.479 |
0.775 |
| H6 |
0.849 |
2.421 |
-0.426 |
| H7 |
0.335 |
-2.479 |
0.775 |
| H8 |
-0.849 |
-2.421 |
-0.426 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2569 | 1.3740 | 2.6296 | 2.0294 | 2.0292 | 3.2157 | 3.1980 |
C2 | 1.2569 | | 2.6296 | 1.3740 | 3.2157 | 3.1980 | 2.0294 | 2.0292 | N3 | 1.3740 | 2.6296 | | 4.0001 | 1.0202 | 1.0204 | 4.5698 | 4.5151 | N4 | 2.6296 | 1.3740 | 4.0001 | | 4.5698 | 4.5151 | 1.0202 | 1.0204 | H5 | 2.0294 | 3.2157 | 1.0202 | 4.5698 | | 1.6875 | 5.0032 | 5.0713 | H6 | 2.0292 | 3.1980 | 1.0204 | 4.5151 | 1.6875 | | 5.0713 | 5.1313 | H7 | 3.2157 | 2.0294 | 4.5698 | 1.0202 | 5.0032 | 5.0713 | | 1.6875 | H8 | 3.1980 | 2.0292 | 4.5151 | 1.0204 | 5.0713 | 5.1313 | 1.6875 | |
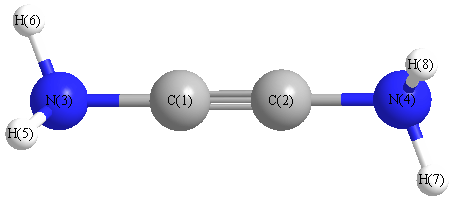 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
176.472 |
|
C1 |
N3 |
H5 |
115.118 |
| C1 |
N3 |
H6 |
115.087 |
|
C2 |
C1 |
N3 |
176.472 |
| C2 |
N4 |
H7 |
115.118 |
|
C2 |
N4 |
H8 |
115.087 |
| H5 |
N3 |
H6 |
111.582 |
|
H7 |
N4 |
H8 |
111.582 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.420 |
|
|
|
| 2 |
C |
-0.420 |
|
|
|
| 3 |
N |
-0.230 |
|
|
|
| 4 |
N |
-0.230 |
|
|
|
| 5 |
H |
0.324 |
|
|
|
| 6 |
H |
0.326 |
|
|
|
| 7 |
H |
0.324 |
|
|
|
| 8 |
H |
0.326 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.641 |
1.641 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.586 |
4.787 |
0.000 |
| y |
4.787 |
-13.089 |
0.000 |
| z |
0.000 |
0.000 |
-23.779 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.152 |
4.787 |
0.000 |
| y |
4.787 |
10.594 |
0.000 |
| z |
0.000 |
0.000 |
-5.441 |
|
| Polar |
|---|
| 3z2-r2 | -10.883 |
|---|
| x2-y2 | -10.497 |
|---|
| xy | 4.787 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.221 |
0.163 |
0.000 |
| y |
0.163 |
9.461 |
0.000 |
| z |
0.000 |
0.000 |
3.217 |
<r2> (average value of r
2) Å
2
| <r2> |
82.609 |
| (<r2>)1/2 |
9.089 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -33.403957 |
| Energy at 298.15K | |
| HF Energy | -33.403957 |
| Nuclear repulsion energy | 54.305361 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3568 |
3446 |
0.00 |
|
|
|
| 2 |
A1 |
2328 |
2249 |
0.00 |
|
|
|
| 3 |
A1 |
1635 |
1579 |
0.00 |
|
|
|
| 4 |
A1 |
838 |
809 |
0.00 |
|
|
|
| 5 |
B1 |
546 |
528 |
0.00 |
|
|
|
| 6 |
B2 |
3574 |
3452 |
11.60 |
|
|
|
| 7 |
B2 |
1643 |
1587 |
35.88 |
|
|
|
| 8 |
B2 |
1418 |
1369 |
159.75 |
|
|
|
| 9 |
E |
3672 |
3546 |
30.66 |
|
|
|
| 9 |
E |
3672 |
3546 |
30.66 |
|
|
|
| 10 |
E |
1113 |
1075 |
2.76 |
|
|
|
| 10 |
E |
1113 |
1075 |
2.76 |
|
|
|
| 11 |
E |
392 |
379 |
2.81 |
|
|
|
| 11 |
E |
392 |
379 |
2.81 |
|
|
|
| 12 |
E |
50 |
48 |
38.37 |
|
|
|
| 12 |
E |
50 |
48 |
38.37 |
|
|
|
| 13 |
E |
429i |
414i |
274.66 |
|
|
|
| 13 |
E |
429i |
414i |
274.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12573.4 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 12142.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.628 |
| C2 |
0.000 |
0.000 |
-0.628 |
| N3 |
0.000 |
0.000 |
1.979 |
| N4 |
0.000 |
0.000 |
-1.979 |
| H5 |
0.000 |
0.868 |
2.508 |
| H6 |
0.000 |
-0.868 |
2.508 |
| H7 |
0.868 |
0.000 |
-2.508 |
| H8 |
-0.868 |
0.000 |
-2.508 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2554 | 1.3517 | 2.6070 | 2.0708 | 2.0708 | 3.2534 | 3.2534 |
C2 | 1.2554 | | 2.6070 | 1.3517 | 3.2534 | 3.2534 | 2.0708 | 2.0708 | N3 | 1.3517 | 2.6070 | | 3.9587 | 1.0163 | 1.0163 | 4.5703 | 4.5703 | N4 | 2.6070 | 1.3517 | 3.9587 | | 4.5703 | 4.5703 | 1.0163 | 1.0163 | H5 | 2.0708 | 3.2534 | 1.0163 | 4.5703 | | 1.7363 | 5.1636 | 5.1636 | H6 | 2.0708 | 3.2534 | 1.0163 | 4.5703 | 1.7363 | | 5.1636 | 5.1636 | H7 | 3.2534 | 2.0708 | 4.5703 | 1.0163 | 5.1636 | 5.1636 | | 1.7363 | H8 | 3.2534 | 2.0708 | 4.5703 | 1.0163 | 5.1636 | 5.1636 | 1.7363 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.326 |
| C1 |
N3 |
H6 |
121.326 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.326 |
|
C2 |
N4 |
H8 |
121.326 |
| H5 |
N3 |
H6 |
117.349 |
|
H7 |
N4 |
H8 |
117.349 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.434 |
|
|
|
| 2 |
C |
-0.434 |
|
|
|
| 3 |
N |
-0.217 |
|
|
|
| 4 |
N |
-0.217 |
|
|
|
| 5 |
H |
0.326 |
|
|
|
| 6 |
H |
0.326 |
|
|
|
| 7 |
H |
0.326 |
|
|
|
| 8 |
H |
0.326 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.873 |
0.000 |
0.000 |
| y |
0.000 |
-23.873 |
0.000 |
| z |
0.000 |
0.000 |
-9.724 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.074 |
0.000 |
0.000 |
| y |
0.000 |
-7.074 |
0.000 |
| z |
0.000 |
0.000 |
14.149 |
|
| Polar |
|---|
| 3z2-r2 | 28.297 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.046 |
0.000 |
0.000 |
| y |
0.000 |
3.046 |
0.000 |
| z |
0.000 |
0.000 |
9.912 |
<r2> (average value of r
2) Å
2
| <r2> |
82.465 |
| (<r2>)1/2 |
9.081 |
