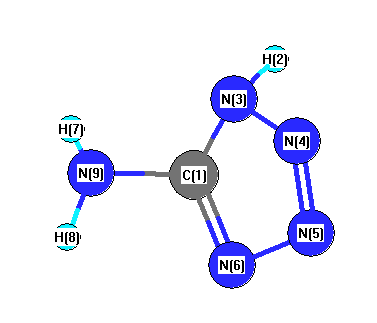
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3647 |
3522 |
72.33 |
|
|
|
| 2 |
A |
3638 |
3513 |
29.42 |
|
|
|
| 3 |
A |
3537 |
3416 |
23.18 |
|
|
|
| 4 |
A |
1672 |
1614 |
151.67 |
|
|
|
| 5 |
A |
1596 |
1541 |
136.37 |
|
|
|
| 6 |
A |
1479 |
1429 |
29.49 |
|
|
|
| 7 |
A |
1388 |
1341 |
23.99 |
|
|
|
| 8 |
A |
1291 |
1247 |
10.02 |
|
|
|
| 9 |
A |
1136 |
1097 |
10.12 |
|
|
|
| 10 |
A |
1106 |
1068 |
13.73 |
|
|
|
| 11 |
A |
1055 |
1019 |
15.93 |
|
|
|
| 12 |
A |
1014 |
980 |
22.73 |
|
|
|
| 13 |
A |
990 |
956 |
1.52 |
|
|
|
| 14 |
A |
771 |
745 |
150.90 |
|
|
|
| 15 |
A |
717 |
692 |
1.96 |
|
|
|
| 16 |
A |
700 |
676 |
16.91 |
|
|
|
| 17 |
A |
674 |
651 |
151.94 |
|
|
|
| 18 |
A |
503 |
486 |
82.29 |
|
|
|
| 19 |
A |
379 |
366 |
8.61 |
|
|
|
| 20 |
A |
300 |
290 |
4.43 |
|
|
|
| 21 |
A |
250 |
241 |
55.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13921.5 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 13444.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.611 |
|
|
|
| 2 |
H |
0.359 |
|
|
|
| 3 |
N |
-0.007 |
|
|
|
| 4 |
N |
-0.071 |
|
|
|
| 5 |
N |
-0.088 |
|
|
|
| 6 |
N |
0.126 |
|
|
|
| 7 |
H |
0.316 |
|
|
|
| 8 |
H |
0.335 |
|
|
|
| 9 |
N |
-0.359 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.949 |
3.562 |
1.032 |
6.184 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.261 |
1.138 |
-3.366 |
| y |
1.138 |
-32.103 |
0.603 |
| z |
-3.366 |
0.603 |
-34.770 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.825 |
1.138 |
-3.366 |
| y |
1.138 |
2.412 |
0.603 |
| z |
-3.366 |
0.603 |
-1.587 |
|
| Polar |
|---|
| 3z2-r2 | -3.175 |
|---|
| x2-y2 | -2.158 |
|---|
| xy | 1.138 |
|---|
| xz | -3.366 |
|---|
| yz | 0.603 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.195 |
-0.042 |
0.017 |
| y |
-0.042 |
6.714 |
0.055 |
| z |
0.017 |
0.055 |
3.392 |
<r2> (average value of r
2) Å
2
| <r2> |
98.700 |
| (<r2>)1/2 |
9.935 |