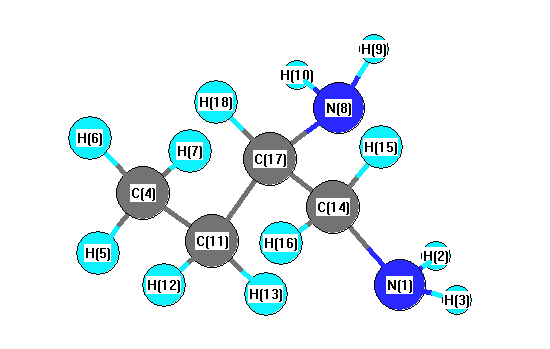
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3566 |
3444 |
1.02 |
|
|
|
| 2 |
A |
3562 |
3440 |
0.32 |
|
|
|
| 3 |
A |
3476 |
3357 |
4.33 |
|
|
|
| 4 |
A |
3474 |
3355 |
3.46 |
|
|
|
| 5 |
A |
3122 |
3015 |
64.24 |
|
|
|
| 6 |
A |
3100 |
2993 |
63.18 |
|
|
|
| 7 |
A |
3086 |
2980 |
34.22 |
|
|
|
| 8 |
A |
3062 |
2957 |
68.48 |
|
|
|
| 9 |
A |
3049 |
2945 |
11.66 |
|
|
|
| 10 |
A |
3029 |
2925 |
38.85 |
|
|
|
| 11 |
A |
2945 |
2844 |
145.42 |
|
|
|
| 12 |
A |
2924 |
2824 |
76.30 |
|
|
|
| 13 |
A |
1675 |
1617 |
41.43 |
|
|
|
| 14 |
A |
1655 |
1599 |
42.42 |
|
|
|
| 15 |
A |
1509 |
1457 |
8.58 |
|
|
|
| 16 |
A |
1508 |
1457 |
5.15 |
|
|
|
| 17 |
A |
1500 |
1448 |
5.59 |
|
|
|
| 18 |
A |
1479 |
1428 |
2.29 |
|
|
|
| 19 |
A |
1431 |
1382 |
0.77 |
|
|
|
| 20 |
A |
1422 |
1373 |
10.76 |
|
|
|
| 21 |
A |
1410 |
1362 |
3.70 |
|
|
|
| 22 |
A |
1379 |
1331 |
14.49 |
|
|
|
| 23 |
A |
1334 |
1288 |
2.59 |
|
|
|
| 24 |
A |
1298 |
1253 |
1.13 |
|
|
|
| 25 |
A |
1279 |
1235 |
1.53 |
|
|
|
| 26 |
A |
1251 |
1209 |
1.60 |
|
|
|
| 27 |
A |
1184 |
1144 |
6.76 |
|
|
|
| 28 |
A |
1141 |
1102 |
8.43 |
|
|
|
| 29 |
A |
1091 |
1054 |
1.49 |
|
|
|
| 30 |
A |
1067 |
1030 |
5.51 |
|
|
|
| 31 |
A |
1049 |
1013 |
3.43 |
|
|
|
| 32 |
A |
1002 |
967 |
3.57 |
|
|
|
| 33 |
A |
967 |
934 |
39.00 |
|
|
|
| 34 |
A |
926 |
894 |
65.39 |
|
|
|
| 35 |
A |
918 |
886 |
38.15 |
|
|
|
| 36 |
A |
871 |
841 |
111.40 |
|
|
|
| 37 |
A |
811 |
783 |
12.21 |
|
|
|
| 38 |
A |
748 |
723 |
0.20 |
|
|
|
| 39 |
A |
610 |
589 |
19.56 |
|
|
|
| 40 |
A |
430 |
415 |
7.02 |
|
|
|
| 41 |
A |
372 |
360 |
5.49 |
|
|
|
| 42 |
A |
333 |
321 |
42.14 |
|
|
|
| 43 |
A |
275 |
265 |
51.67 |
|
|
|
| 44 |
A |
273 |
264 |
18.06 |
|
|
|
| 45 |
A |
232 |
224 |
7.60 |
|
|
|
| 46 |
A |
211 |
204 |
2.51 |
|
|
|
| 47 |
A |
141 |
136 |
5.22 |
|
|
|
| 48 |
A |
85 |
82 |
0.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36629.2 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 35372.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.542 |
|
|
|
| 2 |
H |
0.269 |
|
|
|
| 3 |
H |
0.253 |
|
|
|
| 4 |
C |
-0.443 |
|
|
|
| 5 |
H |
0.184 |
|
|
|
| 6 |
H |
0.133 |
|
|
|
| 7 |
H |
0.121 |
|
|
|
| 8 |
N |
-0.589 |
|
|
|
| 9 |
H |
0.250 |
|
|
|
| 10 |
H |
0.253 |
|
|
|
| 11 |
C |
-0.250 |
|
|
|
| 12 |
H |
0.114 |
|
|
|
| 13 |
H |
0.172 |
|
|
|
| 14 |
C |
-0.261 |
|
|
|
| 15 |
H |
0.137 |
|
|
|
| 16 |
H |
0.131 |
|
|
|
| 17 |
C |
-0.084 |
|
|
|
| 18 |
H |
0.153 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.028 |
1.485 |
1.493 |
2.106 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.356 |
0.570 |
0.970 |
| y |
0.570 |
-38.338 |
-0.338 |
| z |
0.970 |
-0.338 |
-40.326 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.976 |
0.570 |
0.970 |
| y |
0.570 |
-0.497 |
-0.338 |
| z |
0.970 |
-0.338 |
-3.479 |
|
| Polar |
|---|
| 3z2-r2 | -6.957 |
|---|
| x2-y2 | 2.982 |
|---|
| xy | 0.570 |
|---|
| xz | 0.970 |
|---|
| yz | -0.338 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.003 |
-0.095 |
0.183 |
| y |
-0.095 |
8.196 |
-0.094 |
| z |
0.183 |
-0.094 |
7.809 |
<r2> (average value of r
2) Å
2
| <r2> |
174.478 |
| (<r2>)1/2 |
13.209 |