Jump to
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -70.961149 |
| Energy at 298.15K | -70.965928 |
| Nuclear repulsion energy | 123.877878 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3734 |
3606 |
70.98 |
|
|
|
| 2 |
A' |
3727 |
3599 |
70.90 |
|
|
|
| 3 |
A' |
3589 |
3466 |
62.92 |
|
|
|
| 4 |
A' |
1812 |
1750 |
36.21 |
|
|
|
| 5 |
A' |
1791 |
1730 |
614.74 |
|
|
|
| 6 |
A' |
1597 |
1543 |
111.18 |
|
|
|
| 7 |
A' |
1411 |
1363 |
12.28 |
|
|
|
| 8 |
A' |
1303 |
1258 |
43.31 |
|
|
|
| 9 |
A' |
1190 |
1149 |
328.12 |
|
|
|
| 10 |
A' |
1075 |
1038 |
3.26 |
|
|
|
| 11 |
A' |
767 |
740 |
8.76 |
|
|
|
| 12 |
A' |
596 |
576 |
66.94 |
|
|
|
| 13 |
A' |
513 |
496 |
0.17 |
|
|
|
| 14 |
A' |
404 |
390 |
4.23 |
|
|
|
| 15 |
A' |
260 |
251 |
14.84 |
|
|
|
| 16 |
A" |
811 |
783 |
5.63 |
|
|
|
| 17 |
A" |
660 |
637 |
135.70 |
|
|
|
| 18 |
A" |
617 |
595 |
17.46 |
|
|
|
| 19 |
A" |
422 |
408 |
0.61 |
|
|
|
| 20 |
A" |
248 |
240 |
270.37 |
|
|
|
| 21 |
A" |
53 |
51 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13289.5 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 12833.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.766 |
0.000 |
| C2 |
-0.058 |
-0.800 |
0.000 |
| O3 |
-1.111 |
-1.432 |
0.000 |
| O4 |
1.049 |
1.401 |
0.000 |
| O5 |
-1.230 |
1.320 |
0.000 |
| N6 |
1.206 |
-1.333 |
0.000 |
| H7 |
1.309 |
-2.344 |
0.000 |
| H8 |
2.022 |
-0.725 |
0.000 |
| H9 |
-1.092 |
2.291 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5670 | 2.4625 | 1.2263 | 1.3488 | 2.4207 | 3.3744 | 2.5124 | 1.8753 |
C2 | 1.5670 | | 1.2285 | 2.4637 | 2.4225 | 1.3718 | 2.0621 | 2.0811 | 3.2591 | O3 | 2.4625 | 1.2285 | | 3.5625 | 2.7542 | 2.3199 | 2.5868 | 3.2122 | 3.7222 | O4 | 1.2263 | 2.4637 | 3.5625 | | 2.2801 | 2.7384 | 3.7543 | 2.3383 | 2.3183 | O5 | 1.3488 | 2.4225 | 2.7542 | 2.2801 | | 3.6017 | 4.4578 | 3.8415 | 0.9802 | N6 | 2.4207 | 1.3718 | 2.3199 | 2.7384 | 3.6017 | | 1.0166 | 1.0172 | 4.2908 | H7 | 3.3744 | 2.0621 | 2.5868 | 3.7543 | 4.4578 | 1.0166 | | 1.7691 | 5.2197 | H8 | 2.5124 | 2.0811 | 3.2122 | 2.3383 | 3.8415 | 1.0172 | 1.7691 | | 4.3349 | H9 | 1.8753 | 3.2591 | 3.7222 | 2.3183 | 0.9802 | 4.2908 | 5.2197 | 4.3349 | |
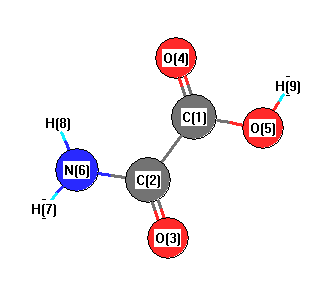 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
123.042 |
|
C1 |
C2 |
N6 |
110.742 |
| C1 |
O5 |
H9 |
106.175 |
|
C2 |
C1 |
O4 |
123.305 |
| C2 |
C1 |
O5 |
112.151 |
|
C2 |
N6 |
H7 |
118.647 |
| C2 |
N6 |
H8 |
120.463 |
|
O3 |
C2 |
N6 |
126.216 |
| O4 |
C1 |
O5 |
124.544 |
|
H7 |
N6 |
H8 |
120.890 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.164 |
|
|
|
| 2 |
C |
-0.126 |
|
|
|
| 3 |
O |
-0.083 |
|
|
|
| 4 |
O |
-0.121 |
|
|
|
| 5 |
O |
-0.257 |
|
|
|
| 6 |
N |
-0.372 |
|
|
|
| 7 |
H |
0.342 |
|
|
|
| 8 |
H |
0.363 |
|
|
|
| 9 |
H |
0.418 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.341 |
0.639 |
0.000 |
2.427 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.417 |
-7.650 |
-0.004 |
| y |
-7.650 |
-39.747 |
0.004 |
| z |
-0.004 |
0.004 |
-33.692 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
13.303 |
-7.650 |
-0.004 |
| y |
-7.650 |
-11.193 |
0.004 |
| z |
-0.004 |
0.004 |
-2.110 |
|
| Polar |
|---|
| 3z2-r2 | -4.221 |
|---|
| x2-y2 | 16.331 |
|---|
| xy | -7.650 |
|---|
| xz | -0.004 |
|---|
| yz | 0.004 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.324 |
-0.199 |
0.002 |
| y |
-0.199 |
6.861 |
0.002 |
| z |
0.002 |
0.002 |
3.072 |
<r2> (average value of r
2) Å
2
| <r2> |
117.120 |
| (<r2>)1/2 |
10.822 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -70.966272 |
| Energy at 298.15K | -70.971309 |
| Nuclear repulsion energy | 124.792448 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3717 |
3590 |
83.27 |
|
|
|
| 2 |
A' |
3593 |
3469 |
130.76 |
|
|
|
| 3 |
A' |
3581 |
3458 |
66.03 |
|
|
|
| 4 |
A' |
1848 |
1785 |
230.25 |
|
|
|
| 5 |
A' |
1778 |
1717 |
371.94 |
|
|
|
| 6 |
A' |
1599 |
1545 |
69.03 |
|
|
|
| 7 |
A' |
1434 |
1384 |
227.18 |
|
|
|
| 8 |
A' |
1329 |
1283 |
360.79 |
|
|
|
| 9 |
A' |
1192 |
1152 |
16.80 |
|
|
|
| 10 |
A' |
1077 |
1040 |
4.75 |
|
|
|
| 11 |
A' |
790 |
762 |
9.93 |
|
|
|
| 12 |
A' |
611 |
590 |
11.30 |
|
|
|
| 13 |
A' |
527 |
509 |
2.18 |
|
|
|
| 14 |
A' |
393 |
380 |
6.57 |
|
|
|
| 15 |
A' |
261 |
252 |
41.30 |
|
|
|
| 16 |
A" |
803 |
776 |
0.06 |
|
|
|
| 17 |
A" |
712 |
687 |
121.94 |
|
|
|
| 18 |
A" |
642 |
620 |
2.47 |
|
|
|
| 19 |
A" |
443 |
428 |
57.88 |
|
|
|
| 20 |
A" |
316 |
305 |
225.50 |
|
|
|
| 21 |
A" |
107 |
103 |
3.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13376.5 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 12917.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.008 |
-0.808 |
0.000 |
| C2 |
0.000 |
0.761 |
0.000 |
| O3 |
-1.082 |
1.367 |
0.000 |
| O4 |
1.030 |
-1.475 |
0.000 |
| O5 |
-1.240 |
-1.310 |
0.000 |
| N6 |
1.245 |
1.306 |
0.000 |
| H7 |
1.354 |
2.318 |
0.000 |
| H8 |
2.058 |
0.693 |
0.000 |
| H9 |
-1.842 |
-0.526 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5693 | 2.4326 | 1.2200 | 1.3455 | 2.4499 | 3.4032 | 2.5408 | 1.8712 |
C2 | 1.5693 | | 1.2396 | 2.4616 | 2.4138 | 1.3596 | 2.0629 | 2.0592 | 2.2471 | O3 | 2.4326 | 1.2396 | | 3.5400 | 2.6810 | 2.3278 | 2.6144 | 3.2111 | 2.0400 | O4 | 1.2200 | 2.4616 | 3.5400 | | 2.2764 | 2.7893 | 3.8061 | 2.3990 | 3.0242 | O5 | 1.3455 | 2.4138 | 2.6810 | 2.2764 | | 3.6087 | 4.4595 | 3.8587 | 0.9874 | N6 | 2.4499 | 1.3596 | 2.3278 | 2.7893 | 3.6087 | | 1.0171 | 1.0181 | 3.5902 | H7 | 3.4032 | 2.0629 | 2.6144 | 3.8061 | 4.4595 | 1.0171 | | 1.7709 | 4.2778 | H8 | 2.5408 | 2.0592 | 3.2111 | 2.3990 | 3.8587 | 1.0181 | 1.7709 | | 4.0859 | H9 | 1.8712 | 2.2471 | 2.0400 | 3.0242 | 0.9874 | 3.5902 | 4.2778 | 4.0859 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.543 |
|
C1 |
C2 |
N6 |
113.347 |
| C1 |
O5 |
H9 |
105.632 |
|
C2 |
C1 |
O4 |
123.409 |
| C2 |
C1 |
O5 |
111.586 |
|
C2 |
N6 |
H7 |
119.752 |
| C2 |
N6 |
H8 |
119.309 |
|
O3 |
C2 |
N6 |
127.110 |
| O4 |
C1 |
O5 |
125.005 |
|
H7 |
N6 |
H8 |
120.938 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.129 |
|
|
|
| 2 |
C |
-0.152 |
|
|
|
| 3 |
O |
-0.152 |
|
|
|
| 4 |
O |
-0.099 |
|
|
|
| 5 |
O |
-0.272 |
|
|
|
| 6 |
N |
-0.335 |
|
|
|
| 7 |
H |
0.347 |
|
|
|
| 8 |
H |
0.371 |
|
|
|
| 9 |
H |
0.422 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.626 |
2.752 |
0.000 |
3.197 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.859 |
7.600 |
0.000 |
| y |
7.600 |
-37.879 |
0.000 |
| z |
0.000 |
0.000 |
-33.631 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.896 |
7.600 |
0.000 |
| y |
7.600 |
-6.134 |
0.000 |
| z |
0.000 |
0.000 |
0.238 |
|
| Polar |
|---|
| 3z2-r2 | 0.476 |
|---|
| x2-y2 | 8.020 |
|---|
| xy | 7.600 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.204 |
-0.076 |
0.000 |
| y |
-0.076 |
5.864 |
0.000 |
| z |
0.000 |
0.000 |
3.059 |
<r2> (average value of r
2) Å
2
| <r2> |
115.071 |
| (<r2>)1/2 |
10.727 |
