Jump to
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -23.536501 |
| Energy at 298.15K | -23.540597 |
| HF Energy | -23.536501 |
| Nuclear repulsion energy | 35.427236 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3441 |
3323 |
3.57 |
|
|
|
| 2 |
A' |
3256 |
3145 |
17.20 |
|
|
|
| 3 |
A' |
3157 |
3049 |
8.45 |
|
|
|
| 4 |
A' |
3053 |
2949 |
59.98 |
|
|
|
| 5 |
A' |
1508 |
1456 |
8.67 |
|
|
|
| 6 |
A' |
1425 |
1376 |
2.60 |
|
|
|
| 7 |
A' |
1345 |
1299 |
35.50 |
|
|
|
| 8 |
A' |
1234 |
1191 |
4.95 |
|
|
|
| 9 |
A' |
1117 |
1079 |
18.61 |
|
|
|
| 10 |
A' |
964 |
931 |
6.66 |
|
|
|
| 11 |
A' |
465 |
449 |
12.04 |
|
|
|
| 12 |
A" |
1025 |
989 |
1.71 |
|
|
|
| 13 |
A" |
783 |
756 |
21.57 |
|
|
|
| 14 |
A" |
665 |
642 |
144.46 |
|
|
|
| 15 |
A" |
514 |
496 |
7.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11975.5 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 11564.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.427 |
0.000 |
| C2 |
1.172 |
-0.394 |
0.000 |
| N3 |
-1.214 |
-0.130 |
0.000 |
| H4 |
0.149 |
1.523 |
0.000 |
| H5 |
2.174 |
0.046 |
0.000 |
| H6 |
1.064 |
-1.484 |
0.000 |
| H7 |
-1.922 |
0.621 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4306 | 1.3352 | 1.1060 | 2.2071 | 2.1872 | 1.9313 |
C2 | 1.4306 | | 2.3997 | 2.1728 | 1.0949 | 1.0950 | 3.2554 | N3 | 1.3352 | 2.3997 | | 2.1417 | 3.3921 | 2.6500 | 1.0320 | H4 | 1.1060 | 2.1728 | 2.1417 | | 2.5065 | 3.1430 | 2.2582 | H5 | 2.2071 | 1.0949 | 3.3921 | 2.5065 | | 1.8900 | 4.1357 | H6 | 2.1872 | 1.0950 | 2.6500 | 3.1430 | 1.8900 | | 3.6532 | H7 | 1.9313 | 3.2554 | 1.0320 | 2.2582 | 4.1357 | 3.6532 | |
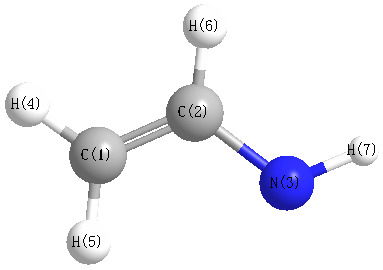 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.264 |
|
C1 |
C2 |
H6 |
119.405 |
| C1 |
N3 |
H7 |
108.674 |
|
C2 |
C1 |
N3 |
120.332 |
| C2 |
C1 |
H4 |
117.295 |
|
N3 |
C1 |
H4 |
122.373 |
| H5 |
C2 |
H6 |
119.331 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.317 |
|
|
|
| 2 |
C |
-0.397 |
|
|
|
| 3 |
N |
-0.172 |
|
|
|
| 4 |
H |
0.212 |
|
|
|
| 5 |
H |
0.207 |
|
|
|
| 6 |
H |
0.224 |
|
|
|
| 7 |
H |
0.243 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.213 |
1.752 |
0.000 |
2.131 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.731 |
-2.861 |
0.000 |
| y |
-2.861 |
-17.135 |
0.000 |
| z |
0.000 |
0.000 |
-20.275 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.974 |
-2.861 |
0.000 |
| y |
-2.861 |
1.368 |
0.000 |
| z |
0.000 |
0.000 |
-3.342 |
|
| Polar |
|---|
| 3z2-r2 | -6.684 |
|---|
| x2-y2 | 0.404 |
|---|
| xy | -2.861 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.555 |
-0.639 |
0.000 |
| y |
-0.639 |
3.920 |
0.000 |
| z |
0.000 |
0.000 |
2.158 |
<r2> (average value of r
2) Å
2
| <r2> |
40.043 |
| (<r2>)1/2 |
6.328 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -23.536062 |
| Energy at 298.15K | -23.540142 |
| HF Energy | -23.536062 |
| Nuclear repulsion energy | 35.485911 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3400 |
3284 |
12.71 |
|
|
|
| 2 |
A' |
3238 |
3127 |
23.84 |
|
|
|
| 3 |
A' |
3142 |
3034 |
7.97 |
|
|
|
| 4 |
A' |
3124 |
3017 |
25.20 |
|
|
|
| 5 |
A' |
1494 |
1442 |
5.15 |
|
|
|
| 6 |
A' |
1428 |
1379 |
7.84 |
|
|
|
| 7 |
A' |
1351 |
1305 |
9.19 |
|
|
|
| 8 |
A' |
1232 |
1190 |
30.53 |
|
|
|
| 9 |
A' |
1136 |
1097 |
24.41 |
|
|
|
| 10 |
A' |
953 |
921 |
1.49 |
|
|
|
| 11 |
A' |
472 |
456 |
8.52 |
|
|
|
| 12 |
A" |
1063 |
1026 |
64.39 |
|
|
|
| 13 |
A" |
789 |
762 |
89.89 |
|
|
|
| 14 |
A" |
666 |
643 |
0.79 |
|
|
|
| 15 |
A" |
474 |
458 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11981.7 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 11570.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.459 |
0.000 |
| C2 |
1.148 |
-0.400 |
0.000 |
| N3 |
-1.279 |
0.077 |
0.000 |
| H4 |
0.175 |
1.546 |
0.000 |
| H5 |
2.163 |
0.013 |
0.000 |
| H6 |
1.034 |
-1.491 |
0.000 |
| H7 |
-1.305 |
-0.958 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4338 | 1.3352 | 1.1008 | 2.2083 | 2.2072 | 1.9265 |
C2 | 1.4338 | | 2.4738 | 2.1754 | 1.0957 | 1.0970 | 2.5161 | N3 | 1.3352 | 2.4738 | | 2.0672 | 3.4428 | 2.7945 | 1.0349 | H4 | 1.1008 | 2.1754 | 2.0672 | | 2.5099 | 3.1559 | 2.9086 | H5 | 2.2083 | 1.0957 | 3.4428 | 2.5099 | | 1.8808 | 3.6017 | H6 | 2.2072 | 1.0970 | 2.7945 | 3.1559 | 1.8808 | | 2.3994 | H7 | 1.9265 | 2.5161 | 1.0349 | 2.9086 | 3.6017 | 2.3994 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.042 |
|
C1 |
C2 |
H6 |
120.831 |
| C1 |
N3 |
H7 |
108.078 |
|
C2 |
C1 |
N3 |
126.565 |
| C2 |
C1 |
H4 |
117.644 |
|
N3 |
C1 |
H4 |
115.790 |
| H5 |
C2 |
H6 |
118.127 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.336 |
|
|
|
| 2 |
C |
-0.423 |
|
|
|
| 3 |
N |
-0.103 |
|
|
|
| 4 |
H |
0.223 |
|
|
|
| 5 |
H |
0.210 |
|
|
|
| 6 |
H |
0.207 |
|
|
|
| 7 |
H |
0.223 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.307 |
-1.102 |
0.000 |
2.556 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.797 |
2.393 |
0.000 |
| y |
2.393 |
-15.819 |
0.000 |
| z |
0.000 |
0.000 |
-20.315 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.730 |
2.393 |
0.000 |
| y |
2.393 |
4.737 |
0.000 |
| z |
0.000 |
0.000 |
-2.006 |
|
| Polar |
|---|
| 3z2-r2 | -4.013 |
|---|
| x2-y2 | -4.978 |
|---|
| xy | 2.393 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.250 |
-0.367 |
0.000 |
| y |
-0.367 |
4.136 |
0.000 |
| z |
0.000 |
0.000 |
2.196 |
<r2> (average value of r
2) Å
2
| <r2> |
39.833 |
| (<r2>)1/2 |
6.311 |
