Jump to
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -16.653027 |
| Energy at 298.15K | -16.654296 |
| HF Energy | -16.653027 |
| Nuclear repulsion energy | 15.425610 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3293 |
3180 |
2.38 |
|
|
|
| 2 |
A' |
2976 |
2874 |
59.60 |
|
|
|
| 3 |
A' |
1813 |
1751 |
34.64 |
|
|
|
| 4 |
A' |
1010 |
976 |
130.21 |
|
|
|
| 5 |
A' |
849 |
820 |
101.61 |
|
|
|
| 6 |
A" |
866 |
836 |
1.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5403.0 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 5217.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.114 |
0.648 |
0.000 |
| N2 |
0.114 |
-0.602 |
0.000 |
| H3 |
-0.685 |
1.421 |
0.000 |
| H4 |
-0.799 |
-1.096 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2491 | 1.1124 | 1.9681 |
N2 | 1.2491 | | 2.1750 | 1.0384 | H3 | 1.1124 | 2.1750 | | 2.5196 | H4 | 1.9681 | 1.0384 | 2.5196 | |
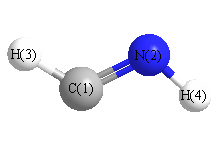 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
118.426 |
|
H3 |
C1 |
N2 |
134.062 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.420 |
|
|
|
| 2 |
N |
-0.052 |
|
|
|
| 3 |
H |
0.206 |
|
|
|
| 4 |
H |
0.266 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.535 |
0.546 |
0.000 |
2.593 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-11.962 |
0.491 |
0.000 |
| y |
0.491 |
-11.097 |
0.000 |
| z |
0.000 |
0.000 |
-12.695 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.067 |
0.491 |
0.000 |
| y |
0.491 |
1.231 |
0.000 |
| z |
0.000 |
0.000 |
-1.165 |
|
| Polar |
|---|
| 3z2-r2 | -2.330 |
|---|
| x2-y2 | -0.865 |
|---|
| xy | 0.491 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.975 |
0.223 |
0.000 |
| y |
0.223 |
3.922 |
0.000 |
| z |
0.000 |
0.000 |
1.476 |
<r2> (average value of r
2) Å
2
| <r2> |
15.377 |
| (<r2>)1/2 |
3.921 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -16.663350 |
| Energy at 298.15K | -16.664683 |
| HF Energy | -16.663350 |
| Nuclear repulsion energy | 15.406022 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3430 |
3312 |
2.64 |
|
|
|
| 2 |
A' |
3035 |
2931 |
34.50 |
|
|
|
| 3 |
A' |
1766 |
1705 |
36.27 |
|
|
|
| 4 |
A' |
1210 |
1168 |
13.12 |
|
|
|
| 5 |
A' |
906 |
875 |
216.29 |
|
|
|
| 6 |
A" |
959 |
926 |
131.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5652.7 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 5458.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.001 |
0.660 |
0.000 |
| N2 |
0.001 |
-0.599 |
0.000 |
| H3 |
0.920 |
1.282 |
0.000 |
| H4 |
-0.927 |
-1.052 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2591 | 1.1096 | 1.9473 |
N2 | 1.2591 | | 2.0933 | 1.0320 | H3 | 1.1096 | 2.0933 | | 2.9759 | H4 | 1.9473 | 1.0320 | 2.9759 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
116.057 |
|
H3 |
C1 |
N2 |
124.068 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.427 |
|
|
|
| 2 |
N |
-0.067 |
|
|
|
| 3 |
H |
0.207 |
|
|
|
| 4 |
H |
0.287 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.411 |
0.252 |
0.000 |
0.482 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-11.052 |
3.438 |
0.000 |
| y |
3.438 |
-11.514 |
0.000 |
| z |
0.000 |
0.000 |
-12.756 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.083 |
3.438 |
0.000 |
| y |
3.438 |
0.390 |
0.000 |
| z |
0.000 |
0.000 |
-1.473 |
|
| Polar |
|---|
| 3z2-r2 | -2.946 |
|---|
| x2-y2 | 0.462 |
|---|
| xy | 3.438 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.163 |
0.309 |
0.000 |
| y |
0.309 |
3.863 |
0.000 |
| z |
0.000 |
0.000 |
1.549 |
<r2> (average value of r
2) Å
2
| <r2> |
15.345 |
| (<r2>)1/2 |
3.917 |
