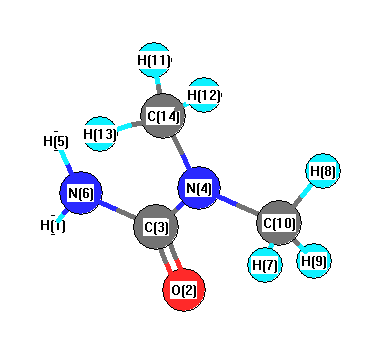
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3663 |
3537 |
18.80 |
|
|
|
| 2 |
A |
3551 |
3429 |
15.50 |
|
|
|
| 3 |
A |
3178 |
3069 |
7.87 |
|
|
|
| 4 |
A |
3152 |
3044 |
39.36 |
|
|
|
| 5 |
A |
3092 |
2986 |
94.35 |
|
|
|
| 6 |
A |
3090 |
2984 |
8.24 |
|
|
|
| 7 |
A |
3018 |
2915 |
97.80 |
|
|
|
| 8 |
A |
3012 |
2908 |
50.36 |
|
|
|
| 9 |
A |
1760 |
1700 |
390.81 |
|
|
|
| 10 |
A |
1640 |
1584 |
146.47 |
|
|
|
| 11 |
A |
1525 |
1473 |
48.71 |
|
|
|
| 12 |
A |
1501 |
1449 |
13.29 |
|
|
|
| 13 |
A |
1497 |
1445 |
7.53 |
|
|
|
| 14 |
A |
1485 |
1434 |
28.12 |
|
|
|
| 15 |
A |
1476 |
1425 |
17.50 |
|
|
|
| 16 |
A |
1441 |
1392 |
4.81 |
|
|
|
| 17 |
A |
1414 |
1366 |
208.28 |
|
|
|
| 18 |
A |
1272 |
1229 |
51.01 |
|
|
|
| 19 |
A |
1251 |
1208 |
14.09 |
|
|
|
| 20 |
A |
1160 |
1120 |
8.34 |
|
|
|
| 21 |
A |
1113 |
1075 |
4.54 |
|
|
|
| 22 |
A |
1095 |
1058 |
79.73 |
|
|
|
| 23 |
A |
1070 |
1033 |
13.90 |
|
|
|
| 24 |
A |
1012 |
977 |
23.08 |
|
|
|
| 25 |
A |
782 |
755 |
87.43 |
|
|
|
| 26 |
A |
761 |
735 |
2.65 |
|
|
|
| 27 |
A |
629 |
607 |
169.54 |
|
|
|
| 28 |
A |
581 |
561 |
23.20 |
|
|
|
| 29 |
A |
493 |
476 |
4.87 |
|
|
|
| 30 |
A |
426 |
412 |
19.63 |
|
|
|
| 31 |
A |
382 |
369 |
21.98 |
|
|
|
| 32 |
A |
314 |
304 |
7.88 |
|
|
|
| 33 |
A |
230 |
223 |
20.77 |
|
|
|
| 34 |
A |
138 |
133 |
1.64 |
|
|
|
| 35 |
A |
102 |
99 |
3.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26152.8 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 25255.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.324 |
|
|
|
| 2 |
O |
-0.169 |
|
|
|
| 3 |
C |
-0.315 |
|
|
|
| 4 |
N |
0.140 |
|
|
|
| 5 |
H |
0.292 |
|
|
|
| 6 |
N |
-0.410 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
| 9 |
H |
0.242 |
|
|
|
| 10 |
C |
-0.495 |
|
|
|
| 11 |
H |
0.180 |
|
|
|
| 12 |
H |
0.181 |
|
|
|
| 13 |
H |
0.164 |
|
|
|
| 14 |
C |
-0.463 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.585 |
3.316 |
-1.050 |
3.822 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.099 |
-4.559 |
2.647 |
| y |
-4.559 |
-37.973 |
-1.183 |
| z |
2.647 |
-1.183 |
-37.387 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.581 |
-4.559 |
2.647 |
| y |
-4.559 |
-3.230 |
-1.183 |
| z |
2.647 |
-1.183 |
-2.351 |
|
| Polar |
|---|
| 3z2-r2 | -4.701 |
|---|
| x2-y2 | 5.874 |
|---|
| xy | -4.559 |
|---|
| xz | 2.647 |
|---|
| yz | -1.183 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.554 |
0.136 |
-0.041 |
| y |
0.136 |
7.877 |
-0.160 |
| z |
-0.041 |
-0.160 |
5.218 |
<r2> (average value of r
2) Å
2
| <r2> |
138.925 |
| (<r2>)1/2 |
11.787 |