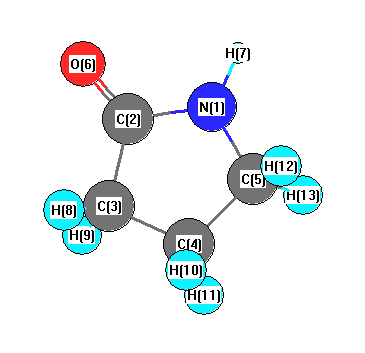
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3614 |
3490 |
25.15 |
|
|
|
| 2 |
A |
3140 |
3032 |
44.66 |
|
|
|
| 3 |
A |
3123 |
3016 |
26.83 |
|
|
|
| 4 |
A |
3089 |
2983 |
54.34 |
|
|
|
| 5 |
A |
3073 |
2968 |
21.41 |
|
|
|
| 6 |
A |
3067 |
2961 |
22.90 |
|
|
|
| 7 |
A |
3015 |
2912 |
83.10 |
|
|
|
| 8 |
A |
1794 |
1733 |
477.47 |
|
|
|
| 9 |
A |
1532 |
1479 |
3.11 |
|
|
|
| 10 |
A |
1497 |
1446 |
7.72 |
|
|
|
| 11 |
A |
1470 |
1420 |
4.35 |
|
|
|
| 12 |
A |
1432 |
1383 |
38.25 |
|
|
|
| 13 |
A |
1357 |
1310 |
8.36 |
|
|
|
| 14 |
A |
1333 |
1287 |
19.43 |
|
|
|
| 15 |
A |
1302 |
1257 |
41.54 |
|
|
|
| 16 |
A |
1254 |
1211 |
66.93 |
|
|
|
| 17 |
A |
1233 |
1190 |
14.90 |
|
|
|
| 18 |
A |
1206 |
1165 |
0.96 |
|
|
|
| 19 |
A |
1173 |
1133 |
3.26 |
|
|
|
| 20 |
A |
1075 |
1039 |
1.29 |
|
|
|
| 21 |
A |
1068 |
1031 |
13.62 |
|
|
|
| 22 |
A |
994 |
960 |
13.98 |
|
|
|
| 23 |
A |
914 |
882 |
0.38 |
|
|
|
| 24 |
A |
892 |
862 |
1.60 |
|
|
|
| 25 |
A |
877 |
847 |
2.97 |
|
|
|
| 26 |
A |
803 |
776 |
6.60 |
|
|
|
| 27 |
A |
677 |
654 |
8.56 |
|
|
|
| 28 |
A |
618 |
596 |
19.38 |
|
|
|
| 29 |
A |
550 |
531 |
39.88 |
|
|
|
| 30 |
A |
484 |
467 |
79.13 |
|
|
|
| 31 |
A |
455 |
440 |
12.50 |
|
|
|
| 32 |
A |
190 |
184 |
6.28 |
|
|
|
| 33 |
A |
135 |
130 |
1.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24216.9 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 23386.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.090 |
|
|
|
| 2 |
C |
-0.386 |
|
|
|
| 3 |
C |
-0.152 |
|
|
|
| 4 |
C |
-0.264 |
|
|
|
| 5 |
C |
-0.313 |
|
|
|
| 6 |
O |
-0.116 |
|
|
|
| 7 |
H |
0.325 |
|
|
|
| 8 |
H |
0.153 |
|
|
|
| 9 |
H |
0.189 |
|
|
|
| 10 |
H |
0.179 |
|
|
|
| 11 |
H |
0.144 |
|
|
|
| 12 |
H |
0.146 |
|
|
|
| 13 |
H |
0.184 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.238 |
-0.572 |
0.471 |
4.302 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.924 |
-0.226 |
0.115 |
| y |
-0.226 |
-30.954 |
-0.372 |
| z |
0.115 |
-0.372 |
-35.075 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.910 |
-0.226 |
0.115 |
| y |
-0.226 |
8.545 |
-0.372 |
| z |
0.115 |
-0.372 |
2.364 |
|
| Polar |
|---|
| 3z2-r2 | 4.729 |
|---|
| x2-y2 | -12.970 |
|---|
| xy | -0.226 |
|---|
| xz | 0.115 |
|---|
| yz | -0.372 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.153 |
-0.177 |
0.369 |
| y |
-0.177 |
7.379 |
0.039 |
| z |
0.369 |
0.039 |
5.579 |
<r2> (average value of r
2) Å
2
| <r2> |
120.392 |
| (<r2>)1/2 |
10.972 |