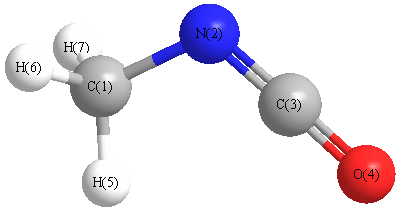
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3144 |
3036 |
29.54 |
78.10 |
0.65 |
0.79 |
| 2 |
A' |
3047 |
2942 |
62.91 |
169.30 |
0.01 |
0.02 |
| 3 |
A' |
2347 |
2267 |
1022.19 |
0.52 |
0.23 |
0.37 |
| 4 |
A' |
1495 |
1444 |
0.33 |
7.50 |
0.38 |
0.55 |
| 5 |
A' |
1468 |
1417 |
15.17 |
17.76 |
0.60 |
0.75 |
| 6 |
A' |
1449 |
1399 |
38.13 |
19.67 |
0.48 |
0.65 |
| 7 |
A' |
1137 |
1098 |
16.62 |
4.51 |
0.71 |
0.83 |
| 8 |
A' |
869 |
840 |
36.19 |
12.27 |
0.15 |
0.26 |
| 9 |
A' |
617 |
595 |
26.58 |
0.61 |
0.75 |
0.85 |
| 10 |
A' |
168 |
163 |
17.97 |
1.75 |
0.68 |
0.81 |
| 11 |
A" |
3113 |
3007 |
39.34 |
69.35 |
0.75 |
0.86 |
| 12 |
A" |
1500 |
1448 |
6.55 |
19.51 |
0.75 |
0.86 |
| 13 |
A" |
1112 |
1074 |
0.01 |
3.75 |
0.75 |
0.86 |
| 14 |
A" |
560 |
541 |
21.13 |
0.95 |
0.75 |
0.86 |
| 15 |
A" |
9 |
9 |
2.39 |
1.18 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11017.3 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 10639.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.448 |
|
|
|
| 2 |
N |
0.296 |
|
|
|
| 3 |
C |
-0.378 |
|
|
|
| 4 |
O |
-0.059 |
|
|
|
| 5 |
H |
0.212 |
|
|
|
| 6 |
H |
0.188 |
|
|
|
| 7 |
H |
0.188 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.755 |
1.479 |
0.000 |
3.128 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.061 |
-0.427 |
0.000 |
| y |
-0.427 |
-24.678 |
0.000 |
| z |
0.000 |
0.000 |
-22.372 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.464 |
-0.427 |
0.000 |
| y |
-0.427 |
-2.461 |
0.000 |
| z |
0.000 |
0.000 |
0.997 |
|
| Polar |
|---|
| 3z2-r2 | 1.995 |
|---|
| x2-y2 | 2.617 |
|---|
| xy | -0.427 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.562 |
1.949 |
0.000 |
| y |
1.949 |
5.415 |
0.000 |
| z |
0.000 |
0.000 |
2.874 |
<r2> (average value of r
2) Å
2
| <r2> |
69.565 |
| (<r2>)1/2 |
8.341 |