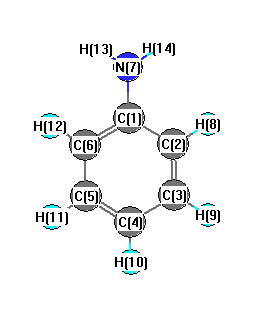
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3539 |
3417 |
7.41 |
|
|
|
| 2 |
A' |
3191 |
3082 |
17.61 |
|
|
|
| 3 |
A' |
3162 |
3053 |
10.04 |
|
|
|
| 4 |
A' |
3138 |
3030 |
13.55 |
|
|
|
| 5 |
A' |
1677 |
1620 |
129.19 |
|
|
|
| 6 |
A' |
1653 |
1596 |
24.07 |
|
|
|
| 7 |
A' |
1519 |
1467 |
65.71 |
|
|
|
| 8 |
A' |
1276 |
1233 |
60.15 |
|
|
|
| 9 |
A' |
1181 |
1141 |
7.73 |
|
|
|
| 10 |
A' |
1041 |
1006 |
5.96 |
|
|
|
| 11 |
A' |
993 |
959 |
0.18 |
|
|
|
| 12 |
A' |
981 |
947 |
1.05 |
|
|
|
| 13 |
A' |
900 |
869 |
11.10 |
|
|
|
| 14 |
A' |
812 |
785 |
7.31 |
|
|
|
| 15 |
A' |
774 |
748 |
143.84 |
|
|
|
| 16 |
A' |
685 |
662 |
13.90 |
|
|
|
| 17 |
A' |
658 |
635 |
305.90 |
|
|
|
| 18 |
A' |
514 |
496 |
1.18 |
|
|
|
| 19 |
A' |
502 |
485 |
47.78 |
|
|
|
| 20 |
A' |
220 |
213 |
5.86 |
|
|
|
| 21 |
A" |
3641 |
3516 |
6.59 |
|
|
|
| 22 |
A" |
3171 |
3062 |
50.57 |
|
|
|
| 23 |
A" |
3144 |
3036 |
1.14 |
|
|
|
| 24 |
A" |
1640 |
1584 |
5.11 |
|
|
|
| 25 |
A" |
1489 |
1437 |
1.76 |
|
|
|
| 26 |
A" |
1390 |
1342 |
8.64 |
|
|
|
| 27 |
A" |
1339 |
1293 |
0.06 |
|
|
|
| 28 |
A" |
1155 |
1116 |
1.16 |
|
|
|
| 29 |
A" |
1126 |
1087 |
3.66 |
|
|
|
| 30 |
A" |
1065 |
1028 |
6.44 |
|
|
|
| 31 |
A" |
980 |
947 |
0.03 |
|
|
|
| 32 |
A" |
847 |
818 |
0.00 |
|
|
|
| 33 |
A" |
607 |
586 |
0.32 |
|
|
|
| 34 |
A" |
406 |
393 |
0.46 |
|
|
|
| 35 |
A" |
370 |
358 |
0.14 |
|
|
|
| 36 |
A" |
253 |
244 |
25.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25519.4 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 24644.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.087 |
|
|
0.000 |
| 2 |
C |
-0.300 |
|
|
0.000 |
| 3 |
C |
-0.340 |
|
|
0.000 |
| 4 |
C |
-0.293 |
|
|
0.000 |
| 5 |
C |
-0.340 |
|
|
0.000 |
| 6 |
C |
-0.300 |
|
|
0.000 |
| 7 |
N |
-0.364 |
|
|
0.000 |
| 8 |
H |
0.289 |
|
|
0.000 |
| 9 |
H |
0.287 |
|
|
0.000 |
| 10 |
H |
0.293 |
|
|
0.000 |
| 11 |
H |
0.287 |
|
|
0.000 |
| 12 |
H |
0.289 |
|
|
0.000 |
| 13 |
H |
0.289 |
|
|
0.000 |
| 14 |
H |
0.289 |
|
|
0.000 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.215 |
1.157 |
0.000 |
1.678 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.478 |
-3.827 |
0.000 |
| y |
-3.827 |
-34.061 |
0.000 |
| z |
0.000 |
0.000 |
-35.371 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.762 |
-3.827 |
0.000 |
| y |
-3.827 |
6.863 |
0.000 |
| z |
0.000 |
0.000 |
4.898 |
|
| Polar |
|---|
| 3z2-r2 | 9.797 |
|---|
| x2-y2 | -12.417 |
|---|
| xy | -3.827 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.824 |
-0.046 |
0.000 |
| y |
-0.046 |
14.091 |
0.000 |
| z |
0.000 |
0.000 |
11.711 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |