Jump to
S1C2
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -25.407078 |
| Energy at 298.15K | -25.415155 |
| Nuclear repulsion energy | 49.430361 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3458 |
3339 |
3.62 |
|
|
|
| 2 |
A' |
3090 |
2984 |
80.27 |
|
|
|
| 3 |
A' |
3050 |
2945 |
40.38 |
|
|
|
| 4 |
A' |
3024 |
2921 |
31.20 |
|
|
|
| 5 |
A' |
1677 |
1620 |
18.26 |
|
|
|
| 6 |
A' |
1501 |
1450 |
3.12 |
|
|
|
| 7 |
A' |
1482 |
1431 |
0.02 |
|
|
|
| 8 |
A' |
1407 |
1359 |
9.14 |
|
|
|
| 9 |
A' |
1364 |
1317 |
16.77 |
|
|
|
| 10 |
A' |
1139 |
1100 |
12.44 |
|
|
|
| 11 |
A' |
1050 |
1014 |
31.92 |
|
|
|
| 12 |
A' |
897 |
866 |
125.36 |
|
|
|
| 13 |
A' |
865 |
835 |
34.13 |
|
|
|
| 14 |
A' |
387 |
374 |
6.48 |
|
|
|
| 15 |
A" |
3543 |
3422 |
1.45 |
|
|
|
| 16 |
A" |
3104 |
2997 |
111.48 |
|
|
|
| 17 |
A" |
3074 |
2968 |
0.11 |
|
|
|
| 18 |
A" |
1494 |
1442 |
8.26 |
|
|
|
| 19 |
A" |
1379 |
1332 |
0.01 |
|
|
|
| 20 |
A" |
1252 |
1209 |
0.02 |
|
|
|
| 21 |
A" |
991 |
957 |
2.31 |
|
|
|
| 22 |
A" |
761 |
735 |
2.35 |
|
|
|
| 23 |
A" |
280 |
270 |
36.93 |
|
|
|
| 24 |
A" |
251 |
243 |
27.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20259.9 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 19565.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.339 |
-0.046 |
0.000 |
| C2 |
0.000 |
0.601 |
0.000 |
| C3 |
1.211 |
-0.373 |
0.000 |
| H4 |
2.171 |
0.177 |
0.000 |
| H5 |
1.194 |
-1.024 |
0.894 |
| H6 |
1.194 |
-1.024 |
-0.894 |
| H7 |
0.052 |
1.261 |
-0.885 |
| H8 |
0.052 |
1.261 |
0.885 |
| H9 |
-1.406 |
-0.665 |
0.816 |
| H10 |
-1.406 |
-0.665 |
-0.816 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4868 | 2.5699 | 3.5168 | 2.8585 | 2.8585 | 2.1034 | 2.1034 | 1.0263 | 1.0263 |
C2 | 1.4868 | | 1.5536 | 2.2121 | 2.2061 | 2.2061 | 1.1051 | 1.1051 | 2.0604 | 2.0604 | C3 | 2.5699 | 1.5536 | | 1.1071 | 1.1066 | 1.1066 | 2.1898 | 2.1898 | 2.7564 | 2.7564 | H4 | 3.5168 | 2.2121 | 1.1071 | | 1.7884 | 1.7884 | 2.5395 | 2.5395 | 3.7645 | 3.7645 | H5 | 2.8585 | 2.2061 | 1.1066 | 1.7884 | | 1.7887 | 3.1133 | 2.5549 | 2.6260 | 3.1330 | H6 | 2.8585 | 2.2061 | 1.1066 | 1.7884 | 1.7887 | | 2.5549 | 3.1133 | 3.1330 | 2.6260 | H7 | 2.1034 | 1.1051 | 2.1898 | 2.5395 | 3.1133 | 2.5549 | | 1.7696 | 2.9541 | 2.4163 | H8 | 2.1034 | 1.1051 | 2.1898 | 2.5395 | 2.5549 | 3.1133 | 1.7696 | | 2.4163 | 2.9541 | H9 | 1.0263 | 2.0604 | 2.7564 | 3.7645 | 2.6260 | 3.1330 | 2.9541 | 2.4163 | | 1.6324 | H10 | 1.0263 | 2.0604 | 2.7564 | 3.7645 | 3.1330 | 2.6260 | 2.4163 | 2.9541 | 1.6324 | |
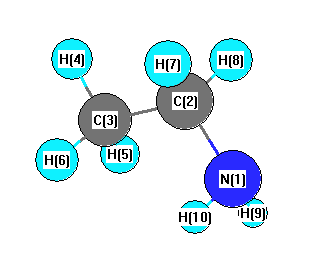 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.384 |
|
N1 |
C2 |
H7 |
107.585 |
| N1 |
C2 |
H8 |
107.585 |
|
C2 |
N1 |
H9 |
108.757 |
| C2 |
N1 |
H10 |
108.757 |
|
C2 |
C3 |
H4 |
111.383 |
| C2 |
C3 |
H5 |
110.942 |
|
C2 |
C3 |
H6 |
110.942 |
| C3 |
C2 |
H7 |
109.750 |
|
C3 |
C2 |
H8 |
109.750 |
| H4 |
C3 |
H5 |
107.786 |
|
H4 |
C3 |
H6 |
107.786 |
| H5 |
C3 |
H6 |
107.847 |
|
H7 |
C2 |
H8 |
106.384 |
| H9 |
N1 |
H10 |
105.357 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.491 |
|
|
|
| 2 |
C |
-0.249 |
|
|
|
| 3 |
C |
-0.457 |
|
|
|
| 4 |
H |
0.148 |
|
|
|
| 5 |
H |
0.138 |
|
|
|
| 6 |
H |
0.138 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.240 |
|
|
|
| 10 |
H |
0.240 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.202 |
-1.130 |
0.000 |
1.650 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.084 |
0.044 |
-0.105 |
| y |
0.044 |
4.373 |
-0.110 |
| z |
-0.105 |
-0.110 |
3.850 |
<r2> (average value of r
2) Å
2
| <r2> |
52.148 |
| (<r2>)1/2 |
7.221 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-31G*
| | hartrees |
|---|
| Energy at 0K | -25.407666 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-31G*
Geometric Data calculated at B3LYP/CEP-31G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
