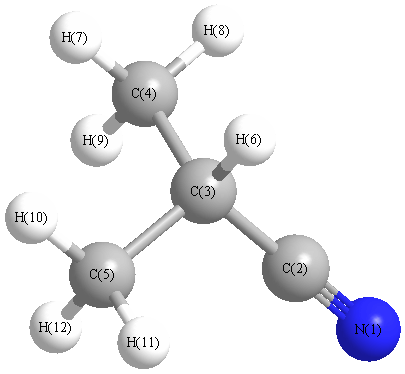
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3139 |
3031 |
53.56 |
|
|
|
| 2 |
A' |
3131 |
3023 |
58.32 |
|
|
|
| 3 |
A' |
3066 |
2961 |
8.60 |
|
|
|
| 4 |
A' |
3055 |
2950 |
24.07 |
|
|
|
| 5 |
A' |
2334 |
2254 |
14.83 |
|
|
|
| 6 |
A' |
1510 |
1458 |
11.46 |
|
|
|
| 7 |
A' |
1500 |
1449 |
9.53 |
|
|
|
| 8 |
A' |
1422 |
1374 |
3.32 |
|
|
|
| 9 |
A' |
1339 |
1293 |
8.32 |
|
|
|
| 10 |
A' |
1182 |
1142 |
4.57 |
|
|
|
| 11 |
A' |
1115 |
1077 |
8.48 |
|
|
|
| 12 |
A' |
920 |
889 |
0.68 |
|
|
|
| 13 |
A' |
762 |
736 |
0.79 |
|
|
|
| 14 |
A' |
535 |
517 |
1.65 |
|
|
|
| 15 |
A' |
343 |
331 |
0.15 |
|
|
|
| 16 |
A' |
277 |
268 |
0.51 |
|
|
|
| 17 |
A' |
213 |
206 |
1.56 |
|
|
|
| 18 |
A" |
3135 |
3027 |
17.55 |
|
|
|
| 19 |
A" |
3126 |
3019 |
5.00 |
|
|
|
| 20 |
A" |
3053 |
2948 |
25.23 |
|
|
|
| 21 |
A" |
1490 |
1439 |
3.28 |
|
|
|
| 22 |
A" |
1488 |
1437 |
1.20 |
|
|
|
| 23 |
A" |
1403 |
1354 |
2.54 |
|
|
|
| 24 |
A" |
1312 |
1267 |
0.44 |
|
|
|
| 25 |
A" |
1123 |
1085 |
2.62 |
|
|
|
| 26 |
A" |
962 |
929 |
0.30 |
|
|
|
| 27 |
A" |
930 |
898 |
3.30 |
|
|
|
| 28 |
A" |
553 |
534 |
0.70 |
|
|
|
| 29 |
A" |
221 |
214 |
0.00 |
|
|
|
| 30 |
A" |
183 |
177 |
3.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22410.7 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 21642.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
0.372 |
|
|
|
| 2 |
C |
-0.650 |
|
|
|
| 3 |
C |
0.093 |
|
|
|
| 4 |
C |
-0.511 |
|
|
|
| 5 |
C |
-0.511 |
|
|
|
| 6 |
H |
0.161 |
|
|
|
| 7 |
H |
0.172 |
|
|
|
| 8 |
H |
0.189 |
|
|
|
| 9 |
H |
0.163 |
|
|
|
| 10 |
H |
0.172 |
|
|
|
| 11 |
H |
0.189 |
|
|
|
| 12 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.199 |
4.002 |
0.000 |
4.178 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.103 |
1.715 |
0.000 |
| y |
1.715 |
-38.940 |
0.000 |
| z |
0.000 |
0.000 |
-29.828 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.281 |
1.715 |
0.000 |
| y |
1.715 |
-8.975 |
0.000 |
| z |
0.000 |
0.000 |
4.694 |
|
| Polar |
|---|
| 3z2-r2 | 9.387 |
|---|
| x2-y2 | 8.837 |
|---|
| xy | 1.715 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.623 |
-0.617 |
0.000 |
| y |
-0.617 |
8.290 |
0.000 |
| z |
0.000 |
0.000 |
6.394 |
<r2> (average value of r
2) Å
2
| <r2> |
107.372 |
| (<r2>)1/2 |
10.362 |