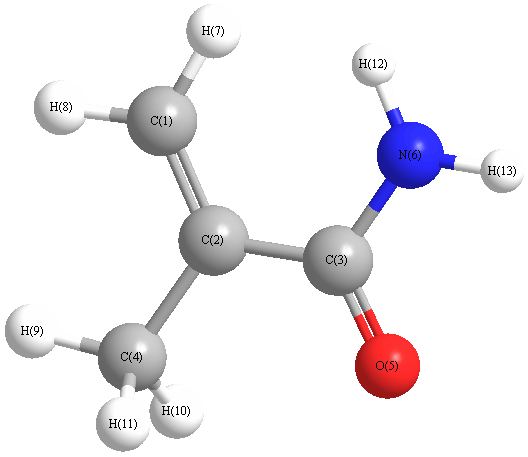
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3699 |
3572 |
24.15 |
|
|
|
| 2 |
A |
3572 |
3449 |
28.63 |
|
|
|
| 3 |
A |
3221 |
3111 |
16.28 |
|
|
|
| 4 |
A |
3149 |
3041 |
10.54 |
|
|
|
| 5 |
A |
3137 |
3030 |
19.97 |
|
|
|
| 6 |
A |
3113 |
3006 |
24.04 |
|
|
|
| 7 |
A |
3050 |
2946 |
22.06 |
|
|
|
| 8 |
A |
1756 |
1696 |
270.10 |
|
|
|
| 9 |
A |
1697 |
1639 |
65.93 |
|
|
|
| 10 |
A |
1619 |
1563 |
90.47 |
|
|
|
| 11 |
A |
1490 |
1439 |
17.64 |
|
|
|
| 12 |
A |
1471 |
1421 |
10.50 |
|
|
|
| 13 |
A |
1441 |
1391 |
11.53 |
|
|
|
| 14 |
A |
1415 |
1366 |
9.90 |
|
|
|
| 15 |
A |
1370 |
1323 |
83.72 |
|
|
|
| 16 |
A |
1211 |
1170 |
62.59 |
|
|
|
| 17 |
A |
1082 |
1045 |
7.27 |
|
|
|
| 18 |
A |
1055 |
1019 |
0.87 |
|
|
|
| 19 |
A |
1006 |
971 |
7.33 |
|
|
|
| 20 |
A |
949 |
917 |
61.17 |
|
|
|
| 21 |
A |
940 |
907 |
13.06 |
|
|
|
| 22 |
A |
800 |
773 |
11.61 |
|
|
|
| 23 |
A |
739 |
714 |
5.00 |
|
|
|
| 24 |
A |
657 |
634 |
12.88 |
|
|
|
| 25 |
A |
559 |
540 |
31.34 |
|
|
|
| 26 |
A |
540 |
522 |
32.00 |
|
|
|
| 27 |
A |
488 |
471 |
8.93 |
|
|
|
| 28 |
A |
403 |
390 |
174.62 |
|
|
|
| 29 |
A |
357 |
345 |
13.48 |
|
|
|
| 30 |
A |
307 |
296 |
4.49 |
|
|
|
| 31 |
A |
254 |
246 |
9.69 |
|
|
|
| 32 |
A |
164 |
158 |
0.20 |
|
|
|
| 33 |
A |
65 |
63 |
4.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23387.0 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 22584.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.367 |
|
|
|
| 2 |
H |
0.326 |
|
|
|
| 3 |
H |
0.317 |
|
|
|
| 4 |
C |
-0.454 |
|
|
|
| 5 |
H |
0.146 |
|
|
|
| 6 |
H |
0.181 |
|
|
|
| 7 |
H |
0.172 |
|
|
|
| 8 |
C |
-0.694 |
|
|
|
| 9 |
H |
0.236 |
|
|
|
| 10 |
H |
0.240 |
|
|
|
| 11 |
C |
-0.337 |
|
|
|
| 12 |
O |
-0.124 |
|
|
|
| 13 |
C |
0.357 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.365 |
3.061 |
1.661 |
3.502 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.473 |
-4.018 |
-2.813 |
| y |
-4.018 |
-37.713 |
-0.016 |
| z |
-2.813 |
-0.016 |
-37.055 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.910 |
-4.018 |
-2.813 |
| y |
-4.018 |
-3.949 |
-0.016 |
| z |
-2.813 |
-0.016 |
-2.962 |
|
| Polar |
|---|
| 3z2-r2 | -5.923 |
|---|
| x2-y2 | 7.239 |
|---|
| xy | -4.018 |
|---|
| xz | -2.813 |
|---|
| yz | -0.016 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.445 |
0.472 |
-0.107 |
| y |
0.472 |
10.210 |
-0.222 |
| z |
-0.107 |
-0.222 |
4.824 |
<r2> (average value of r
2) Å
2
| <r2> |
137.582 |
| (<r2>)1/2 |
11.730 |