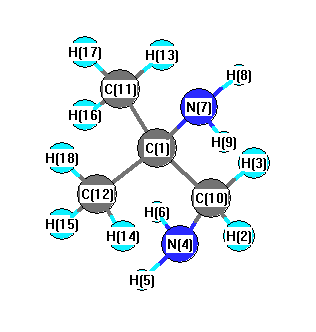
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3478 |
3359 |
1.35 |
|
|
|
| 2 |
A |
3444 |
3326 |
3.80 |
|
|
|
| 3 |
A |
3126 |
3018 |
66.02 |
|
|
|
| 4 |
A |
3100 |
2994 |
85.74 |
|
|
|
| 5 |
A |
3036 |
2932 |
1.36 |
|
|
|
| 6 |
A |
3017 |
2914 |
53.69 |
|
|
|
| 7 |
A |
1674 |
1616 |
9.87 |
|
|
|
| 8 |
A |
1673 |
1616 |
32.41 |
|
|
|
| 9 |
A |
1509 |
1458 |
11.62 |
|
|
|
| 10 |
A |
1503 |
1452 |
6.22 |
|
|
|
| 11 |
A |
1472 |
1422 |
0.49 |
|
|
|
| 12 |
A |
1417 |
1368 |
11.91 |
|
|
|
| 13 |
A |
1365 |
1318 |
7.56 |
|
|
|
| 14 |
A |
1255 |
1212 |
11.80 |
|
|
|
| 15 |
A |
1206 |
1165 |
35.89 |
|
|
|
| 16 |
A |
1085 |
1048 |
13.46 |
|
|
|
| 17 |
A |
1011 |
976 |
0.39 |
|
|
|
| 18 |
A |
924 |
893 |
173.94 |
|
|
|
| 19 |
A |
901 |
870 |
77.36 |
|
|
|
| 20 |
A |
863 |
833 |
146.04 |
|
|
|
| 21 |
A |
719 |
694 |
9.64 |
|
|
|
| 22 |
A |
493 |
477 |
12.00 |
|
|
|
| 23 |
A |
409 |
395 |
10.30 |
|
|
|
| 24 |
A |
350 |
338 |
0.04 |
|
|
|
| 25 |
A |
256 |
247 |
1.44 |
|
|
|
| 26 |
A |
248 |
239 |
5.69 |
|
|
|
| 27 |
A |
3569 |
3446 |
0.25 |
|
|
|
| 28 |
A |
3532 |
3411 |
1.72 |
|
|
|
| 29 |
A |
3120 |
3013 |
2.04 |
|
|
|
| 30 |
A |
3096 |
2989 |
23.54 |
|
|
|
| 31 |
A |
3060 |
2955 |
51.05 |
|
|
|
| 32 |
A |
3030 |
2926 |
55.49 |
|
|
|
| 33 |
A |
1495 |
1443 |
0.82 |
|
|
|
| 34 |
A |
1483 |
1432 |
0.11 |
|
|
|
| 35 |
A |
1413 |
1364 |
9.79 |
|
|
|
| 36 |
A |
1389 |
1341 |
4.42 |
|
|
|
| 37 |
A |
1337 |
1291 |
0.87 |
|
|
|
| 38 |
A |
1175 |
1135 |
1.23 |
|
|
|
| 39 |
A |
1052 |
1016 |
1.92 |
|
|
|
| 40 |
A |
992 |
958 |
0.61 |
|
|
|
| 41 |
A |
934 |
902 |
0.54 |
|
|
|
| 42 |
A |
845 |
816 |
0.18 |
|
|
|
| 43 |
A |
440 |
424 |
2.32 |
|
|
|
| 44 |
A |
343 |
331 |
25.69 |
|
|
|
| 45 |
A |
298 |
288 |
6.99 |
|
|
|
| 46 |
A |
270 |
260 |
78.97 |
|
|
|
| 47 |
A |
221 |
214 |
0.22 |
|
|
|
| 48 |
A |
110 |
106 |
2.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36367.2 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 35119.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.255 |
|
|
|
| 2 |
H |
0.158 |
|
|
|
| 3 |
H |
0.158 |
|
|
|
| 4 |
N |
-0.483 |
|
|
|
| 5 |
H |
0.252 |
|
|
|
| 6 |
H |
0.252 |
|
|
|
| 7 |
N |
-0.540 |
|
|
|
| 8 |
H |
0.250 |
|
|
|
| 9 |
H |
0.250 |
|
|
|
| 10 |
C |
-0.401 |
|
|
|
| 11 |
C |
-0.539 |
|
|
|
| 12 |
C |
-0.539 |
|
|
|
| 13 |
H |
0.146 |
|
|
|
| 14 |
H |
0.146 |
|
|
|
| 15 |
H |
0.131 |
|
|
|
| 16 |
H |
0.131 |
|
|
|
| 17 |
H |
0.186 |
|
|
|
| 18 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.320 |
0.030 |
0.000 |
0.321 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.774 |
4.965 |
0.000 |
| y |
4.965 |
-49.265 |
0.000 |
| z |
0.000 |
0.000 |
-36.808 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.262 |
4.965 |
0.000 |
| y |
4.965 |
-12.473 |
0.000 |
| z |
0.000 |
0.000 |
6.211 |
|
| Polar |
|---|
| 3z2-r2 | 12.422 |
|---|
| x2-y2 | 12.490 |
|---|
| xy | 4.965 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.406 |
0.416 |
0.000 |
| y |
0.416 |
8.864 |
0.000 |
| z |
0.000 |
0.000 |
8.756 |
<r2> (average value of r
2) Å
2
| <r2> |
156.363 |
| (<r2>)1/2 |
12.505 |