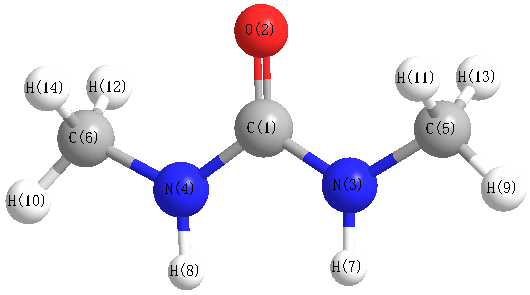
Vibrational Frequencies calculated at B3LYP/CEP-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3585 |
3462 |
8.52 |
|
|
|
| 2 |
A |
3163 |
3055 |
22.34 |
|
|
|
| 3 |
A |
3108 |
3002 |
18.82 |
|
|
|
| 4 |
A |
3033 |
2929 |
5.32 |
|
|
|
| 5 |
A |
1770 |
1709 |
281.54 |
|
|
|
| 6 |
A |
1516 |
1464 |
20.11 |
|
|
|
| 7 |
A |
1490 |
1439 |
0.06 |
|
|
|
| 8 |
A |
1461 |
1411 |
7.63 |
|
|
|
| 9 |
A |
1447 |
1397 |
3.18 |
|
|
|
| 10 |
A |
1186 |
1145 |
0.16 |
|
|
|
| 11 |
A |
1158 |
1119 |
1.49 |
|
|
|
| 12 |
A |
1125 |
1087 |
8.11 |
|
|
|
| 13 |
A |
911 |
880 |
1.84 |
|
|
|
| 14 |
A |
538 |
519 |
97.40 |
|
|
|
| 15 |
A |
427 |
412 |
45.13 |
|
|
|
| 16 |
A |
221 |
214 |
4.31 |
|
|
|
| 17 |
A |
188 |
181 |
0.14 |
|
|
|
| 18 |
A |
111 |
107 |
0.81 |
|
|
|
| 19 |
B |
3582 |
3459 |
7.15 |
|
|
|
| 20 |
B |
3163 |
3055 |
10.14 |
|
|
|
| 21 |
B |
3108 |
3002 |
86.54 |
|
|
|
| 22 |
B |
3032 |
2928 |
146.39 |
|
|
|
| 23 |
B |
1546 |
1493 |
318.10 |
|
|
|
| 24 |
B |
1498 |
1447 |
7.11 |
|
|
|
| 25 |
B |
1482 |
1431 |
91.26 |
|
|
|
| 26 |
B |
1452 |
1402 |
10.39 |
|
|
|
| 27 |
B |
1242 |
1200 |
231.93 |
|
|
|
| 28 |
B |
1141 |
1102 |
29.57 |
|
|
|
| 29 |
B |
1126 |
1087 |
3.37 |
|
|
|
| 30 |
B |
1027 |
992 |
2.72 |
|
|
|
| 31 |
B |
750 |
724 |
27.75 |
|
|
|
| 32 |
B |
717 |
693 |
21.45 |
|
|
|
| 33 |
B |
487 |
471 |
120.76 |
|
|
|
| 34 |
B |
315 |
304 |
43.08 |
|
|
|
| 35 |
B |
125 |
120 |
4.16 |
|
|
|
| 36 |
B |
122 |
118 |
5.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26176.3 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 25278.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.359 |
|
|
|
| 2 |
O |
-0.172 |
|
|
|
| 3 |
N |
-0.125 |
|
|
|
| 4 |
N |
-0.125 |
|
|
|
| 5 |
C |
-0.477 |
|
|
|
| 6 |
C |
-0.477 |
|
|
|
| 7 |
H |
0.291 |
|
|
|
| 8 |
H |
0.291 |
|
|
|
| 9 |
H |
0.195 |
|
|
|
| 10 |
H |
0.195 |
|
|
|
| 11 |
H |
0.212 |
|
|
|
| 12 |
H |
0.212 |
|
|
|
| 13 |
H |
0.169 |
|
|
|
| 14 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.662 |
3.662 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.123 |
-2.410 |
0.000 |
| y |
-2.410 |
-29.458 |
0.000 |
| z |
0.000 |
0.000 |
-37.019 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.885 |
-2.410 |
0.000 |
| y |
-2.410 |
7.613 |
0.000 |
| z |
0.000 |
0.000 |
-3.728 |
|
| Polar |
|---|
| 3z2-r2 | -7.456 |
|---|
| x2-y2 | -7.665 |
|---|
| xy | -2.410 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.276 |
-0.456 |
0.000 |
| y |
-0.456 |
9.313 |
0.000 |
| z |
0.000 |
0.000 |
7.177 |
<r2> (average value of r
2) Å
2
| <r2> |
160.432 |
| (<r2>)1/2 |
12.666 |