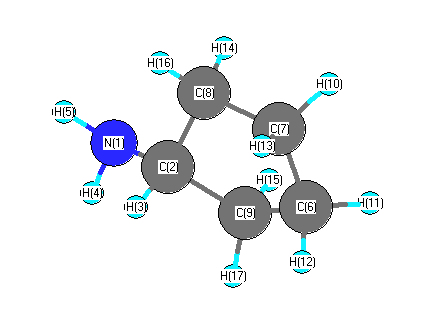
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3469 |
3380 |
5.75 |
|
|
|
| 2 |
A |
3075 |
2996 |
157.73 |
|
|
|
| 3 |
A |
3049 |
2971 |
10.84 |
|
|
|
| 4 |
A |
3018 |
2941 |
34.79 |
|
|
|
| 5 |
A |
2997 |
2920 |
69.07 |
|
|
|
| 6 |
A |
2869 |
2796 |
99.39 |
|
|
|
| 7 |
A |
1692 |
1649 |
39.93 |
|
|
|
| 8 |
A |
1541 |
1502 |
3.23 |
|
|
|
| 9 |
A |
1523 |
1484 |
5.42 |
|
|
|
| 10 |
A |
1391 |
1356 |
27.23 |
|
|
|
| 11 |
A |
1334 |
1300 |
1.20 |
|
|
|
| 12 |
A |
1307 |
1274 |
0.90 |
|
|
|
| 13 |
A |
1239 |
1207 |
2.67 |
|
|
|
| 14 |
A |
1196 |
1165 |
0.25 |
|
|
|
| 15 |
A |
1121 |
1092 |
15.39 |
|
|
|
| 16 |
A |
1054 |
1027 |
0.42 |
|
|
|
| 17 |
A |
984 |
959 |
3.04 |
|
|
|
| 18 |
A |
891 |
868 |
2.25 |
|
|
|
| 19 |
A |
864 |
842 |
0.72 |
|
|
|
| 20 |
A |
768 |
749 |
6.10 |
|
|
|
| 21 |
A |
670 |
653 |
192.34 |
|
|
|
| 22 |
A |
534 |
520 |
4.87 |
|
|
|
| 23 |
A |
459 |
447 |
17.11 |
|
|
|
| 24 |
A |
173 |
169 |
0.80 |
|
|
|
| 25 |
A |
3591 |
3500 |
0.60 |
|
|
|
| 26 |
A |
3056 |
2978 |
16.01 |
|
|
|
| 27 |
A |
3033 |
2956 |
35.80 |
|
|
|
| 28 |
A |
2999 |
2923 |
109.40 |
|
|
|
| 29 |
A |
2990 |
2914 |
3.89 |
|
|
|
| 30 |
A |
1520 |
1481 |
4.65 |
|
|
|
| 31 |
A |
1512 |
1473 |
1.00 |
|
|
|
| 32 |
A |
1384 |
1349 |
3.54 |
|
|
|
| 33 |
A |
1334 |
1300 |
0.23 |
|
|
|
| 34 |
A |
1309 |
1275 |
0.49 |
|
|
|
| 35 |
A |
1271 |
1238 |
0.66 |
|
|
|
| 36 |
A |
1212 |
1181 |
0.03 |
|
|
|
| 37 |
A |
1197 |
1166 |
1.55 |
|
|
|
| 38 |
A |
1030 |
1004 |
1.00 |
|
|
|
| 39 |
A |
991 |
966 |
0.11 |
|
|
|
| 40 |
A |
935 |
911 |
4.15 |
|
|
|
| 41 |
A |
823 |
802 |
0.01 |
|
|
|
| 42 |
A |
636 |
619 |
0.04 |
|
|
|
| 43 |
A |
346 |
337 |
9.01 |
|
|
|
| 44 |
A |
201 |
196 |
39.92 |
|
|
|
| 45 |
A |
53 |
51 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34318.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 33443.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.592 |
|
|
|
| 2 |
C |
0.044 |
|
|
|
| 3 |
H |
0.101 |
|
|
|
| 4 |
H |
0.261 |
|
|
|
| 5 |
H |
0.261 |
|
|
|
| 6 |
C |
-0.260 |
|
|
|
| 7 |
C |
-0.260 |
|
|
|
| 8 |
C |
-0.292 |
|
|
|
| 9 |
C |
-0.292 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.128 |
|
|
|
| 13 |
H |
0.128 |
|
|
|
| 14 |
H |
0.134 |
|
|
|
| 15 |
H |
0.134 |
|
|
|
| 16 |
H |
0.126 |
|
|
|
| 17 |
H |
0.126 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.969 |
0.759 |
0.000 |
1.231 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.524 |
0.693 |
0.000 |
| y |
0.693 |
-35.467 |
0.000 |
| z |
0.000 |
0.000 |
-38.327 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.627 |
0.693 |
0.000 |
| y |
0.693 |
4.959 |
0.000 |
| z |
0.000 |
0.000 |
0.669 |
|
| Polar |
|---|
| 3z2-r2 | 1.337 |
|---|
| x2-y2 | -7.057 |
|---|
| xy | 0.693 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.789 |
-0.761 |
0.000 |
| y |
-0.761 |
9.911 |
0.000 |
| z |
0.000 |
0.000 |
9.596 |
<r2> (average value of r
2) Å
2
| <r2> |
147.196 |
| (<r2>)1/2 |
12.132 |