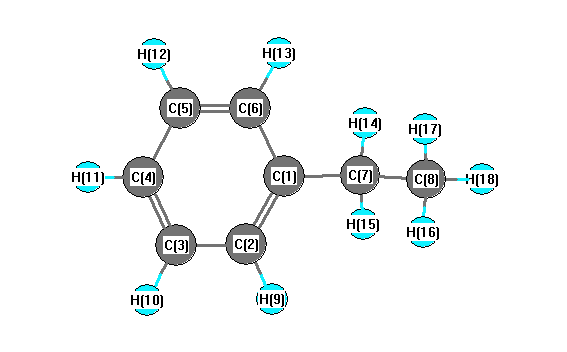
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3176 |
3095 |
30.37 |
|
|
|
| 2 |
A' |
3148 |
3068 |
17.64 |
|
|
|
| 3 |
A' |
3131 |
3051 |
14.18 |
|
|
|
| 4 |
A' |
3070 |
2992 |
74.01 |
|
|
|
| 5 |
A' |
2992 |
2915 |
42.22 |
|
|
|
| 6 |
A' |
2982 |
2906 |
54.86 |
|
|
|
| 7 |
A' |
1631 |
1589 |
6.97 |
|
|
|
| 8 |
A' |
1532 |
1493 |
7.88 |
|
|
|
| 9 |
A' |
1519 |
1480 |
9.42 |
|
|
|
| 10 |
A' |
1511 |
1472 |
4.24 |
|
|
|
| 11 |
A' |
1425 |
1389 |
3.98 |
|
|
|
| 12 |
A' |
1350 |
1315 |
2.54 |
|
|
|
| 13 |
A' |
1225 |
1193 |
0.06 |
|
|
|
| 14 |
A' |
1207 |
1176 |
0.34 |
|
|
|
| 15 |
A' |
1084 |
1056 |
9.16 |
|
|
|
| 16 |
A' |
1038 |
1011 |
3.69 |
|
|
|
| 17 |
A' |
1016 |
990 |
0.38 |
|
|
|
| 18 |
A' |
1005 |
979 |
0.00 |
|
|
|
| 19 |
A' |
966 |
942 |
2.29 |
|
|
|
| 20 |
A' |
946 |
922 |
2.56 |
|
|
|
| 21 |
A' |
790 |
770 |
47.89 |
|
|
|
| 22 |
A' |
763 |
744 |
12.10 |
|
|
|
| 23 |
A' |
715 |
697 |
29.75 |
|
|
|
| 24 |
A' |
564 |
550 |
7.87 |
|
|
|
| 25 |
A' |
497 |
484 |
4.81 |
|
|
|
| 26 |
A' |
310 |
302 |
0.30 |
|
|
|
| 27 |
A' |
134 |
131 |
0.90 |
|
|
|
| 28 |
A" |
3159 |
3078 |
74.66 |
|
|
|
| 29 |
A" |
3133 |
3053 |
8.78 |
|
|
|
| 30 |
A" |
3078 |
3000 |
55.29 |
|
|
|
| 31 |
A" |
3035 |
2957 |
10.75 |
|
|
|
| 32 |
A" |
1608 |
1567 |
1.03 |
|
|
|
| 33 |
A" |
1514 |
1476 |
9.52 |
|
|
|
| 34 |
A" |
1469 |
1432 |
6.11 |
|
|
|
| 35 |
A" |
1359 |
1325 |
0.45 |
|
|
|
| 36 |
A" |
1345 |
1310 |
0.18 |
|
|
|
| 37 |
A" |
1272 |
1240 |
0.07 |
|
|
|
| 38 |
A" |
1189 |
1158 |
0.08 |
|
|
|
| 39 |
A" |
1120 |
1092 |
3.28 |
|
|
|
| 40 |
A" |
1050 |
1023 |
3.57 |
|
|
|
| 41 |
A" |
992 |
967 |
0.02 |
|
|
|
| 42 |
A" |
882 |
860 |
0.01 |
|
|
|
| 43 |
A" |
798 |
777 |
0.79 |
|
|
|
| 44 |
A" |
631 |
615 |
0.09 |
|
|
|
| 45 |
A" |
416 |
406 |
0.01 |
|
|
|
| 46 |
A" |
347 |
338 |
0.09 |
|
|
|
| 47 |
A" |
210 |
205 |
0.04 |
|
|
|
| 48 |
A" |
31 |
31 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34181.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 33310.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.410 |
|
|
|
| 2 |
C |
-0.283 |
|
|
|
| 3 |
C |
-0.127 |
|
|
|
| 4 |
C |
-0.172 |
|
|
|
| 5 |
C |
-0.127 |
|
|
|
| 6 |
C |
-0.283 |
|
|
|
| 7 |
C |
-0.444 |
|
|
|
| 8 |
C |
-0.476 |
|
|
|
| 9 |
H |
0.166 |
|
|
|
| 10 |
H |
0.145 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
| 12 |
H |
0.145 |
|
|
|
| 13 |
H |
0.166 |
|
|
|
| 14 |
H |
0.144 |
|
|
|
| 15 |
H |
0.144 |
|
|
|
| 16 |
H |
0.148 |
|
|
|
| 17 |
H |
0.148 |
|
|
|
| 18 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.067 |
0.376 |
0.000 |
0.382 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-53.460 |
-0.056 |
0.000 |
| y |
-0.056 |
-45.282 |
0.000 |
| z |
0.000 |
0.000 |
-45.384 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.127 |
-0.056 |
0.000 |
| y |
-0.056 |
4.141 |
0.000 |
| z |
0.000 |
0.000 |
3.987 |
|
| Polar |
|---|
| 3z2-r2 | 7.973 |
|---|
| x2-y2 | -8.179 |
|---|
| xy | -0.056 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.856 |
0.750 |
0.000 |
| y |
0.750 |
16.848 |
0.000 |
| z |
0.000 |
0.000 |
13.999 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |