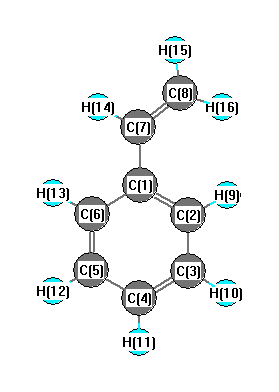
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3211 |
3129 |
32.43 |
|
|
|
| 2 |
A' |
3180 |
3099 |
24.09 |
|
|
|
| 3 |
A' |
3168 |
3087 |
61.82 |
|
|
|
| 4 |
A' |
3157 |
3076 |
30.60 |
|
|
|
| 5 |
A' |
3144 |
3064 |
0.40 |
|
|
|
| 6 |
A' |
3137 |
3057 |
12.23 |
|
|
|
| 7 |
A' |
3117 |
3038 |
7.83 |
|
|
|
| 8 |
A' |
3104 |
3025 |
20.55 |
|
|
|
| 9 |
A' |
1665 |
1622 |
13.76 |
|
|
|
| 10 |
A' |
1630 |
1588 |
2.65 |
|
|
|
| 11 |
A' |
1602 |
1561 |
4.20 |
|
|
|
| 12 |
A' |
1519 |
1480 |
12.90 |
|
|
|
| 13 |
A' |
1478 |
1440 |
2.25 |
|
|
|
| 14 |
A' |
1454 |
1417 |
8.21 |
|
|
|
| 15 |
A' |
1368 |
1333 |
3.25 |
|
|
|
| 16 |
A' |
1354 |
1319 |
1.13 |
|
|
|
| 17 |
A' |
1322 |
1288 |
1.45 |
|
|
|
| 18 |
A' |
1223 |
1192 |
3.14 |
|
|
|
| 19 |
A' |
1213 |
1182 |
0.05 |
|
|
|
| 20 |
A' |
1190 |
1160 |
0.21 |
|
|
|
| 21 |
A' |
1106 |
1078 |
5.56 |
|
|
|
| 22 |
A' |
1052 |
1025 |
1.91 |
|
|
|
| 23 |
A' |
1033 |
1006 |
6.05 |
|
|
|
| 24 |
A' |
1001 |
976 |
0.69 |
|
|
|
| 25 |
A' |
775 |
755 |
0.24 |
|
|
|
| 26 |
A' |
630 |
614 |
0.05 |
|
|
|
| 27 |
A' |
558 |
544 |
4.32 |
|
|
|
| 28 |
A' |
443 |
432 |
0.10 |
|
|
|
| 29 |
A' |
232 |
226 |
0.42 |
|
|
|
| 30 |
A" |
1063 |
1036 |
25.00 |
|
|
|
| 31 |
A" |
1021 |
995 |
0.01 |
|
|
|
| 32 |
A" |
1000 |
975 |
0.01 |
|
|
|
| 33 |
A" |
968 |
943 |
7.26 |
|
|
|
| 34 |
A" |
944 |
920 |
58.57 |
|
|
|
| 35 |
A" |
885 |
863 |
0.41 |
|
|
|
| 36 |
A" |
833 |
812 |
50.28 |
|
|
|
| 37 |
A" |
728 |
709 |
46.20 |
|
|
|
| 38 |
A" |
667 |
650 |
0.68 |
|
|
|
| 39 |
A" |
461 |
449 |
8.42 |
|
|
|
| 40 |
A" |
422 |
411 |
0.18 |
|
|
|
| 41 |
A" |
217 |
212 |
1.95 |
|
|
|
| 42 |
A" |
73 |
71 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29173.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 28429.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.420 |
|
|
|
| 2 |
C |
-0.260 |
|
|
|
| 3 |
C |
-0.157 |
|
|
|
| 4 |
C |
-0.162 |
|
|
|
| 5 |
C |
-0.140 |
|
|
|
| 6 |
C |
-0.368 |
|
|
|
| 7 |
C |
-0.088 |
|
|
|
| 8 |
C |
-0.426 |
|
|
|
| 9 |
H |
0.167 |
|
|
|
| 10 |
H |
0.148 |
|
|
|
| 11 |
H |
0.150 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
H |
0.175 |
|
|
|
| 14 |
H |
0.146 |
|
|
|
| 15 |
H |
0.117 |
|
|
|
| 16 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.104 |
-0.184 |
0.000 |
0.211 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.550 |
0.319 |
0.000 |
| y |
0.319 |
-42.950 |
0.000 |
| z |
0.000 |
0.000 |
-53.537 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.694 |
0.319 |
0.000 |
| y |
0.319 |
5.093 |
0.000 |
| z |
0.000 |
0.000 |
-10.787 |
|
| Polar |
|---|
| 3z2-r2 | -21.574 |
|---|
| x2-y2 | 0.400 |
|---|
| xy | 0.319 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
15.663 |
-2.974 |
0.000 |
| y |
-2.974 |
18.834 |
0.000 |
| z |
0.000 |
0.000 |
5.757 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |