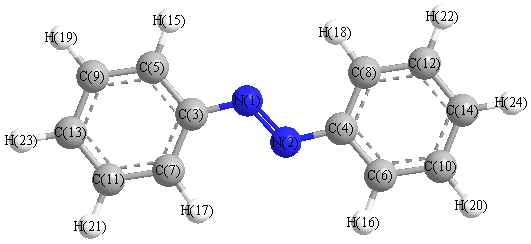
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3205 |
3123 |
0.00 |
|
|
|
| 2 |
Ag |
3189 |
3107 |
0.00 |
|
|
|
| 3 |
Ag |
3174 |
3093 |
0.00 |
|
|
|
| 4 |
Ag |
3160 |
3079 |
0.00 |
|
|
|
| 5 |
Ag |
3146 |
3066 |
0.00 |
|
|
|
| 6 |
Ag |
1621 |
1580 |
0.00 |
|
|
|
| 7 |
Ag |
1608 |
1567 |
0.00 |
|
|
|
| 8 |
Ag |
1500 |
1462 |
0.00 |
|
|
|
| 9 |
Ag |
1480 |
1443 |
0.00 |
|
|
|
| 10 |
Ag |
1435 |
1399 |
0.00 |
|
|
|
| 11 |
Ag |
1356 |
1322 |
0.00 |
|
|
|
| 12 |
Ag |
1341 |
1307 |
0.00 |
|
|
|
| 13 |
Ag |
1202 |
1172 |
0.00 |
|
|
|
| 14 |
Ag |
1188 |
1158 |
0.00 |
|
|
|
| 15 |
Ag |
1154 |
1125 |
0.00 |
|
|
|
| 16 |
Ag |
1089 |
1061 |
0.00 |
|
|
|
| 17 |
Ag |
1029 |
1003 |
0.00 |
|
|
|
| 18 |
Ag |
1003 |
977 |
0.00 |
|
|
|
| 19 |
Ag |
911 |
887 |
0.00 |
|
|
|
| 20 |
Ag |
676 |
659 |
0.00 |
|
|
|
| 21 |
Ag |
623 |
607 |
0.00 |
|
|
|
| 22 |
Ag |
300 |
293 |
0.00 |
|
|
|
| 23 |
Ag |
216 |
211 |
0.00 |
|
|
|
| 24 |
Au |
1043 |
1016 |
3.01 |
|
|
|
| 25 |
Au |
1024 |
998 |
0.02 |
|
|
|
| 26 |
Au |
986 |
961 |
10.52 |
|
|
|
| 27 |
Au |
890 |
868 |
0.14 |
|
|
|
| 28 |
Au |
821 |
800 |
93.73 |
|
|
|
| 29 |
Au |
713 |
695 |
90.26 |
|
|
|
| 30 |
Au |
564 |
550 |
12.58 |
|
|
|
| 31 |
Au |
424 |
413 |
0.00 |
|
|
|
| 32 |
Au |
311 |
303 |
0.73 |
|
|
|
| 33 |
Au |
64 |
63 |
1.73 |
|
|
|
| 34 |
Au |
33 |
32 |
0.06 |
|
|
|
| 35 |
Bg |
1041 |
1015 |
0.00 |
|
|
|
| 36 |
Bg |
1025 |
999 |
0.00 |
|
|
|
| 37 |
Bg |
982 |
957 |
0.00 |
|
|
|
| 38 |
Bg |
891 |
869 |
0.00 |
|
|
|
| 39 |
Bg |
805 |
784 |
0.00 |
|
|
|
| 40 |
Bg |
704 |
686 |
0.00 |
|
|
|
| 41 |
Bg |
503 |
491 |
0.00 |
|
|
|
| 42 |
Bg |
427 |
416 |
0.00 |
|
|
|
| 43 |
Bg |
272 |
265 |
0.00 |
|
|
|
| 44 |
Bg |
126 |
123 |
0.00 |
|
|
|
| 45 |
Bu |
3205 |
3123 |
17.20 |
|
|
|
| 46 |
Bu |
3188 |
3107 |
57.22 |
|
|
|
| 47 |
Bu |
3174 |
3093 |
98.29 |
|
|
|
| 48 |
Bu |
3160 |
3079 |
41.59 |
|
|
|
| 49 |
Bu |
3146 |
3066 |
2.03 |
|
|
|
| 50 |
Bu |
1622 |
1581 |
4.42 |
|
|
|
| 51 |
Bu |
1604 |
1563 |
5.04 |
|
|
|
| 52 |
Bu |
1505 |
1467 |
14.42 |
|
|
|
| 53 |
Bu |
1469 |
1431 |
15.69 |
|
|
|
| 54 |
Bu |
1354 |
1319 |
10.41 |
|
|
|
| 55 |
Bu |
1336 |
1302 |
1.84 |
|
|
|
| 56 |
Bu |
1240 |
1208 |
22.55 |
|
|
|
| 57 |
Bu |
1188 |
1158 |
0.34 |
|
|
|
| 58 |
Bu |
1180 |
1150 |
42.71 |
|
|
|
| 59 |
Bu |
1093 |
1065 |
15.43 |
|
|
|
| 60 |
Bu |
1028 |
1002 |
17.41 |
|
|
|
| 61 |
Bu |
1004 |
978 |
0.82 |
|
|
|
| 62 |
Bu |
820 |
799 |
0.43 |
|
|
|
| 63 |
Bu |
628 |
612 |
0.49 |
|
|
|
| 64 |
Bu |
541 |
528 |
7.30 |
|
|
|
| 65 |
Bu |
519 |
505 |
28.24 |
|
|
|
| 66 |
Bu |
84 |
81 |
2.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41671.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 40608.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
0.114 |
|
|
|
| 2 |
N |
0.114 |
|
|
|
| 3 |
C |
-0.161 |
|
|
|
| 4 |
C |
-0.161 |
|
|
|
| 5 |
C |
-0.196 |
|
|
|
| 6 |
C |
-0.196 |
|
|
|
| 7 |
C |
-0.192 |
|
|
|
| 8 |
C |
-0.192 |
|
|
|
| 9 |
C |
-0.092 |
|
|
|
| 10 |
C |
-0.092 |
|
|
|
| 11 |
C |
-0.076 |
|
|
|
| 12 |
C |
-0.076 |
|
|
|
| 13 |
C |
-0.189 |
|
|
|
| 14 |
C |
-0.189 |
|
|
|
| 15 |
H |
0.157 |
|
|
|
| 16 |
H |
0.157 |
|
|
|
| 17 |
H |
0.167 |
|
|
|
| 18 |
H |
0.167 |
|
|
|
| 19 |
H |
0.158 |
|
|
|
| 20 |
H |
0.158 |
|
|
|
| 21 |
H |
0.157 |
|
|
|
| 22 |
H |
0.157 |
|
|
|
| 23 |
H |
0.154 |
|
|
|
| 24 |
H |
0.154 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-66.087 |
5.868 |
0.000 |
| y |
5.868 |
-69.190 |
0.000 |
| z |
0.000 |
0.000 |
-87.867 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
12.442 |
5.868 |
0.000 |
| y |
5.868 |
7.787 |
0.000 |
| z |
0.000 |
0.000 |
-20.228 |
|
| Polar |
|---|
| 3z2-r2 | -40.456 |
|---|
| x2-y2 | 3.103 |
|---|
| xy | 5.868 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
36.340 |
11.670 |
0.000 |
| y |
11.670 |
32.150 |
0.000 |
| z |
0.000 |
0.000 |
9.169 |
<r2> (average value of r
2) Å
2
| <r2> |
836.008 |
| (<r2>)1/2 |
28.914 |