Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -40.448134 |
| Energy at 298.15K | -40.458205 |
| HF Energy | -40.448134 |
| Nuclear repulsion energy | 59.178615 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3092 |
3013 |
0.00 |
|
|
|
| 2 |
Ag |
1499 |
1460 |
0.00 |
|
|
|
| 3 |
Ag |
1291 |
1259 |
0.00 |
|
|
|
| 4 |
Ag |
1067 |
1040 |
0.00 |
|
|
|
| 5 |
Ag |
620 |
604 |
0.00 |
|
|
|
| 6 |
Ag |
176 |
171 |
0.00 |
|
|
|
| 7 |
Au |
3198 |
3116 |
10.07 |
|
|
|
| 8 |
Au |
1100 |
1072 |
7.75 |
|
|
|
| 9 |
Au |
774 |
755 |
8.53 |
|
|
|
| 10 |
Au |
101 |
98 |
5.18 |
|
|
|
| 11 |
Bg |
3172 |
3091 |
0.00 |
|
|
|
| 12 |
Bg |
1286 |
1254 |
0.00 |
|
|
|
| 13 |
Bg |
948 |
923 |
0.00 |
|
|
|
| 14 |
Bu |
3096 |
3017 |
15.24 |
|
|
|
| 15 |
Bu |
1493 |
1455 |
9.65 |
|
|
|
| 16 |
Bu |
1228 |
1197 |
71.62 |
|
|
|
| 17 |
Bu |
546 |
532 |
99.58 |
|
|
|
| 18 |
Bu |
169 |
165 |
9.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12428.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12111.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.510 |
0.566 |
0.000 |
| C2 |
-0.510 |
-0.566 |
0.000 |
| Br3 |
-0.510 |
2.339 |
0.000 |
| Br4 |
0.510 |
-2.339 |
0.000 |
| H5 |
1.124 |
0.589 |
0.900 |
| H6 |
1.124 |
0.589 |
-0.900 |
| H7 |
-1.124 |
-0.589 |
0.900 |
| H8 |
-1.124 |
-0.589 |
-0.900 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5236 | 2.0446 | 2.9051 | 1.0898 | 1.0898 | 2.1939 | 2.1939 |
C2 | 1.5236 | | 2.9051 | 2.0446 | 2.1939 | 2.1939 | 1.0898 | 1.0898 | Br3 | 2.0446 | 2.9051 | | 4.7873 | 2.5574 | 2.5574 | 3.1239 | 3.1239 | Br4 | 2.9051 | 2.0446 | 4.7873 | | 3.1239 | 3.1239 | 2.5574 | 2.5574 | H5 | 1.0898 | 2.1939 | 2.5574 | 3.1239 | | 1.7992 | 2.5383 | 3.1113 | H6 | 1.0898 | 2.1939 | 2.5574 | 3.1239 | 1.7992 | | 3.1113 | 2.5383 | H7 | 2.1939 | 1.0898 | 3.1239 | 2.5574 | 2.5383 | 3.1113 | | 1.7992 | H8 | 2.1939 | 1.0898 | 3.1239 | 2.5574 | 3.1113 | 2.5383 | 1.7992 | |
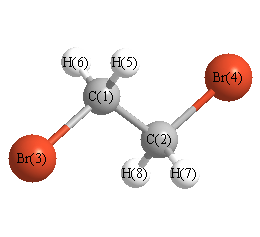 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.126 |
|
C1 |
C2 |
H7 |
113.123 |
| C1 |
C2 |
H8 |
113.123 |
|
C2 |
C1 |
Br3 |
108.126 |
| C2 |
C1 |
H5 |
113.123 |
|
C2 |
C1 |
H6 |
113.123 |
| Br3 |
C1 |
H5 |
105.256 |
|
Br3 |
C1 |
H6 |
105.256 |
| Br4 |
C2 |
H7 |
105.256 |
|
Br4 |
C2 |
H8 |
105.256 |
| H5 |
C1 |
H6 |
111.273 |
|
H7 |
C2 |
H8 |
111.273 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.491 |
|
|
|
| 2 |
C |
-0.491 |
|
|
|
| 3 |
Br |
0.045 |
|
|
|
| 4 |
Br |
0.045 |
|
|
|
| 5 |
H |
0.223 |
|
|
|
| 6 |
H |
0.223 |
|
|
|
| 7 |
H |
0.223 |
|
|
|
| 8 |
H |
0.223 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.472 |
1.855 |
0.000 |
| y |
1.855 |
-51.068 |
0.000 |
| z |
0.000 |
0.000 |
-43.081 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.602 |
1.855 |
0.000 |
| y |
1.855 |
-8.291 |
0.000 |
| z |
0.000 |
0.000 |
3.689 |
|
| Polar |
|---|
| 3z2-r2 | 7.377 |
|---|
| x2-y2 | 8.596 |
|---|
| xy | 1.855 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.970 |
-3.055 |
0.000 |
| y |
-3.055 |
13.849 |
0.000 |
| z |
0.000 |
0.000 |
4.047 |
<r2> (average value of r
2) Å
2
| <r2> |
122.981 |
| (<r2>)1/2 |
11.090 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -40.443114 |
| Energy at 298.15K | |
| HF Energy | -40.443114 |
| Nuclear repulsion energy | 60.077319 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
3065 |
0.97 |
116.47 |
0.75 |
0.86 |
| 2 |
A |
3064 |
2986 |
26.76 |
293.46 |
0.01 |
0.03 |
| 3 |
A |
1476 |
1438 |
1.26 |
6.60 |
0.67 |
0.80 |
| 4 |
A |
1312 |
1279 |
28.00 |
10.02 |
0.52 |
0.68 |
| 5 |
A |
1185 |
1155 |
3.68 |
36.07 |
0.72 |
0.84 |
| 6 |
A |
1044 |
1017 |
1.10 |
4.72 |
0.65 |
0.79 |
| 7 |
A |
908 |
885 |
12.74 |
24.23 |
0.50 |
0.67 |
| 8 |
A |
518 |
505 |
11.74 |
26.02 |
0.09 |
0.17 |
| 9 |
A |
219 |
214 |
1.54 |
3.39 |
0.57 |
0.72 |
| 10 |
A |
72 |
70 |
0.30 |
3.23 |
0.74 |
0.85 |
| 11 |
B |
3159 |
3078 |
8.96 |
33.23 |
0.75 |
0.86 |
| 12 |
B |
3051 |
2973 |
4.86 |
65.34 |
0.75 |
0.86 |
| 13 |
B |
1470 |
1432 |
12.47 |
22.61 |
0.75 |
0.86 |
| 14 |
B |
1290 |
1257 |
85.08 |
3.58 |
0.75 |
0.86 |
| 15 |
B |
1119 |
1090 |
5.83 |
14.03 |
0.75 |
0.86 |
| 16 |
B |
854 |
832 |
28.39 |
2.37 |
0.75 |
0.86 |
| 17 |
B |
550 |
536 |
25.21 |
20.34 |
0.75 |
0.86 |
| 18 |
B |
338 |
329 |
9.60 |
5.54 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12385.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12070.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.327 |
0.690 |
1.208 |
| C2 |
-0.327 |
-0.690 |
1.208 |
| Br3 |
-0.327 |
1.875 |
-0.299 |
| Br4 |
0.327 |
-1.875 |
-0.299 |
| H5 |
0.057 |
1.233 |
2.120 |
| H6 |
1.411 |
0.644 |
1.097 |
| H7 |
-0.057 |
-1.233 |
2.120 |
| H8 |
-1.411 |
-0.644 |
1.097 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5269 | 2.0263 | 2.9752 | 1.0948 | 1.0907 | 2.1623 | 2.1942 |
C2 | 1.5269 | | 2.9752 | 2.0263 | 2.1623 | 2.1942 | 1.0948 | 1.0907 | Br3 | 2.0263 | 2.9752 | | 3.8070 | 2.5323 | 2.5465 | 3.9477 | 3.0776 | Br4 | 2.9752 | 2.0263 | 3.8070 | | 3.9477 | 3.0776 | 2.5323 | 2.5465 | H5 | 1.0948 | 2.1623 | 2.5323 | 3.9477 | | 1.7961 | 2.4685 | 2.5939 | H6 | 1.0907 | 2.1942 | 2.5465 | 3.0776 | 1.7961 | | 2.5939 | 3.1027 | H7 | 2.1623 | 1.0948 | 3.9477 | 2.5323 | 2.4685 | 2.5939 | | 1.7961 | H8 | 2.1942 | 1.0907 | 3.0776 | 2.5465 | 2.5939 | 3.1027 | 1.7961 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.963 |
|
C1 |
C2 |
H7 |
110.037 |
| C1 |
C2 |
H8 |
112.849 |
|
C2 |
C1 |
Br3 |
112.963 |
| C2 |
C1 |
H5 |
110.037 |
|
C2 |
C1 |
H6 |
112.849 |
| Br3 |
C1 |
H5 |
104.460 |
|
Br3 |
C1 |
H6 |
105.609 |
| Br4 |
C2 |
H7 |
104.460 |
|
Br4 |
C2 |
H8 |
105.609 |
| H5 |
C1 |
H6 |
110.538 |
|
H7 |
C2 |
H8 |
110.538 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.490 |
|
|
|
| 2 |
C |
-0.490 |
|
|
|
| 3 |
Br |
0.060 |
|
|
|
| 4 |
Br |
0.060 |
|
|
|
| 5 |
H |
0.208 |
|
|
|
| 6 |
H |
0.222 |
|
|
|
| 7 |
H |
0.208 |
|
|
|
| 8 |
H |
0.222 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
3.063 |
3.063 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.790 |
1.102 |
0.000 |
| y |
1.102 |
-48.109 |
0.000 |
| z |
0.000 |
0.000 |
-38.407 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.468 |
1.102 |
0.000 |
| y |
1.102 |
-7.511 |
0.000 |
| z |
0.000 |
0.000 |
7.042 |
|
| Polar |
|---|
| 3z2-r2 | 14.085 |
|---|
| x2-y2 | 5.319 |
|---|
| xy | 1.102 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.861 |
-1.233 |
0.000 |
| y |
-1.233 |
8.812 |
0.000 |
| z |
0.000 |
0.000 |
7.931 |
<r2> (average value of r
2) Å
2
| <r2> |
114.500 |
| (<r2>)1/2 |
10.700 |
