Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -28.597848 |
| Energy at 298.15K | -28.608434 |
| HF Energy | -28.597848 |
| Nuclear repulsion energy | 76.330083 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
117 |
114 |
0.01 |
|
|
|
|
Au |
207 |
202 |
0.00 |
|
|
|
|
Bg |
242 |
235 |
0.00 |
|
|
|
|
Bu |
254 |
247 |
0.05 |
|
|
|
|
Ag |
418 |
407 |
0.00 |
|
|
|
|
Au |
742 |
724 |
4.50 |
|
|
|
|
Bg |
820 |
799 |
0.00 |
|
|
|
|
Ag |
834 |
813 |
0.00 |
|
|
|
|
Au |
968 |
944 |
1.23 |
|
|
|
|
Bu |
984 |
959 |
8.86 |
|
|
|
|
Bu |
1015 |
989 |
0.44 |
|
|
|
|
Ag |
1070 |
1043 |
0.00 |
|
|
|
|
Ag |
1170 |
1140 |
0.00 |
|
|
|
|
Bg |
1216 |
1185 |
0.00 |
|
|
|
|
Au |
1290 |
1257 |
0.16 |
|
|
|
|
Bg |
1327 |
1293 |
0.00 |
|
|
|
|
Bu |
1332 |
1298 |
2.28 |
|
|
|
|
Ag |
1392 |
1356 |
0.00 |
|
|
|
|
Ag |
1428 |
1392 |
0.00 |
|
|
|
|
Bu |
1430 |
1393 |
9.97 |
|
|
|
|
Ag |
1513 |
1475 |
0.00 |
|
|
|
|
Bu |
1515 |
1477 |
3.38 |
|
|
|
|
Bg |
1520 |
1482 |
0.00 |
|
|
|
|
Au |
1521 |
1482 |
16.63 |
|
|
|
|
Ag |
1526 |
1487 |
0.00 |
|
|
|
|
Bu |
1536 |
1497 |
11.31 |
|
|
|
|
Ag |
2964 |
2888 |
0.00 |
|
|
|
|
Bu |
2972 |
2897 |
97.40 |
|
|
|
|
Bu |
2974 |
2899 |
92.53 |
|
|
|
|
Ag |
2978 |
2902 |
0.00 |
|
|
|
|
Bg |
2995 |
2918 |
0.00 |
|
|
|
|
Au |
3020 |
2943 |
21.06 |
|
|
|
|
Bg |
3057 |
2979 |
0.00 |
|
|
|
|
Ag |
3062 |
2984 |
0.00 |
|
|
|
|
Bu |
3063 |
2985 |
137.15 |
|
|
|
|
Au |
3065 |
2987 |
202.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28768.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 28035.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.428 |
0.648 |
0.000 |
| C2 |
0.428 |
-0.648 |
0.000 |
| C3 |
0.428 |
1.942 |
0.000 |
| C4 |
-0.428 |
-1.942 |
0.000 |
| H5 |
-1.089 |
0.644 |
0.882 |
| H6 |
-1.089 |
0.644 |
-0.882 |
| H7 |
1.089 |
-0.644 |
0.882 |
| H8 |
1.089 |
-0.644 |
-0.882 |
| H9 |
-0.205 |
2.840 |
0.000 |
| H10 |
1.075 |
1.987 |
0.888 |
| H11 |
1.075 |
1.987 |
-0.888 |
| H12 |
0.205 |
-2.840 |
0.000 |
| H13 |
-1.075 |
-1.987 |
0.888 |
| H14 |
-1.075 |
-1.987 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5531 | 1.5508 | 2.5899 | 1.1023 | 1.1023 | 2.1790 | 2.1790 | 2.2034 | 2.1997 | 2.1997 | 3.5453 | 2.8548 | 2.8548 |
C2 | 1.5531 | | 2.5899 | 1.5508 | 2.1790 | 2.1790 | 1.1023 | 1.1023 | 3.5453 | 2.8548 | 2.8548 | 2.2034 | 2.1997 | 2.1997 | C3 | 1.5508 | 2.5899 | | 3.9766 | 2.1820 | 2.1820 | 2.8111 | 2.8111 | 1.0988 | 1.0997 | 1.0997 | 4.7872 | 4.2988 | 4.2988 | C4 | 2.5899 | 1.5508 | 3.9766 | | 2.8111 | 2.8111 | 2.1820 | 2.1820 | 4.7872 | 4.2988 | 4.2988 | 1.0988 | 1.0997 | 1.0997 | H5 | 1.1023 | 2.1790 | 2.1820 | 2.8111 | | 1.7638 | 2.5304 | 3.0845 | 2.5261 | 2.5467 | 3.1012 | 3.8201 | 2.6310 | 3.1707 | H6 | 1.1023 | 2.1790 | 2.1820 | 2.8111 | 1.7638 | | 3.0845 | 2.5304 | 2.5261 | 3.1012 | 2.5467 | 3.8201 | 3.1707 | 2.6310 | H7 | 2.1790 | 1.1023 | 2.8111 | 2.1820 | 2.5304 | 3.0845 | | 1.7638 | 3.8201 | 2.6310 | 3.1707 | 2.5261 | 2.5467 | 3.1012 | H8 | 2.1790 | 1.1023 | 2.8111 | 2.1820 | 3.0845 | 2.5304 | 1.7638 | | 3.8201 | 3.1707 | 2.6310 | 2.5261 | 3.1012 | 2.5467 | H9 | 2.2034 | 3.5453 | 1.0988 | 4.7872 | 2.5261 | 2.5261 | 3.8201 | 3.8201 | | 1.7764 | 1.7764 | 5.6952 | 4.9844 | 4.9844 | H10 | 2.1997 | 2.8548 | 1.0997 | 4.2988 | 2.5467 | 3.1012 | 2.6310 | 3.1707 | 1.7764 | | 1.7754 | 4.9844 | 4.5180 | 4.8543 | H11 | 2.1997 | 2.8548 | 1.0997 | 4.2988 | 3.1012 | 2.5467 | 3.1707 | 2.6310 | 1.7764 | 1.7754 | | 4.9844 | 4.8543 | 4.5180 | H12 | 3.5453 | 2.2034 | 4.7872 | 1.0988 | 3.8201 | 3.8201 | 2.5261 | 2.5261 | 5.6952 | 4.9844 | 4.9844 | | 1.7764 | 1.7764 | H13 | 2.8548 | 2.1997 | 4.2988 | 1.0997 | 2.6310 | 3.1707 | 2.5467 | 3.1012 | 4.9844 | 4.5180 | 4.8543 | 1.7764 | | 1.7754 | H14 | 2.8548 | 2.1997 | 4.2988 | 1.0997 | 3.1707 | 2.6310 | 3.1012 | 2.5467 | 4.9844 | 4.8543 | 4.5180 | 1.7764 | 1.7754 | |
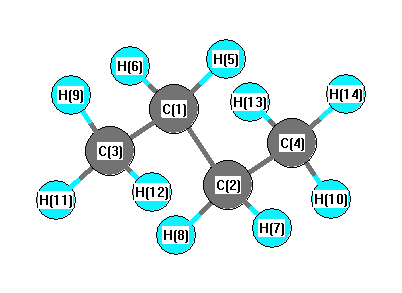 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
113.109 |
|
C1 |
C2 |
H7 |
109.112 |
| C1 |
C2 |
H8 |
109.112 |
|
C1 |
C3 |
H9 |
111.376 |
| C1 |
C3 |
H11 |
111.030 |
|
C1 |
C3 |
H12 |
30.807 |
| C2 |
C1 |
C3 |
113.109 |
|
C2 |
C1 |
H5 |
109.112 |
| C2 |
C1 |
H6 |
109.112 |
|
C2 |
C4 |
H10 |
17.231 |
| C2 |
C4 |
H13 |
111.030 |
|
C2 |
C4 |
H14 |
111.030 |
| C3 |
C1 |
H5 |
109.500 |
|
C3 |
C1 |
H6 |
109.500 |
| C4 |
C2 |
H7 |
109.500 |
|
C4 |
C2 |
H8 |
109.500 |
| H5 |
C1 |
H6 |
106.277 |
|
H7 |
C2 |
H8 |
106.277 |
| H9 |
C3 |
H11 |
107.798 |
|
H9 |
C3 |
H12 |
142.182 |
| H10 |
C4 |
H13 |
94.389 |
|
H10 |
C4 |
H14 |
114.198 |
| H11 |
C3 |
H12 |
93.910 |
|
H13 |
C4 |
H14 |
107.642 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.196 |
|
|
|
| 2 |
C |
-0.196 |
|
|
|
| 3 |
C |
-0.467 |
|
|
|
| 4 |
C |
-0.467 |
|
|
|
| 5 |
H |
0.121 |
|
|
|
| 6 |
H |
0.121 |
|
|
|
| 7 |
H |
0.121 |
|
|
|
| 8 |
H |
0.121 |
|
|
|
| 9 |
H |
0.144 |
|
|
|
| 10 |
H |
0.138 |
|
|
|
| 11 |
H |
0.138 |
|
|
|
| 12 |
H |
0.144 |
|
|
|
| 13 |
H |
0.138 |
|
|
|
| 14 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.021 |
-0.365 |
0.000 |
| y |
-0.365 |
-29.294 |
0.000 |
| z |
0.000 |
0.000 |
-28.254 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.247 |
-0.365 |
0.000 |
| y |
-0.365 |
-0.656 |
0.000 |
| z |
0.000 |
0.000 |
0.903 |
|
| Polar |
|---|
| 3z2-r2 | 1.806 |
|---|
| x2-y2 | 0.273 |
|---|
| xy | -0.365 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.974 |
0.185 |
0.000 |
| y |
0.185 |
8.173 |
0.000 |
| z |
0.000 |
0.000 |
6.774 |
<r2> (average value of r
2) Å
2
| <r2> |
103.771 |
| (<r2>)1/2 |
10.187 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -28.596590 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 77.315708 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3075 |
2996 |
69.39 |
|
|
|
| 2 |
A |
3059 |
2981 |
1.04 |
|
|
|
| 3 |
A |
3010 |
2933 |
40.23 |
|
|
|
| 4 |
A |
2982 |
2906 |
21.14 |
|
|
|
| 5 |
A |
2969 |
2893 |
51.81 |
|
|
|
| 6 |
A |
1534 |
1495 |
1.38 |
|
|
|
| 7 |
A |
1527 |
1488 |
7.18 |
|
|
|
| 8 |
A |
1511 |
1472 |
0.00 |
|
|
|
| 9 |
A |
1433 |
1396 |
5.90 |
|
|
|
| 10 |
A |
1371 |
1336 |
0.96 |
|
|
|
| 11 |
A |
1310 |
1276 |
0.39 |
|
|
|
| 12 |
A |
1200 |
1170 |
0.09 |
|
|
|
| 13 |
A |
1097 |
1069 |
0.20 |
|
|
|
| 14 |
A |
995 |
969 |
0.44 |
|
|
|
| 15 |
A |
832 |
811 |
0.28 |
|
|
|
| 16 |
A |
791 |
771 |
2.00 |
|
|
|
| 17 |
A |
314 |
306 |
0.04 |
|
|
|
| 18 |
A |
253 |
246 |
0.01 |
|
|
|
| 19 |
A |
105 |
103 |
0.01 |
|
|
|
| 20 |
B |
3067 |
2989 |
103.04 |
|
|
|
| 21 |
B |
3061 |
2983 |
144.34 |
|
|
|
| 22 |
B |
3015 |
2938 |
39.98 |
|
|
|
| 23 |
B |
2978 |
2902 |
40.72 |
|
|
|
| 24 |
B |
2964 |
2889 |
41.61 |
|
|
|
| 25 |
B |
1529 |
1490 |
15.13 |
|
|
|
| 26 |
B |
1521 |
1482 |
9.90 |
|
|
|
| 27 |
B |
1511 |
1472 |
0.69 |
|
|
|
| 28 |
B |
1427 |
1390 |
8.69 |
|
|
|
| 29 |
B |
1383 |
1347 |
3.13 |
|
|
|
| 30 |
B |
1290 |
1257 |
0.32 |
|
|
|
| 31 |
B |
1157 |
1128 |
2.97 |
|
|
|
| 32 |
B |
974 |
950 |
3.76 |
|
|
|
| 33 |
B |
963 |
939 |
2.59 |
|
|
|
| 34 |
B |
756 |
737 |
4.94 |
|
|
|
| 35 |
B |
428 |
417 |
0.46 |
|
|
|
| 36 |
B |
202 |
196 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28795.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 28061.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
