Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -35.246556 |
| Energy at 298.15K | |
| HF Energy | -35.246556 |
| Nuclear repulsion energy | 54.406258 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3215 |
3133 |
18.34 |
72.76 |
0.53 |
0.69 |
| 2 |
A' |
3160 |
3079 |
3.62 |
110.00 |
0.32 |
0.49 |
| 3 |
A' |
3105 |
3026 |
8.96 |
81.03 |
0.13 |
0.23 |
| 4 |
A' |
2929 |
2855 |
96.87 |
108.25 |
0.30 |
0.46 |
| 5 |
A' |
1668 |
1626 |
126.73 |
118.65 |
0.29 |
0.44 |
| 6 |
A' |
1625 |
1583 |
86.56 |
4.97 |
0.75 |
0.86 |
| 7 |
A' |
1455 |
1418 |
5.28 |
11.57 |
0.45 |
0.62 |
| 8 |
A' |
1385 |
1349 |
9.61 |
25.06 |
0.40 |
0.57 |
| 9 |
A' |
1297 |
1263 |
1.55 |
16.62 |
0.26 |
0.42 |
| 10 |
A' |
1177 |
1147 |
31.95 |
14.36 |
0.62 |
0.76 |
| 11 |
A' |
926 |
902 |
24.67 |
1.56 |
0.07 |
0.13 |
| 12 |
A' |
558 |
544 |
7.18 |
8.49 |
0.36 |
0.53 |
| 13 |
A' |
316 |
308 |
10.75 |
1.08 |
0.32 |
0.48 |
| 14 |
A" |
1034 |
1007 |
24.77 |
0.41 |
0.75 |
0.86 |
| 15 |
A" |
1019 |
993 |
24.86 |
5.43 |
0.75 |
0.86 |
| 16 |
A" |
1001 |
975 |
32.78 |
2.02 |
0.75 |
0.86 |
| 17 |
A" |
616 |
601 |
12.94 |
3.27 |
0.75 |
0.86 |
| 18 |
A" |
177 |
173 |
5.53 |
1.35 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13330.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12990.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.132 |
-0.757 |
0.000 |
| C2 |
0.000 |
0.725 |
0.000 |
| C3 |
1.219 |
1.324 |
0.000 |
| O4 |
-1.235 |
-1.357 |
0.000 |
| H5 |
0.819 |
-1.328 |
0.000 |
| H6 |
-0.929 |
1.294 |
0.000 |
| H7 |
1.329 |
2.406 |
0.000 |
| H8 |
2.139 |
0.737 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4876 | 2.4809 | 1.2559 | 1.1097 | 2.1999 | 3.4838 | 2.7186 |
C2 | 1.4876 | | 1.3585 | 2.4210 | 2.2100 | 1.0891 | 2.1428 | 2.1392 | C3 | 2.4809 | 1.3585 | | 3.6349 | 2.6811 | 2.1483 | 1.0877 | 1.0908 | O4 | 1.2559 | 2.4210 | 3.6349 | | 2.0550 | 2.6688 | 4.5537 | 3.9716 | H5 | 1.1097 | 2.2100 | 2.6811 | 2.0550 | | 3.1508 | 3.7679 | 2.4504 | H6 | 2.1999 | 1.0891 | 2.1483 | 2.6688 | 3.1508 | | 2.5167 | 3.1179 | H7 | 3.4838 | 2.1428 | 1.0877 | 4.5537 | 3.7679 | 2.5167 | | 1.8547 | H8 | 2.7186 | 2.1392 | 1.0908 | 3.9716 | 2.4504 | 3.1179 | 1.8547 | |
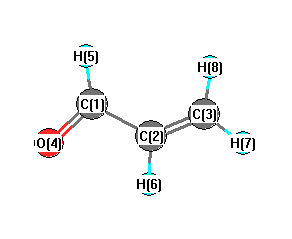 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.247 |
|
C1 |
C2 |
H6 |
116.395 |
| C2 |
C1 |
O4 |
123.659 |
|
C2 |
C1 |
H5 |
115.853 |
| C2 |
C3 |
H7 |
121.942 |
|
C2 |
C3 |
H8 |
121.328 |
| C3 |
C2 |
H6 |
122.358 |
|
O4 |
C1 |
H5 |
120.488 |
| H7 |
C3 |
H8 |
116.730 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.169 |
|
|
|
| 2 |
C |
0.158 |
|
|
|
| 3 |
C |
-0.403 |
|
|
|
| 4 |
O |
-0.153 |
|
|
|
| 5 |
H |
0.091 |
|
|
|
| 6 |
H |
0.171 |
|
|
|
| 7 |
H |
0.150 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.866 |
2.368 |
0.000 |
3.718 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.117 |
-3.286 |
0.000 |
| y |
-3.286 |
-24.484 |
0.000 |
| z |
0.000 |
0.000 |
-25.255 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.247 |
-3.286 |
0.000 |
| y |
-3.286 |
0.702 |
0.000 |
| z |
0.000 |
0.000 |
-0.454 |
|
| Polar |
|---|
| 3z2-r2 | -0.909 |
|---|
| x2-y2 | -0.633 |
|---|
| xy | -3.286 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.471 |
2.083 |
0.000 |
| y |
2.083 |
6.692 |
0.000 |
| z |
0.000 |
0.000 |
2.657 |
<r2> (average value of r
2) Å
2
| <r2> |
70.855 |
| (<r2>)1/2 |
8.418 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -35.243965 |
| Energy at 298.15K | |
| HF Energy | -35.243965 |
| Nuclear repulsion energy | 55.184026 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3226 |
3144 |
9.36 |
57.99 |
0.63 |
0.77 |
| 2 |
A' |
3148 |
3067 |
19.23 |
138.69 |
0.24 |
0.39 |
| 3 |
A' |
3113 |
3034 |
8.54 |
58.94 |
0.17 |
0.29 |
| 4 |
A' |
2950 |
2875 |
136.99 |
168.46 |
0.32 |
0.49 |
| 5 |
A' |
1670 |
1627 |
25.92 |
7.15 |
0.41 |
0.58 |
| 6 |
A' |
1612 |
1571 |
139.25 |
76.28 |
0.26 |
0.41 |
| 7 |
A' |
1440 |
1403 |
45.27 |
15.79 |
0.37 |
0.53 |
| 8 |
A' |
1414 |
1378 |
5.20 |
12.35 |
0.34 |
0.51 |
| 9 |
A' |
1315 |
1282 |
2.80 |
22.10 |
0.38 |
0.55 |
| 10 |
A' |
1075 |
1048 |
7.13 |
5.24 |
0.74 |
0.85 |
| 11 |
A' |
932 |
908 |
56.59 |
7.87 |
0.21 |
0.35 |
| 12 |
A' |
678 |
660 |
11.39 |
1.51 |
0.75 |
0.86 |
| 13 |
A' |
278 |
271 |
7.80 |
4.97 |
0.49 |
0.66 |
| 14 |
A" |
1041 |
1014 |
11.50 |
7.85 |
0.75 |
0.86 |
| 15 |
A" |
1030 |
1003 |
58.20 |
2.29 |
0.75 |
0.86 |
| 16 |
A" |
1010 |
984 |
0.58 |
1.88 |
0.75 |
0.86 |
| 17 |
A" |
572 |
558 |
14.33 |
4.71 |
0.75 |
0.86 |
| 18 |
A" |
186 |
181 |
9.09 |
0.36 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13344.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 13003.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.904 |
-0.281 |
0.000 |
| C2 |
0.000 |
0.911 |
0.000 |
| C3 |
1.353 |
0.789 |
0.000 |
| O4 |
-0.506 |
-1.474 |
0.000 |
| H5 |
-1.990 |
-0.065 |
0.000 |
| H6 |
-0.485 |
1.887 |
0.000 |
| H7 |
2.006 |
1.659 |
0.000 |
| H8 |
1.816 |
-0.197 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4958 | 2.4975 | 1.2572 | 1.1075 | 2.2081 | 3.4974 | 2.7210 |
C2 | 1.4958 | | 1.3585 | 2.4376 | 2.2160 | 1.0899 | 2.1411 | 2.1270 | C3 | 2.4975 | 1.3585 | | 2.9283 | 3.4501 | 2.1409 | 1.0880 | 1.0889 | O4 | 1.2572 | 2.4376 | 2.9283 | | 2.0465 | 3.3607 | 4.0155 | 2.6499 | H5 | 1.1075 | 2.2160 | 3.4501 | 2.0465 | | 2.4643 | 4.3519 | 3.8081 | H6 | 2.2081 | 1.0899 | 2.1409 | 3.3607 | 2.4643 | | 2.5014 | 3.1040 | H7 | 3.4974 | 2.1411 | 1.0880 | 4.0155 | 4.3519 | 2.5014 | | 1.8654 | H8 | 2.7210 | 2.1270 | 1.0889 | 2.6499 | 3.8081 | 3.1040 | 1.8654 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
122.024 |
|
C1 |
C2 |
H6 |
116.414 |
| C2 |
C1 |
O4 |
124.382 |
|
C2 |
C1 |
H5 |
115.892 |
| C2 |
C3 |
H7 |
121.747 |
|
C2 |
C3 |
H8 |
120.310 |
| C3 |
C2 |
H6 |
121.562 |
|
O4 |
C1 |
H5 |
119.726 |
| H7 |
C3 |
H8 |
117.943 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.142 |
|
|
|
| 2 |
C |
0.088 |
|
|
|
| 3 |
C |
-0.356 |
|
|
|
| 4 |
O |
-0.149 |
|
|
|
| 5 |
H |
0.085 |
|
|
|
| 6 |
H |
0.157 |
|
|
|
| 7 |
H |
0.141 |
|
|
|
| 8 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.273 |
3.145 |
0.000 |
3.157 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.926 |
-0.745 |
0.000 |
| y |
-0.745 |
-27.218 |
0.000 |
| z |
0.000 |
0.000 |
-25.209 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.287 |
-0.745 |
0.000 |
| y |
-0.745 |
-4.651 |
0.000 |
| z |
0.000 |
0.000 |
-1.636 |
|
| Polar |
|---|
| 3z2-r2 | -3.273 |
|---|
| x2-y2 | 7.292 |
|---|
| xy | -0.745 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.487 |
0.538 |
0.000 |
| y |
0.538 |
6.082 |
0.000 |
| z |
0.000 |
0.000 |
2.641 |
<r2> (average value of r
2) Å
2
| <r2> |
64.216 |
| (<r2>)1/2 |
8.013 |
