Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -41.984428 |
| Energy at 298.15K | |
| HF Energy | -41.984428 |
| Nuclear repulsion energy | 60.339145 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3096 |
3017 |
15.60 |
16.82 |
0.05 |
0.10 |
| 2 |
A' |
3091 |
3012 |
1.70 |
175.48 |
0.06 |
0.11 |
| 3 |
A' |
1499 |
1461 |
2.07 |
13.88 |
0.74 |
0.85 |
| 4 |
A' |
1496 |
1458 |
7.33 |
4.99 |
0.73 |
0.85 |
| 5 |
A' |
1317 |
1284 |
1.71 |
34.96 |
0.55 |
0.71 |
| 6 |
A' |
1242 |
1210 |
53.30 |
4.72 |
0.40 |
0.57 |
| 7 |
A' |
1069 |
1042 |
1.06 |
26.90 |
0.66 |
0.80 |
| 8 |
A' |
666 |
649 |
23.02 |
104.66 |
0.29 |
0.45 |
| 9 |
A' |
588 |
573 |
88.28 |
11.93 |
0.30 |
0.46 |
| 10 |
A' |
229 |
223 |
1.17 |
10.50 |
0.39 |
0.56 |
| 11 |
A' |
181 |
176 |
11.39 |
0.30 |
0.68 |
0.81 |
| 12 |
A" |
3197 |
3115 |
11.04 |
1.39 |
0.75 |
0.86 |
| 13 |
A" |
3171 |
3090 |
0.29 |
102.47 |
0.75 |
0.86 |
| 14 |
A" |
1290 |
1257 |
0.00 |
9.13 |
0.75 |
0.86 |
| 15 |
A" |
1115 |
1087 |
4.15 |
0.14 |
0.75 |
0.86 |
| 16 |
A" |
971 |
946 |
1.27 |
18.31 |
0.75 |
0.86 |
| 17 |
A" |
781 |
761 |
5.83 |
0.18 |
0.75 |
0.86 |
| 18 |
A" |
106 |
103 |
7.40 |
0.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12551.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12231.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.665 |
0.000 |
| C2 |
1.249 |
-0.210 |
0.000 |
| Br3 |
-1.631 |
-0.559 |
0.000 |
| Cl4 |
2.771 |
0.937 |
0.000 |
| H5 |
-0.092 |
1.273 |
0.900 |
| H6 |
-0.092 |
1.273 |
-0.900 |
| H7 |
1.340 |
-0.820 |
0.899 |
| H8 |
1.340 |
-0.820 |
-0.899 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5255 | 2.0396 | 2.7841 | 1.0898 | 1.0898 | 2.1929 | 2.1929 |
C2 | 1.5255 | | 2.9015 | 1.9058 | 2.1931 | 2.1931 | 1.0900 | 1.0900 | Br3 | 2.0396 | 2.9015 | | 4.6494 | 2.5564 | 2.5564 | 3.1152 | 3.1152 | Cl4 | 2.7841 | 1.9058 | 4.6494 | | 3.0198 | 3.0198 | 2.4376 | 2.4376 | H5 | 1.0898 | 2.1931 | 2.5564 | 3.0198 | | 1.7996 | 2.5358 | 3.1092 | H6 | 1.0898 | 2.1931 | 2.5564 | 3.0198 | 1.7996 | | 3.1092 | 2.5358 | H7 | 2.1929 | 1.0900 | 3.1152 | 2.4376 | 2.5358 | 3.1092 | | 1.7987 | H8 | 2.1929 | 1.0900 | 3.1152 | 2.4376 | 3.1092 | 2.5358 | 1.7987 | |
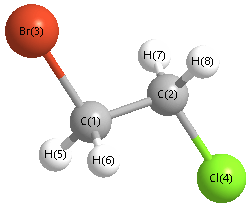 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
107.949 |
|
C1 |
C2 |
H7 |
112.889 |
| C1 |
C2 |
H8 |
112.889 |
|
C2 |
C1 |
Br3 |
108.079 |
| C2 |
C1 |
Cl4 |
40.633 |
|
C2 |
C1 |
H6 |
112.918 |
| Br3 |
C1 |
H5 |
105.497 |
|
Br3 |
C1 |
H6 |
105.497 |
| Cl4 |
C2 |
H7 |
105.665 |
|
Cl4 |
C2 |
H8 |
105.665 |
| H5 |
C1 |
H6 |
111.314 |
|
H7 |
C2 |
H8 |
111.195 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.495 |
|
|
|
| 2 |
C |
-0.415 |
|
|
|
| 3 |
Br |
0.054 |
|
|
|
| 4 |
Cl |
-0.031 |
|
|
|
| 5 |
H |
0.222 |
|
|
|
| 6 |
H |
0.222 |
|
|
|
| 7 |
H |
0.221 |
|
|
|
| 8 |
H |
0.221 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.317 |
-0.175 |
0.000 |
0.362 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.614 |
-3.558 |
0.000 |
| y |
-3.558 |
-40.113 |
0.000 |
| z |
0.000 |
0.000 |
-39.794 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.660 |
-3.558 |
0.000 |
| y |
-3.558 |
4.091 |
0.000 |
| z |
0.000 |
0.000 |
4.569 |
|
| Polar |
|---|
| 3z2-r2 | 9.138 |
|---|
| x2-y2 | -8.501 |
|---|
| xy | -3.558 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.241 |
3.142 |
0.000 |
| y |
3.142 |
6.376 |
0.000 |
| z |
0.000 |
0.000 |
4.106 |
<r2> (average value of r
2) Å
2
| <r2> |
127.079 |
| (<r2>)1/2 |
11.273 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -41.980083 |
| Energy at 298.15K | |
| HF Energy | -41.980083 |
| Nuclear repulsion energy | 61.273661 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3160 |
3079 |
9.08 |
31.67 |
0.75 |
0.85 |
| 2 |
A |
3145 |
3064 |
2.06 |
108.56 |
0.75 |
0.86 |
| 3 |
A |
3066 |
2988 |
25.50 |
250.93 |
0.01 |
0.03 |
| 4 |
A |
3051 |
2973 |
6.93 |
78.76 |
0.47 |
0.64 |
| 5 |
A |
1478 |
1440 |
2.52 |
8.15 |
0.74 |
0.85 |
| 6 |
A |
1470 |
1433 |
12.26 |
18.79 |
0.74 |
0.85 |
| 7 |
A |
1335 |
1301 |
28.29 |
7.52 |
0.70 |
0.82 |
| 8 |
A |
1301 |
1268 |
61.86 |
3.84 |
0.51 |
0.68 |
| 9 |
A |
1199 |
1169 |
2.68 |
29.89 |
0.74 |
0.85 |
| 10 |
A |
1136 |
1107 |
3.26 |
11.99 |
0.74 |
0.85 |
| 11 |
A |
1049 |
1022 |
2.03 |
4.60 |
0.70 |
0.82 |
| 12 |
A |
930 |
906 |
10.53 |
16.76 |
0.48 |
0.65 |
| 13 |
A |
871 |
848 |
26.95 |
3.07 |
0.72 |
0.84 |
| 14 |
A |
616 |
600 |
29.52 |
19.77 |
0.45 |
0.62 |
| 15 |
A |
534 |
520 |
16.50 |
24.43 |
0.21 |
0.35 |
| 16 |
A |
365 |
355 |
11.39 |
4.84 |
0.75 |
0.86 |
| 17 |
A |
233 |
227 |
1.65 |
2.73 |
0.52 |
0.68 |
| 18 |
A |
89 |
86 |
0.81 |
3.48 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 12512.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12193.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.025 |
1.126 |
-0.408 |
| C2 |
1.291 |
0.944 |
0.425 |
| Br3 |
-1.410 |
-0.233 |
0.037 |
| Cl4 |
2.272 |
-0.588 |
-0.086 |
| H5 |
-0.433 |
2.095 |
-0.188 |
| H6 |
0.209 |
1.021 |
-1.478 |
| H7 |
1.972 |
1.785 |
0.258 |
| H8 |
1.083 |
0.826 |
1.489 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5260 | 2.0261 | 2.8436 | 1.0943 | 1.0906 | 2.1603 | 2.1922 |
C2 | 1.5260 | | 2.9718 | 1.8887 | 2.1624 | 2.1905 | 1.0950 | 1.0908 | Br3 | 2.0261 | 2.9718 | | 3.7006 | 2.5344 | 2.5464 | 3.9446 | 3.0733 | Cl4 | 2.8436 | 1.8887 | 3.7006 | | 3.8111 | 2.9632 | 2.4157 | 2.4278 | H5 | 1.0943 | 2.1624 | 2.5344 | 3.8111 | | 1.7970 | 2.4664 | 2.5925 | H6 | 1.0906 | 2.1905 | 2.5464 | 2.9632 | 1.7970 | | 2.5897 | 3.0991 | H7 | 2.1603 | 1.0950 | 3.9446 | 2.4157 | 2.4664 | 2.5897 | | 1.7962 | H8 | 2.1922 | 1.0908 | 3.0733 | 2.4278 | 2.5925 | 3.0991 | 1.7962 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.327 |
|
C1 |
C2 |
H7 |
109.927 |
| C1 |
C2 |
H8 |
112.741 |
|
C2 |
C1 |
Br3 |
112.815 |
| C2 |
C1 |
Cl4 |
37.909 |
|
C2 |
C1 |
H6 |
112.615 |
| Br3 |
C1 |
H5 |
104.641 |
|
Br3 |
C1 |
H6 |
105.617 |
| Cl4 |
C2 |
H7 |
104.983 |
|
Cl4 |
C2 |
H8 |
106.021 |
| H5 |
C1 |
H6 |
110.665 |
|
H7 |
C2 |
H8 |
110.514 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.493 |
|
|
|
| 2 |
C |
-0.414 |
|
|
|
| 3 |
Br |
0.061 |
|
|
|
| 4 |
Cl |
-0.017 |
|
|
|
| 5 |
H |
0.207 |
|
|
|
| 6 |
H |
0.223 |
|
|
|
| 7 |
H |
0.208 |
|
|
|
| 8 |
H |
0.223 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.208 |
3.273 |
0.170 |
3.284 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.205 |
3.411 |
0.633 |
| y |
3.411 |
-36.691 |
-0.128 |
| z |
0.633 |
-0.128 |
-39.330 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.195 |
3.411 |
0.633 |
| y |
3.411 |
5.577 |
-0.128 |
| z |
0.633 |
-0.128 |
1.618 |
|
| Polar |
|---|
| 3z2-r2 | 3.236 |
|---|
| x2-y2 | -8.514 |
|---|
| xy | 3.411 |
|---|
| xz | 0.633 |
|---|
| yz | -0.128 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.356 |
0.583 |
-0.516 |
| y |
0.583 |
7.272 |
-0.012 |
| z |
-0.516 |
-0.012 |
4.568 |
<r2> (average value of r
2) Å
2
| <r2> |
113.922 |
| (<r2>)1/2 |
10.673 |
