Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -43.520741 |
| Energy at 298.15K | -43.525294 |
| Nuclear repulsion energy | 61.505648 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3090 |
3011 |
0.00 |
|
|
|
| 2 |
Ag |
1499 |
1461 |
0.00 |
|
|
|
| 3 |
Ag |
1335 |
1301 |
0.00 |
|
|
|
| 4 |
Ag |
1073 |
1045 |
0.00 |
|
|
|
| 5 |
Ag |
690 |
672 |
0.00 |
|
|
|
| 6 |
Ag |
277 |
270 |
0.00 |
|
|
|
| 7 |
Au |
3194 |
3113 |
12.59 |
|
|
|
| 8 |
Au |
1127 |
1098 |
2.54 |
|
|
|
| 9 |
Au |
789 |
769 |
3.40 |
|
|
|
| 10 |
Au |
111 |
108 |
9.80 |
|
|
|
| 11 |
Bg |
3169 |
3088 |
0.00 |
|
|
|
| 12 |
Bg |
1293 |
1260 |
0.00 |
|
|
|
| 13 |
Bg |
1000 |
975 |
0.00 |
|
|
|
| 14 |
Bu |
3094 |
3015 |
19.08 |
|
|
|
| 15 |
Bu |
1501 |
1463 |
9.10 |
|
|
|
| 16 |
Bu |
1266 |
1234 |
41.29 |
|
|
|
| 17 |
Bu |
647 |
631 |
118.58 |
|
|
|
| 18 |
Bu |
198 |
193 |
15.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12676.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12352.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.763 |
0.000 |
| C2 |
0.000 |
-0.763 |
0.000 |
| Cl3 |
-1.809 |
1.349 |
0.000 |
| Cl4 |
1.809 |
-1.349 |
0.000 |
| H5 |
0.450 |
1.183 |
0.900 |
| H6 |
0.450 |
1.183 |
-0.900 |
| H7 |
-0.450 |
-1.183 |
0.900 |
| H8 |
-0.450 |
-1.183 |
-0.900 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5268 | 1.9015 | 2.7815 | 1.0901 | 1.0901 | 2.1912 | 2.1912 |
C2 | 1.5268 | | 2.7815 | 1.9015 | 2.1912 | 2.1912 | 1.0901 | 1.0901 | Cl3 | 1.9015 | 2.7815 | | 4.5137 | 2.4375 | 2.4375 | 3.0115 | 3.0115 | Cl4 | 2.7815 | 1.9015 | 4.5137 | | 3.0115 | 3.0115 | 2.4375 | 2.4375 | H5 | 1.0901 | 2.1912 | 2.4375 | 3.0115 | | 1.7992 | 2.5320 | 3.1061 | H6 | 1.0901 | 2.1912 | 2.4375 | 3.0115 | 1.7992 | | 3.1061 | 2.5320 | H7 | 2.1912 | 1.0901 | 3.0115 | 2.4375 | 2.5320 | 3.1061 | | 1.7992 | H8 | 2.1912 | 1.0901 | 3.0115 | 2.4375 | 3.1061 | 2.5320 | 1.7992 | |
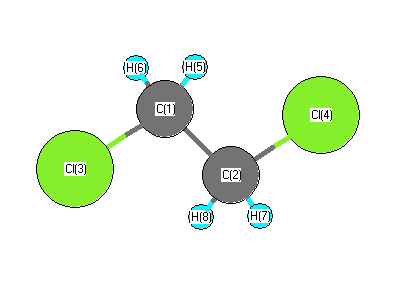 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
107.951 |
|
C1 |
C2 |
H7 |
112.648 |
| C1 |
C2 |
H8 |
112.648 |
|
C2 |
C1 |
Cl3 |
107.951 |
| C2 |
C1 |
H5 |
112.648 |
|
C2 |
C1 |
H6 |
112.648 |
| Cl3 |
C1 |
H5 |
105.922 |
|
Cl3 |
C1 |
H6 |
105.922 |
| Cl4 |
C2 |
H7 |
105.922 |
|
Cl4 |
C2 |
H8 |
105.922 |
| H5 |
C1 |
H6 |
111.230 |
|
H7 |
C2 |
H8 |
111.230 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.419 |
|
|
|
| 2 |
C |
-0.419 |
|
|
|
| 3 |
Cl |
-0.023 |
|
|
|
| 4 |
Cl |
-0.023 |
|
|
|
| 5 |
H |
0.221 |
|
|
|
| 6 |
H |
0.221 |
|
|
|
| 7 |
H |
0.221 |
|
|
|
| 8 |
H |
0.221 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.722 |
5.584 |
0.000 |
| y |
5.584 |
-39.378 |
0.000 |
| z |
0.000 |
0.000 |
-36.518 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.774 |
5.584 |
0.000 |
| y |
5.584 |
0.741 |
0.000 |
| z |
0.000 |
0.000 |
5.033 |
|
| Polar |
|---|
| 3z2-r2 | 10.065 |
|---|
| x2-y2 | -4.344 |
|---|
| xy | 5.584 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.498 |
-2.590 |
0.000 |
| y |
-2.590 |
6.139 |
0.000 |
| z |
0.000 |
0.000 |
4.163 |
<r2> (average value of r
2) Å
2
| <r2> |
110.520 |
| (<r2>)1/2 |
10.513 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -43.517003 |
| Energy at 298.15K | -43.521704 |
| HF Energy | -43.517003 |
| Nuclear repulsion energy | 62.472885 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
3065 |
1.79 |
|
|
|
| 2 |
A |
3066 |
2988 |
27.95 |
|
|
|
| 3 |
A |
1477 |
1439 |
1.58 |
|
|
|
| 4 |
A |
1342 |
1308 |
21.82 |
|
|
|
| 5 |
A |
1215 |
1184 |
0.78 |
|
|
|
| 6 |
A |
1056 |
1029 |
1.83 |
|
|
|
| 7 |
A |
947 |
923 |
10.74 |
|
|
|
| 8 |
A |
606 |
590 |
23.52 |
|
|
|
| 9 |
A |
251 |
244 |
1.22 |
|
|
|
| 10 |
A |
102 |
100 |
1.41 |
|
|
|
| 11 |
B |
3159 |
3078 |
10.49 |
|
|
|
| 12 |
B |
3053 |
2975 |
5.08 |
|
|
|
| 13 |
B |
1473 |
1436 |
13.96 |
|
|
|
| 14 |
B |
1326 |
1292 |
48.28 |
|
|
|
| 15 |
B |
1151 |
1121 |
1.89 |
|
|
|
| 16 |
B |
892 |
869 |
20.59 |
|
|
|
| 17 |
B |
626 |
610 |
34.80 |
|
|
|
| 18 |
B |
387 |
377 |
12.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12636.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12314.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.318 |
0.693 |
0.930 |
| C2 |
-0.318 |
-0.693 |
0.930 |
| Cl3 |
-0.318 |
1.770 |
-0.485 |
| Cl4 |
0.318 |
-1.770 |
-0.485 |
| H5 |
0.045 |
1.230 |
1.844 |
| H6 |
1.402 |
0.655 |
0.816 |
| H7 |
-0.045 |
-1.230 |
1.844 |
| H8 |
-1.402 |
-0.655 |
0.816 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5253 | 1.8886 | 2.8408 | 1.0946 | 1.0909 | 2.1604 | 2.1887 |
C2 | 1.5253 | | 2.8408 | 1.8886 | 2.1604 | 2.1887 | 1.0946 | 1.0909 | Cl3 | 1.8886 | 2.8408 | | 3.5972 | 2.4179 | 2.4279 | 3.8081 | 2.9580 | Cl4 | 2.8408 | 1.8886 | 3.5972 | | 3.8081 | 2.9580 | 2.4179 | 2.4279 | H5 | 1.0946 | 2.1604 | 2.4179 | 3.8081 | | 1.7971 | 2.4626 | 2.5899 | H6 | 1.0909 | 2.1887 | 2.4279 | 2.9580 | 1.7971 | | 2.5899 | 3.0955 | H7 | 2.1604 | 1.0946 | 3.8081 | 2.4179 | 2.4626 | 2.5899 | | 1.7971 | H8 | 2.1887 | 1.0909 | 2.9580 | 2.4279 | 2.5899 | 3.0955 | 1.7971 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.203 |
|
C1 |
C2 |
H7 |
110.010 |
| C1 |
C2 |
H8 |
112.508 |
|
C2 |
C1 |
Cl3 |
112.203 |
| C2 |
C1 |
H5 |
110.010 |
|
C2 |
C1 |
H6 |
112.508 |
| Cl3 |
C1 |
H5 |
105.160 |
|
Cl3 |
C1 |
H6 |
106.033 |
| Cl4 |
C2 |
H7 |
105.160 |
|
Cl4 |
C2 |
H8 |
106.033 |
| H5 |
C1 |
H6 |
110.627 |
|
H7 |
C2 |
H8 |
110.627 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.417 |
|
|
|
| 2 |
C |
-0.417 |
|
|
|
| 3 |
Cl |
-0.015 |
|
|
|
| 4 |
Cl |
-0.015 |
|
|
|
| 5 |
H |
0.208 |
|
|
|
| 6 |
H |
0.224 |
|
|
|
| 7 |
H |
0.208 |
|
|
|
| 8 |
H |
0.224 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
3.490 |
3.490 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.414 |
1.718 |
0.000 |
| y |
1.718 |
-42.788 |
0.000 |
| z |
0.000 |
0.000 |
-34.203 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.082 |
1.718 |
0.000 |
| y |
1.718 |
-7.479 |
0.000 |
| z |
0.000 |
0.000 |
5.397 |
|
| Polar |
|---|
| 3z2-r2 | 10.795 |
|---|
| x2-y2 | 6.374 |
|---|
| xy | 1.718 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.731 |
-0.828 |
0.000 |
| y |
-0.828 |
7.211 |
0.000 |
| z |
0.000 |
0.000 |
6.873 |
<r2> (average value of r
2) Å
2
| <r2> |
99.719 |
| (<r2>)1/2 |
9.986 |
