Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -43.932676 |
| Energy at 298.15K | -43.935928 |
| HF Energy | -43.932676 |
| Nuclear repulsion energy | 57.261785 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3165 |
3084 |
4.01 |
|
|
|
| 2 |
A |
3060 |
2982 |
8.31 |
|
|
|
| 3 |
A |
2987 |
2911 |
60.93 |
|
|
|
| 4 |
A |
1672 |
1630 |
125.75 |
|
|
|
| 5 |
A |
1463 |
1426 |
12.21 |
|
|
|
| 6 |
A |
1402 |
1366 |
7.63 |
|
|
|
| 7 |
A |
1279 |
1246 |
20.89 |
|
|
|
| 8 |
A |
1180 |
1150 |
8.59 |
|
|
|
| 9 |
A |
1050 |
1024 |
25.10 |
|
|
|
| 10 |
A |
1013 |
987 |
3.14 |
|
|
|
| 11 |
A |
809 |
788 |
9.40 |
|
|
|
| 12 |
A |
671 |
654 |
31.79 |
|
|
|
| 13 |
A |
442 |
430 |
13.06 |
|
|
|
| 14 |
A |
276 |
269 |
12.60 |
|
|
|
| 15 |
A |
72 |
70 |
16.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10270.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 10008.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.054 |
0.731 |
0.137 |
| C2 |
1.188 |
-0.281 |
0.287 |
| Cl3 |
-1.581 |
-0.189 |
-0.089 |
| O4 |
2.287 |
-0.130 |
-0.284 |
| H5 |
-0.064 |
1.328 |
1.046 |
| H6 |
0.204 |
1.366 |
-0.737 |
| H7 |
0.990 |
-1.147 |
0.943 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5273 | 1.8897 | 2.4300 | 1.0947 | 1.0904 | 2.2479 |
C2 | 1.5273 | | 2.7963 | 1.2476 | 2.1761 | 2.1744 | 1.1050 | Cl3 | 1.8897 | 2.7963 | | 3.8735 | 2.4274 | 2.4540 | 2.9313 | O4 | 2.4300 | 1.2476 | 3.8735 | | 3.0703 | 2.6040 | 2.0558 | H5 | 1.0947 | 2.1761 | 2.4274 | 3.0703 | | 1.8041 | 2.6924 | H6 | 1.0904 | 2.1744 | 2.4540 | 2.6040 | 1.8041 | | 3.1233 | H7 | 2.2479 | 1.1050 | 2.9313 | 2.0558 | 2.6924 | 3.1233 | |
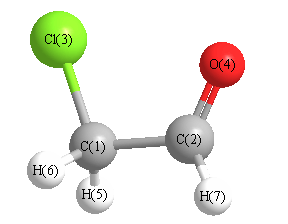 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
121.929 |
|
C1 |
C2 |
H7 |
116.369 |
| C2 |
C1 |
Cl3 |
109.383 |
|
C2 |
C1 |
H5 |
111.108 |
| C2 |
C1 |
H6 |
111.236 |
|
Cl3 |
C1 |
H5 |
105.750 |
| Cl3 |
C1 |
H6 |
107.838 |
|
O4 |
C2 |
H7 |
121.694 |
| H5 |
C1 |
H6 |
111.309 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.483 |
|
|
|
| 2 |
C |
0.052 |
|
|
|
| 3 |
Cl |
-0.016 |
|
|
|
| 4 |
O |
-0.138 |
|
|
|
| 5 |
H |
0.227 |
|
|
|
| 6 |
H |
0.226 |
|
|
|
| 7 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.679 |
1.035 |
1.476 |
1.926 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.296 |
-0.093 |
2.173 |
| y |
-0.093 |
-27.795 |
-0.125 |
| z |
2.173 |
-0.125 |
-28.905 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.946 |
-0.093 |
2.173 |
| y |
-0.093 |
6.805 |
-0.125 |
| z |
2.173 |
-0.125 |
5.141 |
|
| Polar |
|---|
| 3z2-r2 | 10.282 |
|---|
| x2-y2 | -12.501 |
|---|
| xy | -0.093 |
|---|
| xz | 2.173 |
|---|
| yz | -0.125 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.421 |
0.718 |
-0.303 |
| y |
0.718 |
4.355 |
-0.230 |
| z |
-0.303 |
-0.230 |
3.694 |
<r2> (average value of r
2) Å
2
| <r2> |
86.983 |
| (<r2>)1/2 |
9.326 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -43.929400 |
| Energy at 298.15K | -43.932761 |
| HF Energy | -43.929400 |
| Nuclear repulsion energy | 58.157924 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3038 |
2961 |
18.95 |
|
|
|
| 2 |
A' |
2910 |
2836 |
86.76 |
|
|
|
| 3 |
A' |
1698 |
1654 |
117.36 |
|
|
|
| 4 |
A' |
1456 |
1419 |
31.17 |
|
|
|
| 5 |
A' |
1400 |
1364 |
17.39 |
|
|
|
| 6 |
A' |
1300 |
1267 |
19.91 |
|
|
|
| 7 |
A' |
944 |
920 |
3.05 |
|
|
|
| 8 |
A' |
742 |
723 |
5.59 |
|
|
|
| 9 |
A' |
584 |
569 |
48.18 |
|
|
|
| 10 |
A' |
203 |
197 |
2.24 |
|
|
|
| 11 |
A" |
3109 |
3029 |
5.44 |
|
|
|
| 12 |
A" |
1193 |
1163 |
4.98 |
|
|
|
| 13 |
A" |
1014 |
988 |
0.58 |
|
|
|
| 14 |
A" |
708 |
690 |
4.49 |
|
|
|
| 15 |
A" |
145 |
141 |
1.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10220.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 9960.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.957 |
0.000 |
| C2 |
1.394 |
0.332 |
0.000 |
| Cl3 |
-1.388 |
-0.291 |
0.000 |
| O4 |
1.659 |
-0.881 |
0.000 |
| H5 |
-0.124 |
1.583 |
0.892 |
| H6 |
-0.124 |
1.583 |
-0.892 |
| H7 |
2.204 |
1.093 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5278 | 1.8664 | 2.4763 | 1.0961 | 1.0961 | 2.2085 |
C2 | 1.5278 | | 2.8506 | 1.2421 | 2.1594 | 2.1594 | 1.1117 | Cl3 | 1.8664 | 2.8506 | | 3.1030 | 2.4293 | 2.4293 | 3.8494 | O4 | 2.4763 | 1.2421 | 3.1030 | | 3.1692 | 3.1692 | 2.0488 | H5 | 1.0961 | 2.1594 | 2.4293 | 3.1692 | | 1.7836 | 2.5408 | H6 | 1.0961 | 2.1594 | 2.4293 | 3.1692 | 1.7836 | | 2.5408 | H7 | 2.2085 | 1.1117 | 3.8494 | 2.0488 | 2.5408 | 2.5408 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
126.454 |
|
C1 |
C2 |
H7 |
112.633 |
| C2 |
C1 |
Cl3 |
113.874 |
|
C2 |
C1 |
H5 |
109.670 |
| C2 |
C1 |
H6 |
109.670 |
|
Cl3 |
C1 |
H5 |
107.296 |
| Cl3 |
C1 |
H6 |
107.296 |
|
O4 |
C2 |
H7 |
120.913 |
| H5 |
C1 |
H6 |
108.896 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.510 |
|
|
|
| 2 |
C |
0.056 |
|
|
|
| 3 |
Cl |
0.016 |
|
|
|
| 4 |
O |
-0.114 |
|
|
|
| 5 |
H |
0.220 |
|
|
|
| 6 |
H |
0.220 |
|
|
|
| 7 |
H |
0.113 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.692 |
4.112 |
0.000 |
4.170 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.870 |
4.146 |
0.000 |
| y |
4.146 |
-29.236 |
0.000 |
| z |
0.000 |
0.000 |
-28.949 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.778 |
4.146 |
0.000 |
| y |
4.146 |
1.673 |
0.000 |
| z |
0.000 |
0.000 |
2.104 |
|
| Polar |
|---|
| 3z2-r2 | 4.208 |
|---|
| x2-y2 | -3.634 |
|---|
| xy | 4.146 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.172 |
0.874 |
0.000 |
| y |
0.874 |
5.935 |
0.000 |
| z |
0.000 |
0.000 |
3.126 |
<r2> (average value of r
2) Å
2
| <r2> |
78.762 |
| (<r2>)1/2 |
8.875 |
