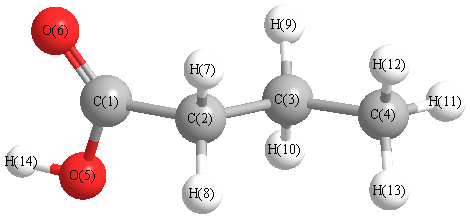Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3620 |
3527 |
25.65 |
|
|
|
| 2 |
A |
3104 |
3025 |
45.61 |
|
|
|
| 3 |
A |
3074 |
2996 |
50.07 |
|
|
|
| 4 |
A |
3072 |
2994 |
55.31 |
|
|
|
| 5 |
A |
3047 |
2969 |
0.86 |
|
|
|
| 6 |
A |
3026 |
2949 |
24.05 |
|
|
|
| 7 |
A |
3002 |
2925 |
17.69 |
|
|
|
| 8 |
A |
2982 |
2906 |
42.30 |
|
|
|
| 9 |
A |
1684 |
1641 |
278.38 |
|
|
|
| 10 |
A |
1533 |
1494 |
11.26 |
|
|
|
| 11 |
A |
1522 |
1483 |
9.13 |
|
|
|
| 12 |
A |
1519 |
1480 |
2.04 |
|
|
|
| 13 |
A |
1498 |
1460 |
6.41 |
|
|
|
| 14 |
A |
1434 |
1398 |
4.22 |
|
|
|
| 15 |
A |
1381 |
1346 |
7.81 |
|
|
|
| 16 |
A |
1350 |
1315 |
44.39 |
|
|
|
| 17 |
A |
1322 |
1288 |
5.90 |
|
|
|
| 18 |
A |
1306 |
1272 |
13.62 |
|
|
|
| 19 |
A |
1257 |
1224 |
1.23 |
|
|
|
| 20 |
A |
1177 |
1147 |
78.56 |
|
|
|
| 21 |
A |
1116 |
1088 |
10.15 |
|
|
|
| 22 |
A |
1066 |
1039 |
133.94 |
|
|
|
| 23 |
A |
1041 |
1014 |
45.86 |
|
|
|
| 24 |
A |
910 |
887 |
2.28 |
|
|
|
| 25 |
A |
896 |
873 |
5.82 |
|
|
|
| 26 |
A |
844 |
822 |
23.72 |
|
|
|
| 27 |
A |
762 |
743 |
3.26 |
|
|
|
| 28 |
A |
688 |
671 |
53.31 |
|
|
|
| 29 |
A |
600 |
585 |
114.87 |
|
|
|
| 30 |
A |
549 |
535 |
50.98 |
|
|
|
| 31 |
A |
413 |
402 |
3.69 |
|
|
|
| 32 |
A |
325 |
317 |
1.89 |
|
|
|
| 33 |
A |
233 |
227 |
0.04 |
|
|
|
| 34 |
A |
183 |
178 |
0.12 |
|
|
|
| 35 |
A |
91 |
88 |
0.25 |
|
|
|
| 36 |
A |
42 |
41 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25832.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 25174.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.092 |
|
|
|
| 2 |
C |
-0.305 |
|
|
|
| 3 |
C |
-0.233 |
|
|
|
| 4 |
C |
-0.473 |
|
|
|
| 5 |
O |
-0.378 |
|
|
|
| 6 |
O |
-0.152 |
|
|
|
| 7 |
H |
0.164 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
H |
0.143 |
|
|
|
| 10 |
H |
0.161 |
|
|
|
| 11 |
H |
0.156 |
|
|
|
| 12 |
H |
0.147 |
|
|
|
| 13 |
H |
0.145 |
|
|
|
| 14 |
H |
0.374 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.458 |
1.316 |
0.312 |
1.988 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.522 |
6.125 |
0.239 |
| y |
6.125 |
-40.627 |
-1.863 |
| z |
0.239 |
-1.863 |
-36.705 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.144 |
6.125 |
0.239 |
| y |
6.125 |
-4.013 |
-1.863 |
| z |
0.239 |
-1.863 |
1.870 |
|
| Polar |
|---|
| 3z2-r2 | 3.739 |
|---|
| x2-y2 | 4.105 |
|---|
| xy | 6.125 |
|---|
| xz | 0.239 |
|---|
| yz | -1.863 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.287 |
-0.146 |
-0.357 |
| y |
-0.146 |
7.771 |
0.114 |
| z |
-0.357 |
0.114 |
6.402 |
<r2> (average value of r
2) Å
2
| <r2> |
174.918 |
| (<r2>)1/2 |
13.226 |