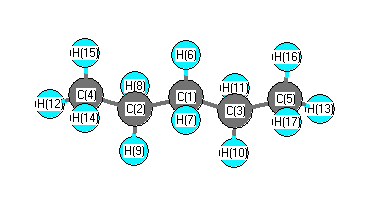
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3063 |
2984 |
90.92 |
|
|
|
| 2 |
A1 |
2978 |
2902 |
0.00 |
|
|
|
| 3 |
A1 |
2974 |
2898 |
166.59 |
|
|
|
| 4 |
A1 |
2957 |
2882 |
6.32 |
|
|
|
| 5 |
A1 |
1538 |
1499 |
11.47 |
|
|
|
| 6 |
A1 |
1520 |
1481 |
0.69 |
|
|
|
| 7 |
A1 |
1512 |
1473 |
0.00 |
|
|
|
| 8 |
A1 |
1429 |
1393 |
4.59 |
|
|
|
| 9 |
A1 |
1372 |
1337 |
0.19 |
|
|
|
| 10 |
A1 |
1161 |
1132 |
1.55 |
|
|
|
| 11 |
A1 |
1046 |
1019 |
0.65 |
|
|
|
| 12 |
A1 |
869 |
847 |
1.33 |
|
|
|
| 13 |
A1 |
390 |
380 |
0.00 |
|
|
|
| 14 |
A1 |
174 |
170 |
0.02 |
|
|
|
| 15 |
A2 |
3059 |
2981 |
0.00 |
|
|
|
| 16 |
A2 |
3005 |
2928 |
0.00 |
|
|
|
| 17 |
A2 |
1521 |
1482 |
0.00 |
|
|
|
| 18 |
A2 |
1327 |
1293 |
0.00 |
|
|
|
| 19 |
A2 |
1269 |
1236 |
0.00 |
|
|
|
| 20 |
A2 |
999 |
973 |
0.00 |
|
|
|
| 21 |
A2 |
774 |
754 |
0.00 |
|
|
|
| 22 |
A2 |
233 |
227 |
0.00 |
|
|
|
| 23 |
A2 |
104 |
102 |
0.00 |
|
|
|
| 24 |
B1 |
3063 |
2985 |
229.44 |
|
|
|
| 25 |
B1 |
3027 |
2950 |
44.49 |
|
|
|
| 26 |
B1 |
2987 |
2911 |
0.07 |
|
|
|
| 27 |
B1 |
1520 |
1481 |
16.46 |
|
|
|
| 28 |
B1 |
1323 |
1290 |
0.64 |
|
|
|
| 29 |
B1 |
1215 |
1184 |
0.06 |
|
|
|
| 30 |
B1 |
875 |
853 |
1.56 |
|
|
|
| 31 |
B1 |
739 |
720 |
4.90 |
|
|
|
| 32 |
B1 |
225 |
220 |
0.00 |
|
|
|
| 33 |
B1 |
109 |
106 |
0.00 |
|
|
|
| 34 |
B2 |
3062 |
2984 |
51.02 |
|
|
|
| 35 |
B2 |
2975 |
2899 |
61.53 |
|
|
|
| 36 |
B2 |
2965 |
2890 |
7.34 |
|
|
|
| 37 |
B2 |
1528 |
1490 |
2.19 |
|
|
|
| 38 |
B2 |
1514 |
1475 |
1.51 |
|
|
|
| 39 |
B2 |
1429 |
1393 |
3.83 |
|
|
|
| 40 |
B2 |
1394 |
1358 |
2.59 |
|
|
|
| 41 |
B2 |
1302 |
1269 |
1.51 |
|
|
|
| 42 |
B2 |
1083 |
1056 |
2.03 |
|
|
|
| 43 |
B2 |
1039 |
1013 |
2.32 |
|
|
|
| 44 |
B2 |
922 |
899 |
3.38 |
|
|
|
| 45 |
B2 |
390 |
380 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34979.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 34087.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.230 |
|
|
|
| 2 |
C |
-0.190 |
|
|
|
| 3 |
C |
-0.190 |
|
|
|
| 4 |
C |
-0.472 |
|
|
|
| 5 |
C |
-0.472 |
|
|
|
| 6 |
H |
0.113 |
|
|
|
| 7 |
H |
0.113 |
|
|
|
| 8 |
H |
0.121 |
|
|
|
| 9 |
H |
0.121 |
|
|
|
| 10 |
H |
0.121 |
|
|
|
| 11 |
H |
0.121 |
|
|
|
| 12 |
H |
0.144 |
|
|
|
| 13 |
H |
0.144 |
|
|
|
| 14 |
H |
0.139 |
|
|
|
| 15 |
H |
0.139 |
|
|
|
| 16 |
H |
0.139 |
|
|
|
| 17 |
H |
0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.067 |
0.067 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.861 |
0.000 |
0.000 |
| y |
0.000 |
-36.234 |
0.000 |
| z |
0.000 |
0.000 |
-35.902 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.206 |
0.000 |
0.000 |
| y |
0.000 |
-0.852 |
0.000 |
| z |
0.000 |
0.000 |
-0.355 |
|
| Polar |
|---|
| 3z2-r2 | -0.709 |
|---|
| x2-y2 | 1.372 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.236 |
0.000 |
0.000 |
| y |
0.000 |
10.436 |
0.000 |
| z |
0.000 |
0.000 |
8.505 |
<r2> (average value of r
2) Å
2
| <r2> |
171.196 |
| (<r2>)1/2 |
13.084 |