Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -34.209838 |
| Energy at 298.15K | -34.220000 |
| Nuclear repulsion energy | 95.074686 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3196 |
3114 |
41.06 |
|
|
|
| 2 |
A |
3100 |
3021 |
9.09 |
|
|
|
| 3 |
A |
3085 |
3006 |
54.55 |
|
|
|
| 4 |
A |
3066 |
2988 |
65.88 |
|
|
|
| 5 |
A |
3065 |
2987 |
97.55 |
|
|
|
| 6 |
A |
3033 |
2956 |
17.21 |
|
|
|
| 7 |
A |
3012 |
2935 |
7.27 |
|
|
|
| 8 |
A |
2981 |
2905 |
22.20 |
|
|
|
| 9 |
A |
2976 |
2900 |
82.82 |
|
|
|
| 10 |
A |
2961 |
2885 |
26.81 |
|
|
|
| 11 |
A |
1672 |
1629 |
15.11 |
|
|
|
| 12 |
A |
1532 |
1493 |
9.97 |
|
|
|
| 13 |
A |
1521 |
1482 |
8.16 |
|
|
|
| 14 |
A |
1517 |
1478 |
1.08 |
|
|
|
| 15 |
A |
1507 |
1469 |
2.93 |
|
|
|
| 16 |
A |
1461 |
1424 |
1.87 |
|
|
|
| 17 |
A |
1428 |
1392 |
4.29 |
|
|
|
| 18 |
A |
1379 |
1344 |
1.20 |
|
|
|
| 19 |
A |
1329 |
1295 |
0.48 |
|
|
|
| 20 |
A |
1319 |
1285 |
0.66 |
|
|
|
| 21 |
A |
1306 |
1273 |
1.34 |
|
|
|
| 22 |
A |
1268 |
1236 |
0.15 |
|
|
|
| 23 |
A |
1201 |
1171 |
0.55 |
|
|
|
| 24 |
A |
1116 |
1088 |
1.27 |
|
|
|
| 25 |
A |
1057 |
1030 |
4.70 |
|
|
|
| 26 |
A |
1035 |
1008 |
7.50 |
|
|
|
| 27 |
A |
1024 |
998 |
9.93 |
|
|
|
| 28 |
A |
956 |
932 |
58.88 |
|
|
|
| 29 |
A |
949 |
925 |
1.57 |
|
|
|
| 30 |
A |
888 |
865 |
7.42 |
|
|
|
| 31 |
A |
880 |
858 |
0.77 |
|
|
|
| 32 |
A |
756 |
737 |
3.02 |
|
|
|
| 33 |
A |
639 |
623 |
11.69 |
|
|
|
| 34 |
A |
433 |
422 |
1.25 |
|
|
|
| 35 |
A |
378 |
368 |
0.22 |
|
|
|
| 36 |
A |
239 |
233 |
0.03 |
|
|
|
| 37 |
A |
224 |
218 |
0.09 |
|
|
|
| 38 |
A |
98 |
95 |
0.04 |
|
|
|
| 39 |
A |
91 |
88 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29837.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 29076.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.518 |
-0.221 |
-0.305 |
| C2 |
1.434 |
0.349 |
0.274 |
| C3 |
0.075 |
-0.318 |
0.432 |
| C4 |
-1.072 |
0.458 |
-0.285 |
| C5 |
-2.459 |
-0.202 |
-0.078 |
| H6 |
3.464 |
0.310 |
-0.395 |
| H7 |
2.485 |
-1.236 |
-0.702 |
| H8 |
1.513 |
1.371 |
0.658 |
| H9 |
0.120 |
-1.348 |
0.045 |
| H10 |
-0.174 |
-0.392 |
1.505 |
| H11 |
-1.097 |
1.494 |
0.089 |
| H12 |
-0.844 |
0.520 |
-1.360 |
| H13 |
-3.249 |
0.362 |
-0.593 |
| H14 |
-2.471 |
-1.229 |
-0.469 |
| H15 |
-2.720 |
-0.248 |
0.989 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3543 | 2.5535 | 3.6534 | 4.9826 | 1.0888 | 1.0908 | 2.1143 | 2.6727 | 3.2481 | 4.0201 | 3.6004 | 5.8034 | 5.0922 | 5.3959 |
C2 | 1.3543 | | 1.5222 | 2.5699 | 3.9480 | 2.1381 | 2.1373 | 1.0947 | 2.1581 | 2.1563 | 2.7842 | 2.8085 | 4.7627 | 4.2767 | 4.2576 | C3 | 2.5535 | 1.5222 | | 1.5596 | 2.5879 | 3.5451 | 2.8169 | 2.2297 | 1.1005 | 1.1038 | 2.1854 | 2.1812 | 3.5444 | 2.8499 | 2.8513 | C4 | 3.6534 | 2.5699 | 1.5596 | | 1.5505 | 4.5401 | 3.9615 | 2.8987 | 2.1888 | 2.1757 | 1.1017 | 1.1003 | 2.2009 | 2.1996 | 2.2000 | C5 | 4.9826 | 3.9480 | 2.5879 | 1.5505 | | 5.9545 | 5.0897 | 4.3351 | 2.8252 | 2.7866 | 2.1820 | 2.1851 | 1.0986 | 1.0995 | 1.0997 | H6 | 1.0888 | 2.1381 | 3.5451 | 4.5401 | 5.9545 | | 1.8558 | 2.4584 | 3.7587 | 4.1642 | 4.7373 | 4.4200 | 6.7166 | 6.1320 | 6.3624 | H7 | 1.0908 | 2.1373 | 2.8169 | 3.9615 | 5.0897 | 1.8558 | | 3.0968 | 2.4826 | 3.5566 | 4.5724 | 3.8206 | 5.9533 | 4.9610 | 5.5616 | H8 | 2.1143 | 1.0947 | 2.2297 | 2.8987 | 4.3351 | 2.4584 | 3.0968 | | 3.1155 | 2.5829 | 2.6737 | 3.2168 | 5.0255 | 4.8886 | 4.5441 | H9 | 2.6727 | 2.1581 | 1.1005 | 2.1888 | 2.8252 | 3.7587 | 2.4826 | 3.1155 | | 1.7689 | 3.0917 | 2.5282 | 3.8316 | 2.6438 | 3.1889 | H10 | 3.2481 | 2.1563 | 1.1038 | 2.1757 | 2.7866 | 4.1642 | 3.5566 | 2.5829 | 1.7689 | | 2.5330 | 3.0801 | 3.7983 | 3.1419 | 2.6022 | H11 | 4.0201 | 2.7842 | 2.1854 | 1.1017 | 2.1820 | 4.7373 | 4.5724 | 2.6737 | 3.0917 | 2.5330 | | 1.7638 | 2.5255 | 3.1010 | 2.5458 | H12 | 3.6004 | 2.8085 | 2.1812 | 1.1003 | 2.1851 | 4.4200 | 3.8206 | 3.2168 | 2.5282 | 3.0801 | 1.7638 | | 2.5293 | 2.5497 | 3.1031 | H13 | 5.8034 | 4.7627 | 3.5444 | 2.2009 | 1.0986 | 6.7166 | 5.9533 | 5.0255 | 3.8316 | 3.7983 | 2.5255 | 2.5293 | | 1.7760 | 1.7764 | H14 | 5.0922 | 4.2767 | 2.8499 | 2.1996 | 1.0995 | 6.1320 | 4.9610 | 4.8886 | 2.6438 | 3.1419 | 3.1010 | 2.5497 | 1.7760 | | 1.7757 | H15 | 5.3959 | 4.2576 | 2.8513 | 2.2000 | 1.0997 | 6.3624 | 5.5616 | 4.5441 | 3.1889 | 2.6022 | 2.5458 | 3.1031 | 1.7764 | 1.7757 | |
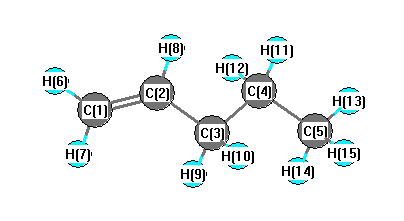 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.077 |
|
C1 |
C2 |
H8 |
119.011 |
| C2 |
C1 |
H6 |
121.756 |
|
C2 |
C1 |
H7 |
121.508 |
| C2 |
C3 |
C4 |
112.996 |
|
C2 |
C3 |
H9 |
109.693 |
| C2 |
C3 |
H10 |
109.361 |
|
C3 |
C2 |
H8 |
115.910 |
| C3 |
C4 |
C5 |
112.627 |
|
C3 |
C4 |
H11 |
109.193 |
| C3 |
C4 |
H12 |
108.950 |
|
C4 |
C3 |
H9 |
109.524 |
| C4 |
C3 |
H10 |
108.332 |
|
C4 |
C5 |
H13 |
111.221 |
| C4 |
C5 |
H14 |
111.064 |
|
C4 |
C5 |
H15 |
111.085 |
| C5 |
C4 |
H11 |
109.552 |
|
C5 |
C4 |
H12 |
109.873 |
| H6 |
C1 |
H7 |
116.736 |
|
H9 |
C3 |
H10 |
106.734 |
| H11 |
C4 |
H12 |
106.452 |
|
H13 |
C5 |
H14 |
107.799 |
| H13 |
C5 |
H15 |
107.820 |
|
H14 |
C5 |
H15 |
107.690 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.414 |
|
|
|
| 2 |
C |
0.147 |
|
|
|
| 3 |
C |
-0.369 |
|
|
|
| 4 |
C |
-0.206 |
|
|
|
| 5 |
C |
-0.473 |
|
|
|
| 6 |
H |
0.116 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.131 |
|
|
|
| 10 |
H |
0.129 |
|
|
|
| 11 |
H |
0.127 |
|
|
|
| 12 |
H |
0.133 |
|
|
|
| 13 |
H |
0.147 |
|
|
|
| 14 |
H |
0.141 |
|
|
|
| 15 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.431 |
0.094 |
0.085 |
0.449 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.200 |
0.183 |
-0.597 |
| y |
0.183 |
-32.999 |
1.220 |
| z |
-0.597 |
1.220 |
-34.713 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.344 |
0.183 |
-0.597 |
| y |
0.183 |
1.458 |
1.220 |
| z |
-0.597 |
1.220 |
-1.114 |
|
| Polar |
|---|
| 3z2-r2 | -2.228 |
|---|
| x2-y2 | -1.201 |
|---|
| xy | 0.183 |
|---|
| xz | -0.597 |
|---|
| yz | 1.220 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.393 |
-0.475 |
-0.972 |
| y |
-0.475 |
8.014 |
0.580 |
| z |
-0.972 |
0.580 |
7.242 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -34.209270 |
| Energy at 298.15K | -34.219498 |
| HF Energy | -34.209270 |
| Nuclear repulsion energy | 96.778355 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3194 |
3113 |
39.86 |
|
|
|
| 2 |
A |
3098 |
3019 |
7.78 |
|
|
|
| 3 |
A |
3081 |
3003 |
55.95 |
|
|
|
| 4 |
A |
3076 |
2998 |
63.91 |
|
|
|
| 5 |
A |
3062 |
2984 |
68.67 |
|
|
|
| 6 |
A |
3026 |
2949 |
40.05 |
|
|
|
| 7 |
A |
3014 |
2937 |
34.89 |
|
|
|
| 8 |
A |
2982 |
2906 |
16.90 |
|
|
|
| 9 |
A |
2975 |
2899 |
75.33 |
|
|
|
| 10 |
A |
2961 |
2886 |
33.19 |
|
|
|
| 11 |
A |
1672 |
1630 |
12.61 |
|
|
|
| 12 |
A |
1530 |
1491 |
5.47 |
|
|
|
| 13 |
A |
1520 |
1482 |
11.75 |
|
|
|
| 14 |
A |
1514 |
1476 |
1.02 |
|
|
|
| 15 |
A |
1506 |
1468 |
6.32 |
|
|
|
| 16 |
A |
1462 |
1424 |
2.44 |
|
|
|
| 17 |
A |
1431 |
1395 |
7.42 |
|
|
|
| 18 |
A |
1380 |
1345 |
3.88 |
|
|
|
| 19 |
A |
1355 |
1320 |
1.09 |
|
|
|
| 20 |
A |
1322 |
1288 |
0.61 |
|
|
|
| 21 |
A |
1290 |
1257 |
0.23 |
|
|
|
| 22 |
A |
1259 |
1227 |
1.75 |
|
|
|
| 23 |
A |
1196 |
1166 |
0.42 |
|
|
|
| 24 |
A |
1101 |
1073 |
5.01 |
|
|
|
| 25 |
A |
1077 |
1050 |
2.62 |
|
|
|
| 26 |
A |
1031 |
1005 |
13.23 |
|
|
|
| 27 |
A |
1017 |
992 |
2.90 |
|
|
|
| 28 |
A |
957 |
932 |
63.74 |
|
|
|
| 29 |
A |
929 |
905 |
0.88 |
|
|
|
| 30 |
A |
892 |
869 |
0.45 |
|
|
|
| 31 |
A |
857 |
835 |
3.25 |
|
|
|
| 32 |
A |
773 |
753 |
4.40 |
|
|
|
| 33 |
A |
638 |
622 |
11.42 |
|
|
|
| 34 |
A |
431 |
420 |
0.49 |
|
|
|
| 35 |
A |
384 |
375 |
1.71 |
|
|
|
| 36 |
A |
290 |
283 |
0.14 |
|
|
|
| 37 |
A |
211 |
206 |
0.08 |
|
|
|
| 38 |
A |
113 |
110 |
0.02 |
|
|
|
| 39 |
A |
85 |
83 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29847.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 29085.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.149 |
0.574 |
0.121 |
| C2 |
-1.206 |
-0.240 |
-0.408 |
| C3 |
-0.075 |
-0.901 |
0.371 |
| C4 |
1.345 |
-0.532 |
-0.163 |
| C5 |
1.712 |
0.962 |
0.027 |
| H6 |
-2.936 |
1.012 |
-0.491 |
| H7 |
-2.160 |
0.821 |
1.183 |
| H8 |
-1.233 |
-0.458 |
-1.481 |
| H9 |
-0.193 |
-1.996 |
0.311 |
| H10 |
-0.151 |
-0.630 |
1.435 |
| H11 |
2.087 |
-1.160 |
0.354 |
| H12 |
1.408 |
-0.794 |
-1.232 |
| H13 |
2.718 |
1.173 |
-0.361 |
| H14 |
1.001 |
1.616 |
-0.495 |
| H15 |
1.697 |
1.238 |
1.092 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3539 | 2.5568 | 3.6764 | 3.8816 | 1.0890 | 1.0907 | 2.1141 | 3.2351 | 2.6775 | 4.5832 | 4.0444 | 4.9275 | 3.3749 | 4.0216 |
C2 | 1.3539 | | 1.5229 | 2.5793 | 3.1853 | 2.1380 | 2.1376 | 1.0951 | 2.1504 | 2.1588 | 3.5024 | 2.7959 | 4.1707 | 2.8848 | 3.5855 | C3 | 2.5568 | 1.5229 | | 1.5619 | 2.6042 | 3.5476 | 2.8230 | 2.2282 | 1.1032 | 1.1008 | 2.1778 | 2.1863 | 3.5550 | 2.8706 | 2.8692 | C4 | 3.6764 | 2.5793 | 1.5619 | | 1.5500 | 4.5636 | 3.9914 | 2.8971 | 2.1761 | 2.1916 | 1.1010 | 1.1022 | 2.1978 | 2.2003 | 2.1976 | C5 | 3.8816 | 3.1853 | 2.6042 | 1.5500 | | 4.6774 | 4.0434 | 3.6008 | 3.5298 | 2.8264 | 2.1801 | 2.1821 | 1.0987 | 1.0977 | 1.0999 | H6 | 1.0890 | 2.1380 | 3.5476 | 4.5636 | 4.6774 | | 1.8553 | 2.4579 | 4.1491 | 3.7634 | 5.5377 | 4.7632 | 5.6583 | 3.9838 | 4.9011 | H7 | 1.0907 | 2.1376 | 2.8230 | 3.9914 | 4.0434 | 1.8553 | | 3.0971 | 3.5443 | 2.4909 | 4.7590 | 4.6015 | 5.1287 | 3.6666 | 3.8803 | H8 | 2.1141 | 1.0951 | 2.2282 | 2.8971 | 3.6008 | 2.4579 | 3.0971 | | 2.5804 | 3.1150 | 3.8582 | 2.6746 | 4.4193 | 3.2040 | 4.2519 | H9 | 3.2351 | 2.1504 | 1.1032 | 2.1761 | 3.5298 | 4.1491 | 3.5443 | 2.5804 | | 1.7690 | 2.4286 | 2.5277 | 4.3554 | 3.8885 | 3.8258 | H10 | 2.6775 | 2.1588 | 1.1008 | 2.1916 | 2.8264 | 3.7634 | 2.4909 | 3.1150 | 1.7690 | | 2.5410 | 3.0935 | 3.8352 | 3.1778 | 2.6498 | H11 | 4.5832 | 3.5024 | 2.1778 | 1.1010 | 2.1801 | 5.5377 | 4.7590 | 3.8582 | 2.4286 | 2.5410 | | 1.7637 | 2.5211 | 3.0996 | 2.5390 | H12 | 4.0444 | 2.7959 | 2.1863 | 1.1022 | 2.1821 | 4.7632 | 4.6015 | 2.6746 | 2.5277 | 3.0935 | 1.7637 | | 2.5189 | 2.5526 | 3.1000 | H13 | 4.9275 | 4.1707 | 3.5550 | 2.1978 | 1.0987 | 5.6583 | 5.1287 | 4.4193 | 4.3554 | 3.8352 | 2.5211 | 2.5189 | | 1.7779 | 1.7768 | H14 | 3.3749 | 2.8848 | 2.8706 | 2.2003 | 1.0977 | 3.9838 | 3.6666 | 3.2040 | 3.8885 | 3.1778 | 3.0996 | 2.5526 | 1.7779 | | 1.7734 | H15 | 4.0216 | 3.5855 | 2.8692 | 2.1976 | 1.0999 | 4.9011 | 3.8803 | 4.2519 | 3.8258 | 2.6498 | 2.5390 | 3.1000 | 1.7768 | 1.7734 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.327 |
|
C1 |
C2 |
H8 |
118.982 |
| C2 |
C1 |
H6 |
121.754 |
|
C2 |
C1 |
H7 |
121.570 |
| C2 |
C3 |
C4 |
113.462 |
|
C2 |
C3 |
H9 |
108.887 |
| C2 |
C3 |
H10 |
109.681 |
|
C3 |
C2 |
H8 |
115.691 |
| C3 |
C4 |
C5 |
113.618 |
|
C3 |
C4 |
H11 |
108.494 |
| C3 |
C4 |
H12 |
109.080 |
|
C4 |
C3 |
H9 |
108.248 |
| C4 |
C3 |
H10 |
109.570 |
|
C4 |
C5 |
H13 |
111.000 |
| C4 |
C5 |
H14 |
111.258 |
|
C4 |
C5 |
H15 |
110.912 |
| C5 |
C4 |
H11 |
109.477 |
|
C5 |
C4 |
H12 |
109.558 |
| H6 |
C1 |
H7 |
116.675 |
|
H9 |
C3 |
H10 |
106.763 |
| H11 |
C4 |
H12 |
106.354 |
|
H13 |
C5 |
H14 |
108.087 |
| H13 |
C5 |
H15 |
107.827 |
|
H14 |
C5 |
H15 |
107.606 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.415 |
|
|
|
| 2 |
C |
0.117 |
|
|
|
| 3 |
C |
-0.341 |
|
|
|
| 4 |
C |
-0.214 |
|
|
|
| 5 |
C |
-0.476 |
|
|
|
| 6 |
H |
0.117 |
|
|
|
| 7 |
H |
0.129 |
|
|
|
| 8 |
H |
0.119 |
|
|
|
| 9 |
H |
0.140 |
|
|
|
| 10 |
H |
0.134 |
|
|
|
| 11 |
H |
0.134 |
|
|
|
| 12 |
H |
0.127 |
|
|
|
| 13 |
H |
0.142 |
|
|
|
| 14 |
H |
0.153 |
|
|
|
| 15 |
H |
0.136 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.343 |
-0.244 |
-0.058 |
0.425 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.500 |
-1.313 |
0.392 |
| y |
-1.313 |
-34.519 |
0.431 |
| z |
0.392 |
0.431 |
-32.527 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.978 |
-1.313 |
0.392 |
| y |
-1.313 |
-1.006 |
0.431 |
| z |
0.392 |
0.431 |
1.983 |
|
| Polar |
|---|
| 3z2-r2 | 3.967 |
|---|
| x2-y2 | 0.019 |
|---|
| xy | -1.313 |
|---|
| xz | 0.392 |
|---|
| yz | 0.431 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.148 |
-1.393 |
-0.351 |
| y |
-1.393 |
8.237 |
0.479 |
| z |
-0.351 |
0.479 |
7.995 |
<r2> (average value of r
2) Å
2
| <r2> |
130.887 |
| (<r2>)1/2 |
11.441 |
